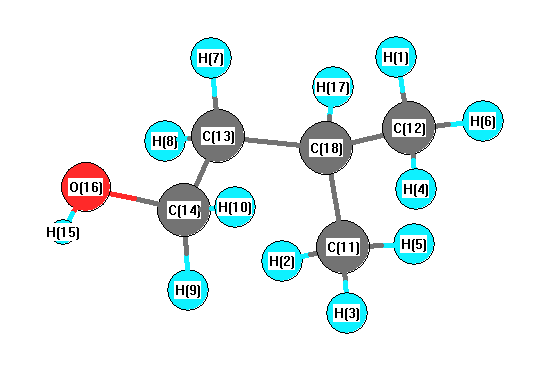
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3892 |
3729 |
32.49 |
|
|
|
| 2 |
A' |
3133 |
3003 |
49.05 |
|
|
|
| 3 |
A' |
3129 |
2999 |
75.57 |
|
|
|
| 4 |
A' |
3060 |
2933 |
31.23 |
|
|
|
| 5 |
A' |
3051 |
2924 |
32.77 |
|
|
|
| 6 |
A' |
3042 |
2915 |
11.50 |
|
|
|
| 7 |
A' |
3020 |
2895 |
40.24 |
|
|
|
| 8 |
A' |
1506 |
1443 |
1.36 |
|
|
|
| 9 |
A' |
1491 |
1429 |
12.78 |
|
|
|
| 10 |
A' |
1490 |
1428 |
6.65 |
|
|
|
| 11 |
A' |
1475 |
1414 |
2.87 |
|
|
|
| 12 |
A' |
1445 |
1385 |
2.91 |
|
|
|
| 13 |
A' |
1409 |
1350 |
4.49 |
|
|
|
| 14 |
A' |
1361 |
1305 |
1.16 |
|
|
|
| 15 |
A' |
1338 |
1282 |
30.59 |
|
|
|
| 16 |
A' |
1249 |
1197 |
20.13 |
|
|
|
| 17 |
A' |
1206 |
1156 |
4.86 |
|
|
|
| 18 |
A' |
1124 |
1077 |
0.60 |
|
|
|
| 19 |
A' |
1083 |
1037 |
103.86 |
|
|
|
| 20 |
A' |
1008 |
966 |
7.09 |
|
|
|
| 21 |
A' |
970 |
930 |
8.17 |
|
|
|
| 22 |
A' |
816 |
782 |
4.53 |
|
|
|
| 23 |
A' |
560 |
537 |
1.36 |
|
|
|
| 24 |
A' |
406 |
389 |
6.09 |
|
|
|
| 25 |
A' |
328 |
314 |
5.10 |
|
|
|
| 26 |
A' |
249 |
239 |
0.11 |
|
|
|
| 27 |
A' |
207 |
198 |
1.95 |
|
|
|
| 28 |
A" |
3132 |
3001 |
20.53 |
|
|
|
| 29 |
A" |
3124 |
2994 |
2.49 |
|
|
|
| 30 |
A" |
3107 |
2977 |
46.64 |
|
|
|
| 31 |
A" |
3060 |
2933 |
40.96 |
|
|
|
| 32 |
A" |
3046 |
2919 |
19.43 |
|
|
|
| 33 |
A" |
1478 |
1417 |
0.97 |
|
|
|
| 34 |
A" |
1465 |
1404 |
0.45 |
|
|
|
| 35 |
A" |
1393 |
1335 |
11.01 |
|
|
|
| 36 |
A" |
1358 |
1302 |
0.04 |
|
|
|
| 37 |
A" |
1312 |
1257 |
0.03 |
|
|
|
| 38 |
A" |
1243 |
1191 |
0.02 |
|
|
|
| 39 |
A" |
1179 |
1129 |
3.39 |
|
|
|
| 40 |
A" |
1011 |
969 |
0.22 |
|
|
|
| 41 |
A" |
963 |
923 |
0.01 |
|
|
|
| 42 |
A" |
919 |
881 |
0.60 |
|
|
|
| 43 |
A" |
787 |
754 |
0.15 |
|
|
|
| 44 |
A" |
368 |
352 |
0.00 |
|
|
|
| 45 |
A" |
293 |
281 |
103.39 |
|
|
|
| 46 |
A" |
227 |
218 |
0.78 |
|
|
|
| 47 |
A" |
102 |
98 |
0.02 |
|
|
|
| 48 |
A" |
59 |
57 |
9.52 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36337.0 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 34821.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
-0.141 |
|
|
|
| 2 |
H |
-0.141 |
|
|
|
| 3 |
H |
-0.101 |
|
|
|
| 4 |
H |
-0.101 |
|
|
|
| 5 |
H |
-0.182 |
|
|
|
| 6 |
H |
-0.182 |
|
|
|
| 7 |
H |
-0.246 |
|
|
|
| 8 |
H |
-0.246 |
|
|
|
| 9 |
H |
-0.296 |
|
|
|
| 10 |
H |
-0.296 |
|
|
|
| 11 |
C |
0.855 |
|
|
|
| 12 |
C |
0.855 |
|
|
|
| 13 |
C |
0.561 |
|
|
|
| 14 |
C |
1.142 |
|
|
|
| 15 |
H |
0.118 |
|
|
|
| 16 |
O |
-0.548 |
|
|
|
| 17 |
H |
-0.268 |
|
|
|
| 18 |
C |
-0.784 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.177 |
-1.430 |
0.000 |
1.441 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.367 |
2.983 |
0.000 |
| y |
2.983 |
-44.232 |
0.000 |
| z |
0.000 |
0.000 |
-40.610 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
7.054 |
2.983 |
0.000 |
| y |
2.983 |
-6.244 |
0.000 |
| z |
0.000 |
0.000 |
-0.810 |
|
| Polar |
|---|
| 3z2-r2 | -1.621 |
|---|
| x2-y2 | 8.865 |
|---|
| xy | 2.983 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.756 |
-0.896 |
0.000 |
| y |
-0.896 |
10.053 |
0.000 |
| z |
0.000 |
0.000 |
9.957 |
<r2> (average value of r
2) Å
2
| <r2> |
223.244 |
| (<r2>)1/2 |
14.941 |