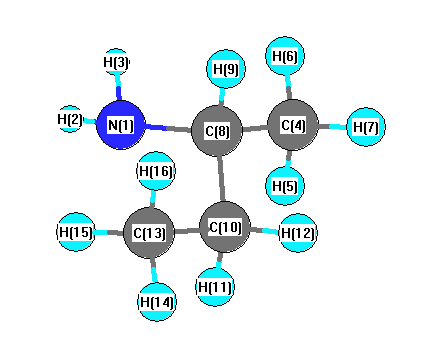
Vibrational Frequencies calculated at HF/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3810 |
3470 |
1.14 |
|
|
|
| 2 |
A |
3731 |
3398 |
0.23 |
|
|
|
| 3 |
A |
3241 |
2951 |
82.40 |
|
|
|
| 4 |
A |
3232 |
2944 |
53.75 |
|
|
|
| 5 |
A |
3225 |
2937 |
71.33 |
|
|
|
| 6 |
A |
3223 |
2935 |
36.03 |
|
|
|
| 7 |
A |
3199 |
2913 |
6.88 |
|
|
|
| 8 |
A |
3167 |
2884 |
25.55 |
|
|
|
| 9 |
A |
3159 |
2877 |
47.04 |
|
|
|
| 10 |
A |
3158 |
2875 |
34.03 |
|
|
|
| 11 |
A |
3083 |
2807 |
69.83 |
|
|
|
| 12 |
A |
1784 |
1624 |
46.59 |
|
|
|
| 13 |
A |
1605 |
1462 |
6.87 |
|
|
|
| 14 |
A |
1598 |
1455 |
0.26 |
|
|
|
| 15 |
A |
1596 |
1453 |
7.34 |
|
|
|
| 16 |
A |
1590 |
1448 |
2.67 |
|
|
|
| 17 |
A |
1582 |
1440 |
0.17 |
|
|
|
| 18 |
A |
1550 |
1411 |
4.60 |
|
|
|
| 19 |
A |
1521 |
1385 |
12.21 |
|
|
|
| 20 |
A |
1514 |
1379 |
7.11 |
|
|
|
| 21 |
A |
1492 |
1358 |
11.71 |
|
|
|
| 22 |
A |
1440 |
1311 |
3.43 |
|
|
|
| 23 |
A |
1394 |
1269 |
1.96 |
|
|
|
| 24 |
A |
1348 |
1228 |
1.63 |
|
|
|
| 25 |
A |
1282 |
1167 |
3.73 |
|
|
|
| 26 |
A |
1255 |
1143 |
13.12 |
|
|
|
| 27 |
A |
1136 |
1034 |
3.79 |
|
|
|
| 28 |
A |
1097 |
999 |
4.73 |
|
|
|
| 29 |
A |
1087 |
990 |
2.52 |
|
|
|
| 30 |
A |
1049 |
956 |
3.28 |
|
|
|
| 31 |
A |
998 |
909 |
4.44 |
|
|
|
| 32 |
A |
932 |
849 |
107.57 |
|
|
|
| 33 |
A |
863 |
786 |
8.48 |
|
|
|
| 34 |
A |
821 |
748 |
0.76 |
|
|
|
| 35 |
A |
513 |
467 |
7.04 |
|
|
|
| 36 |
A |
483 |
440 |
5.27 |
|
|
|
| 37 |
A |
402 |
366 |
0.63 |
|
|
|
| 38 |
A |
311 |
283 |
32.72 |
|
|
|
| 39 |
A |
278 |
253 |
0.01 |
|
|
|
| 40 |
A |
247 |
225 |
0.24 |
|
|
|
| 41 |
A |
241 |
219 |
15.97 |
|
|
|
| 42 |
A |
115 |
104 |
1.55 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 34675.1 cm
-1
Scaled (by 0.9106) Zero Point Vibrational Energy (zpe) 31575.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.293 |
|
|
|
| 2 |
H |
-0.038 |
|
|
|
| 3 |
H |
-0.044 |
|
|
|
| 4 |
C |
0.747 |
|
|
|
| 5 |
H |
0.005 |
|
|
|
| 6 |
H |
-0.092 |
|
|
|
| 7 |
H |
-0.067 |
|
|
|
| 8 |
C |
-0.387 |
|
|
|
| 9 |
H |
-0.194 |
|
|
|
| 10 |
C |
0.445 |
|
|
|
| 11 |
H |
-0.140 |
|
|
|
| 12 |
H |
-0.129 |
|
|
|
| 13 |
C |
0.436 |
|
|
|
| 14 |
H |
-0.082 |
|
|
|
| 15 |
H |
-0.090 |
|
|
|
| 16 |
H |
-0.077 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.046 |
0.045 |
1.244 |
1.245 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.772 |
0.363 |
1.115 |
| y |
0.363 |
-33.014 |
2.013 |
| z |
1.115 |
2.013 |
-36.070 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.770 |
0.363 |
1.115 |
| y |
0.363 |
1.907 |
2.013 |
| z |
1.115 |
2.013 |
-2.676 |
|
| Polar |
|---|
| 3z2-r2 | -5.353 |
|---|
| x2-y2 | -0.758 |
|---|
| xy | 0.363 |
|---|
| xz | 1.115 |
|---|
| yz | 2.013 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.460 |
-0.077 |
-0.028 |
| y |
-0.077 |
8.514 |
-0.009 |
| z |
-0.028 |
-0.009 |
7.783 |
<r2> (average value of r
2) Å
2
| <r2> |
151.406 |
| (<r2>)1/2 |
12.305 |