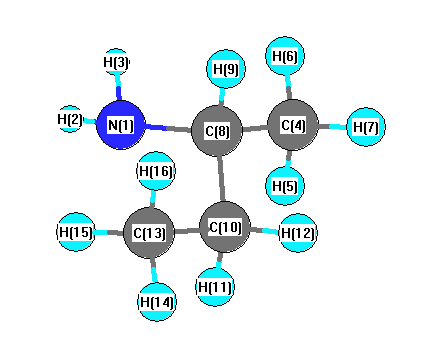
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3617 |
3466 |
0.65 |
|
|
|
| 2 |
A |
3533 |
3385 |
0.91 |
|
|
|
| 3 |
A |
3144 |
3013 |
35.21 |
|
|
|
| 4 |
A |
3135 |
3004 |
37.17 |
|
|
|
| 5 |
A |
3129 |
2998 |
30.77 |
|
|
|
| 6 |
A |
3128 |
2998 |
41.28 |
|
|
|
| 7 |
A |
3097 |
2968 |
7.62 |
|
|
|
| 8 |
A |
3054 |
2926 |
40.40 |
|
|
|
| 9 |
A |
3049 |
2922 |
7.96 |
|
|
|
| 10 |
A |
3048 |
2921 |
37.80 |
|
|
|
| 11 |
A |
2949 |
2826 |
68.95 |
|
|
|
| 12 |
A |
1643 |
1574 |
41.85 |
|
|
|
| 13 |
A |
1487 |
1425 |
8.74 |
|
|
|
| 14 |
A |
1479 |
1418 |
1.13 |
|
|
|
| 15 |
A |
1477 |
1415 |
8.53 |
|
|
|
| 16 |
A |
1470 |
1409 |
4.39 |
|
|
|
| 17 |
A |
1457 |
1397 |
0.69 |
|
|
|
| 18 |
A |
1420 |
1361 |
5.64 |
|
|
|
| 19 |
A |
1393 |
1335 |
15.74 |
|
|
|
| 20 |
A |
1389 |
1331 |
7.11 |
|
|
|
| 21 |
A |
1369 |
1312 |
10.40 |
|
|
|
| 22 |
A |
1321 |
1266 |
1.32 |
|
|
|
| 23 |
A |
1289 |
1236 |
1.92 |
|
|
|
| 24 |
A |
1252 |
1200 |
0.82 |
|
|
|
| 25 |
A |
1198 |
1148 |
4.08 |
|
|
|
| 26 |
A |
1159 |
1111 |
7.33 |
|
|
|
| 27 |
A |
1081 |
1036 |
0.92 |
|
|
|
| 28 |
A |
1040 |
996 |
3.65 |
|
|
|
| 29 |
A |
1013 |
971 |
4.61 |
|
|
|
| 30 |
A |
974 |
933 |
2.59 |
|
|
|
| 31 |
A |
949 |
910 |
6.53 |
|
|
|
| 32 |
A |
851 |
815 |
92.82 |
|
|
|
| 33 |
A |
817 |
783 |
10.11 |
|
|
|
| 34 |
A |
766 |
734 |
3.05 |
|
|
|
| 35 |
A |
479 |
459 |
6.58 |
|
|
|
| 36 |
A |
453 |
434 |
5.70 |
|
|
|
| 37 |
A |
374 |
358 |
1.46 |
|
|
|
| 38 |
A |
282 |
270 |
30.99 |
|
|
|
| 39 |
A |
252 |
241 |
0.25 |
|
|
|
| 40 |
A |
223 |
214 |
1.96 |
|
|
|
| 41 |
A |
219 |
209 |
11.70 |
|
|
|
| 42 |
A |
104 |
100 |
1.48 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 32780.3 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 31413.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.115 |
|
|
|
| 2 |
H |
-0.042 |
|
|
|
| 3 |
H |
-0.039 |
|
|
|
| 4 |
C |
0.679 |
|
|
|
| 5 |
H |
-0.044 |
|
|
|
| 6 |
H |
-0.163 |
|
|
|
| 7 |
H |
-0.127 |
|
|
|
| 8 |
C |
-0.022 |
|
|
|
| 9 |
H |
-0.362 |
|
|
|
| 10 |
C |
0.767 |
|
|
|
| 11 |
H |
-0.276 |
|
|
|
| 12 |
H |
-0.275 |
|
|
|
| 13 |
C |
0.433 |
|
|
|
| 14 |
H |
-0.150 |
|
|
|
| 15 |
H |
-0.142 |
|
|
|
| 16 |
H |
-0.123 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.059 |
0.046 |
1.193 |
1.195 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.465 |
0.353 |
1.078 |
| y |
0.353 |
-32.826 |
1.972 |
| z |
1.078 |
1.972 |
-35.971 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.933 |
0.353 |
1.078 |
| y |
0.353 |
1.892 |
1.972 |
| z |
1.078 |
1.972 |
-2.825 |
|
| Polar |
|---|
| 3z2-r2 | -5.650 |
|---|
| x2-y2 | -0.639 |
|---|
| xy | 0.353 |
|---|
| xz | 1.078 |
|---|
| yz | 1.972 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.219 |
-0.082 |
-0.041 |
| y |
-0.082 |
9.167 |
-0.012 |
| z |
-0.041 |
-0.012 |
8.402 |
<r2> (average value of r
2) Å
2
| <r2> |
150.710 |
| (<r2>)1/2 |
12.276 |