Jump to
S1C2
Energy calculated at B2PLYP=FULLultrafine/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -454.549119 |
| Energy at 298.15K | |
| HF Energy | -454.434698 |
| Nuclear repulsion energy | 56.910504 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULLultrafine/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3506 |
3374 |
5.53 |
|
|
|
| 2 |
A' |
2671 |
2570 |
7.97 |
|
|
|
| 3 |
A' |
1606 |
1546 |
12.27 |
|
|
|
| 4 |
A' |
1026 |
987 |
10.73 |
|
|
|
| 5 |
A' |
884 |
850 |
44.59 |
|
|
|
| 6 |
A' |
658 |
633 |
49.13 |
|
|
|
| 7 |
A" |
3606 |
3470 |
21.32 |
|
|
|
| 8 |
A" |
1125 |
1083 |
0.45 |
|
|
|
| 9 |
A" |
396 |
381 |
48.13 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 7739.3 cm
-1
Scaled (by 0.9623) Zero Point Vibrational Energy (zpe) 7447.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULLultrafine/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
0.013 |
1.129 |
0.000 |
| S2 |
0.013 |
-0.627 |
0.000 |
| H3 |
-1.333 |
-0.777 |
0.000 |
| H4 |
0.514 |
1.454 |
0.824 |
| H5 |
0.514 |
1.454 |
-0.824 |
Atom - Atom Distances (Å)
| |
N1 |
S2 |
H3 |
H4 |
H5 |
| N1 | | 1.7568 | 2.3336 | 1.0172 | 1.0172 |
S2 | 1.7568 | | 1.3547 | 2.2942 | 2.2942 | H3 | 2.3336 | 1.3547 | | 3.0113 | 3.0113 | H4 | 1.0172 | 2.2942 | 3.0113 | | 1.6473 | H5 | 1.0172 | 2.2942 | 3.0113 | 1.6473 | |
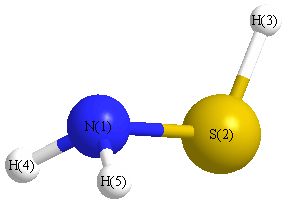 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
S2 |
H3 |
96.321 |
|
S2 |
N1 |
H4 |
108.637 |
| S2 |
N1 |
H5 |
108.637 |
|
H4 |
N1 |
H5 |
108.126 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B2PLYP=FULLultrafine/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -454.548370 |
| Energy at 298.15K | |
| HF Energy | -454.434295 |
| Nuclear repulsion energy | 57.109438 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULLultrafine/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3510 |
3377 |
2.26 |
|
|
|
| 2 |
A' |
2601 |
2503 |
29.08 |
|
|
|
| 3 |
A' |
1594 |
1534 |
13.14 |
|
|
|
| 4 |
A' |
1002 |
964 |
22.76 |
|
|
|
| 5 |
A' |
870 |
837 |
25.65 |
|
|
|
| 6 |
A' |
634 |
610 |
84.39 |
|
|
|
| 7 |
A" |
3620 |
3483 |
21.31 |
|
|
|
| 8 |
A" |
1110 |
1068 |
0.69 |
|
|
|
| 9 |
A" |
485 |
467 |
2.77 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 7712.2 cm
-1
Scaled (by 0.9623) Zero Point Vibrational Energy (zpe) 7421.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULLultrafine/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
0.086 |
1.121 |
0.000 |
| S2 |
0.086 |
-0.620 |
0.000 |
| H3 |
-1.246 |
-0.907 |
0.000 |
| H4 |
-0.368 |
1.491 |
0.830 |
| H5 |
-0.368 |
1.491 |
-0.830 |
Atom - Atom Distances (Å)
| |
N1 |
S2 |
H3 |
H4 |
H5 |
| N1 | | 1.7407 | 2.4266 | 1.0163 | 1.0163 |
S2 | 1.7407 | | 1.3630 | 2.3132 | 2.3132 | H3 | 2.4266 | 1.3630 | | 2.6852 | 2.6852 | H4 | 1.0163 | 2.3132 | 2.6852 | | 1.6605 | H5 | 1.0163 | 2.3132 | 2.6852 | 1.6605 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
S2 |
H3 |
102.173 |
|
S2 |
N1 |
H4 |
111.345 |
| S2 |
N1 |
H5 |
111.345 |
|
H4 |
N1 |
H5 |
109.567 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
