Jump to
S1C2
S1C3
Vibrational Frequencies calculated at HF/aug-cc-pVDZ
Geometric Data calculated at HF/aug-cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Vibrational Frequencies calculated at HF/aug-cc-pVDZ
Geometric Data calculated at HF/aug-cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at HF/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -209.144685 |
| Energy at 298.15K | -209.153945 |
| HF Energy | -209.144685 |
| Nuclear repulsion energy | 134.639616 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
4134 |
3764 |
69.21 |
|
|
|
| 2 |
A |
3814 |
3473 |
5.00 |
|
|
|
| 3 |
A |
3728 |
3395 |
1.04 |
|
|
|
| 4 |
A |
3237 |
2948 |
61.64 |
|
|
|
| 5 |
A |
3223 |
2935 |
42.12 |
|
|
|
| 6 |
A |
3141 |
2860 |
99.07 |
|
|
|
| 7 |
A |
3126 |
2847 |
53.81 |
|
|
|
| 8 |
A |
1779 |
1620 |
34.67 |
|
|
|
| 9 |
A |
1618 |
1473 |
0.62 |
|
|
|
| 10 |
A |
1605 |
1462 |
4.80 |
|
|
|
| 11 |
A |
1565 |
1426 |
45.53 |
|
|
|
| 12 |
A |
1519 |
1384 |
28.91 |
|
|
|
| 13 |
A |
1473 |
1341 |
9.61 |
|
|
|
| 14 |
A |
1426 |
1298 |
3.56 |
|
|
|
| 15 |
A |
1350 |
1230 |
32.39 |
|
|
|
| 16 |
A |
1270 |
1156 |
8.69 |
|
|
|
| 17 |
A |
1214 |
1105 |
74.94 |
|
|
|
| 18 |
A |
1154 |
1051 |
31.55 |
|
|
|
| 19 |
A |
1082 |
985 |
15.87 |
|
|
|
| 20 |
A |
1001 |
911 |
59.75 |
|
|
|
| 21 |
A |
937 |
854 |
22.46 |
|
|
|
| 22 |
A |
885 |
806 |
83.87 |
|
|
|
| 23 |
A |
564 |
513 |
27.22 |
|
|
|
| 24 |
A |
485 |
442 |
132.70 |
|
|
|
| 25 |
A |
342 |
311 |
2.47 |
|
|
|
| 26 |
A |
276 |
251 |
10.78 |
|
|
|
| 27 |
A |
187 |
171 |
4.82 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23067.0 cm
-1
Scaled (by 0.9106) Zero Point Vibrational Energy (zpe) 21004.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/aug-cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
-1.399 |
-0.555 |
0.118 |
| C2 |
-0.637 |
0.624 |
-0.283 |
| C3 |
0.772 |
0.540 |
0.282 |
| O4 |
1.452 |
-0.596 |
-0.172 |
| H5 |
-1.675 |
-0.500 |
1.081 |
| H6 |
-2.234 |
-0.657 |
-0.425 |
| H7 |
-1.095 |
1.566 |
0.035 |
| H8 |
-0.577 |
0.634 |
-1.371 |
| H9 |
1.350 |
1.409 |
-0.031 |
| H10 |
0.730 |
0.548 |
1.378 |
| H11 |
0.864 |
-1.333 |
-0.107 |
Atom - Atom Distances (Å)
| |
N1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| N1 | | 1.4598 | 2.4366 | 2.8658 | 1.0032 | 1.0014 | 2.1445 | 2.0745 | 3.3810 | 2.7081 | 2.4029 |
C2 | 1.4598 | | 1.5198 | 2.4215 | 2.0493 | 2.0524 | 1.0956 | 1.0897 | 2.1507 | 2.1519 | 2.4721 | C3 | 2.4366 | 1.5198 | | 1.4006 | 2.7755 | 3.3118 | 2.1446 | 2.1360 | 1.0889 | 1.0963 | 1.9156 | O4 | 2.8658 | 2.4215 | 1.4006 | | 3.3698 | 3.6952 | 3.3481 | 2.6588 | 2.0124 | 2.0576 | 0.9452 | H5 | 1.0032 | 2.0493 | 2.7755 | 3.3698 | | 1.6139 | 2.3869 | 2.9154 | 3.7448 | 2.6394 | 2.9235 | H6 | 1.0014 | 2.0524 | 3.3118 | 3.6952 | 1.6139 | | 2.5401 | 2.3032 | 4.1552 | 3.6722 | 3.1865 | H7 | 2.1445 | 1.0956 | 2.1446 | 3.3481 | 2.3869 | 2.5401 | | 1.7654 | 2.4512 | 2.4839 | 3.5021 | H8 | 2.0745 | 1.0897 | 2.1360 | 2.6588 | 2.9154 | 2.3032 | 1.7654 | | 2.4723 | 3.0450 | 2.7467 | H9 | 3.3810 | 2.1507 | 1.0889 | 2.0124 | 3.7448 | 4.1552 | 2.4512 | 2.4723 | | 1.7631 | 2.7854 | H10 | 2.7081 | 2.1519 | 1.0963 | 2.0576 | 2.6394 | 3.6722 | 2.4839 | 3.0450 | 1.7631 | | 2.3998 | H11 | 2.4029 | 2.4721 | 1.9156 | 0.9452 | 2.9235 | 3.1865 | 3.5021 | 2.7467 | 2.7854 | 2.3998 | |
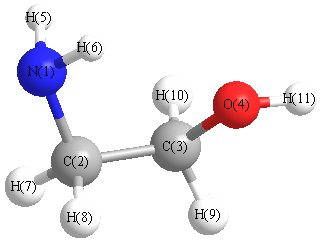 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
C2 |
C3 |
109.712 |
|
N1 |
C2 |
H7 |
113.348 |
| N1 |
C2 |
H8 |
108.040 |
|
C2 |
N1 |
H5 |
111.267 |
| C2 |
N1 |
H6 |
111.652 |
|
C2 |
C3 |
O4 |
111.967 |
| C2 |
C3 |
H9 |
109.962 |
|
C2 |
C3 |
H10 |
109.622 |
| C3 |
C2 |
H7 |
109.095 |
|
C3 |
C2 |
H8 |
108.763 |
| C3 |
O4 |
H11 |
107.912 |
|
O4 |
C3 |
H9 |
107.208 |
| O4 |
C3 |
H10 |
110.393 |
|
H5 |
N1 |
H6 |
107.245 |
| H7 |
C2 |
H8 |
107.771 |
|
H9 |
C3 |
H10 |
107.568 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.477 |
|
|
|
| 2 |
C |
0.613 |
|
|
|
| 3 |
C |
0.961 |
|
|
|
| 4 |
O |
-0.677 |
|
|
|
| 5 |
H |
0.021 |
|
|
|
| 6 |
H |
-0.026 |
|
|
|
| 7 |
H |
-0.078 |
|
|
|
| 8 |
H |
-0.178 |
|
|
|
| 9 |
H |
-0.159 |
|
|
|
| 10 |
H |
-0.163 |
|
|
|
| 11 |
H |
0.163 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.723 |
1.093 |
0.661 |
3.008 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-27.160 |
1.519 |
0.565 |
| y |
1.519 |
-25.507 |
-0.292 |
| z |
0.565 |
-0.292 |
-24.677 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.068 |
1.519 |
0.565 |
| y |
1.519 |
0.412 |
-0.292 |
| z |
0.565 |
-0.292 |
1.656 |
|
| Polar |
|---|
| 3z2-r2 | 3.311 |
|---|
| x2-y2 | -1.653 |
|---|
| xy | 1.519 |
|---|
| xz | 0.565 |
|---|
| yz | -0.292 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.970 |
0.142 |
-0.002 |
| y |
0.142 |
5.807 |
0.031 |
| z |
-0.002 |
0.031 |
5.284 |
<r2> (average value of r
2) Å
2
| <r2> |
88.217 |
| (<r2>)1/2 |
9.392 |
