Jump to
S1C2
S1C3
Vibrational Frequencies calculated at QCISD(T)/aug-cc-pVDZ
Geometric Data calculated at QCISD(T)/aug-cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Vibrational Frequencies calculated at QCISD(T)/aug-cc-pVDZ
Geometric Data calculated at QCISD(T)/aug-cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at QCISD(T)/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -209.895042 |
| Energy at 298.15K | -209.904106 |
| HF Energy | -209.141113 |
| Nuclear repulsion energy | 133.276227 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3724 |
3642 |
|
|
|
|
| 2 |
A |
3559 |
3480 |
|
|
|
|
| 3 |
A |
3466 |
3389 |
|
|
|
|
| 4 |
A |
3087 |
3018 |
|
|
|
|
| 5 |
A |
3076 |
3007 |
|
|
|
|
| 6 |
A |
2990 |
2924 |
|
|
|
|
| 7 |
A |
2969 |
2903 |
|
|
|
|
| 8 |
A |
1639 |
1603 |
|
|
|
|
| 9 |
A |
1491 |
1458 |
|
|
|
|
| 10 |
A |
1478 |
1446 |
|
|
|
|
| 11 |
A |
1433 |
1401 |
|
|
|
|
| 12 |
A |
1391 |
1360 |
|
|
|
|
| 13 |
A |
1354 |
1324 |
|
|
|
|
| 14 |
A |
1310 |
1281 |
|
|
|
|
| 15 |
A |
1242 |
1215 |
|
|
|
|
| 16 |
A |
1170 |
1144 |
|
|
|
|
| 17 |
A |
1105 |
1080 |
|
|
|
|
| 18 |
A |
1051 |
1027 |
|
|
|
|
| 19 |
A |
1001 |
979 |
|
|
|
|
| 20 |
A |
929 |
908 |
|
|
|
|
| 21 |
A |
878 |
859 |
|
|
|
|
| 22 |
A |
823 |
805 |
|
|
|
|
| 23 |
A |
536 |
524 |
|
|
|
|
| 24 |
A |
510 |
498 |
|
|
|
|
| 25 |
A |
320 |
313 |
|
|
|
|
| 26 |
A |
255 |
249 |
|
|
|
|
| 27 |
A |
177 |
173 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21481.5 cm
-1
Scaled (by 0.9778) Zero Point Vibrational Energy (zpe) 21004.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD(T)/aug-cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
-1.374 |
-0.573 |
0.123 |
| C2 |
-0.640 |
0.650 |
-0.286 |
| C3 |
0.781 |
0.561 |
0.282 |
| O4 |
1.436 |
-0.628 |
-0.173 |
| H5 |
-1.670 |
-0.491 |
1.099 |
| H6 |
-2.229 |
-0.681 |
-0.426 |
| H7 |
-1.120 |
1.596 |
0.044 |
| H8 |
-0.584 |
0.657 |
-1.390 |
| H9 |
1.389 |
1.419 |
-0.055 |
| H10 |
0.739 |
0.584 |
1.393 |
| H11 |
0.758 |
-1.322 |
-0.112 |
Atom - Atom Distances (Å)
| |
N1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| N1 | | 1.4837 | 2.4403 | 2.8265 | 1.0239 | 1.0217 | 2.1849 | 2.1035 | 3.4111 | 2.7237 | 2.2718 |
C2 | 1.4837 | | 1.5331 | 2.4411 | 2.0695 | 2.0773 | 1.1106 | 1.1051 | 2.1828 | 2.1747 | 2.4236 | C3 | 2.4403 | 1.5331 | | 1.4319 | 2.7894 | 3.3321 | 2.1768 | 2.1600 | 1.1044 | 1.1127 | 1.9240 | O4 | 2.8265 | 2.4411 | 1.4319 | | 3.3593 | 3.6745 | 3.3948 | 2.6854 | 2.0506 | 2.0995 | 0.9726 | H5 | 1.0239 | 2.0695 | 2.7894 | 3.3593 | | 1.6362 | 2.4024 | 2.9487 | 3.7867 | 2.6540 | 2.8374 | H6 | 1.0217 | 2.0773 | 3.3321 | 3.6745 | 1.6362 | | 2.5761 | 2.3293 | 4.2000 | 3.7041 | 3.0708 | H7 | 2.1849 | 1.1106 | 2.1768 | 3.3948 | 2.4024 | 2.5761 | | 1.7964 | 2.5172 | 2.5098 | 3.4730 | H8 | 2.1035 | 1.1051 | 2.1600 | 2.6854 | 2.9487 | 2.3293 | 1.7964 | | 2.5011 | 3.0827 | 2.7104 | H9 | 3.4111 | 2.1828 | 1.1044 | 2.0506 | 3.7867 | 4.2000 | 2.5172 | 2.5011 | | 1.7942 | 2.8132 | H10 | 2.7237 | 2.1747 | 1.1127 | 2.0995 | 2.6540 | 3.7041 | 2.5098 | 3.0827 | 1.7942 | | 2.4288 | H11 | 2.2718 | 2.4236 | 1.9240 | 0.9726 | 2.8374 | 3.0708 | 3.4730 | 2.7104 | 2.8132 | 2.4288 | |
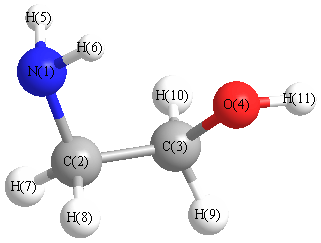 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
C2 |
C3 |
107.964 |
|
N1 |
C2 |
H7 |
113.976 |
| N1 |
C2 |
H8 |
107.795 |
|
C2 |
N1 |
H5 |
109.880 |
| C2 |
N1 |
H6 |
110.671 |
|
C2 |
C3 |
O4 |
110.789 |
| C2 |
C3 |
H9 |
110.652 |
|
C2 |
C3 |
H10 |
109.532 |
| C3 |
C2 |
H7 |
109.817 |
|
C3 |
C2 |
H8 |
108.832 |
| C3 |
O4 |
H11 |
104.675 |
|
O4 |
C3 |
H9 |
107.200 |
| O4 |
C3 |
H10 |
110.565 |
|
H5 |
N1 |
H6 |
106.238 |
| H7 |
C2 |
H8 |
108.338 |
|
H9 |
C3 |
H10 |
108.045 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
