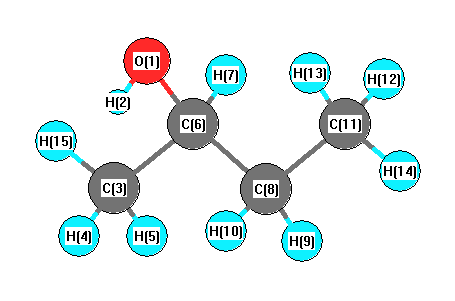
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3850 |
3690 |
17.90 |
|
|
|
| 2 |
A |
3153 |
3022 |
35.17 |
|
|
|
| 3 |
A |
3151 |
3020 |
12.00 |
|
|
|
| 4 |
A |
3134 |
3004 |
49.10 |
|
|
|
| 5 |
A |
3127 |
2997 |
32.03 |
|
|
|
| 6 |
A |
3087 |
2959 |
24.62 |
|
|
|
| 7 |
A |
3069 |
2941 |
24.07 |
|
|
|
| 8 |
A |
3057 |
2930 |
26.77 |
|
|
|
| 9 |
A |
3046 |
2919 |
23.14 |
|
|
|
| 10 |
A |
3031 |
2905 |
19.11 |
|
|
|
| 11 |
A |
1485 |
1423 |
7.35 |
|
|
|
| 12 |
A |
1476 |
1415 |
16.90 |
|
|
|
| 13 |
A |
1474 |
1412 |
1.53 |
|
|
|
| 14 |
A |
1473 |
1412 |
0.87 |
|
|
|
| 15 |
A |
1459 |
1398 |
0.29 |
|
|
|
| 16 |
A |
1413 |
1354 |
37.34 |
|
|
|
| 17 |
A |
1396 |
1338 |
8.36 |
|
|
|
| 18 |
A |
1391 |
1333 |
14.50 |
|
|
|
| 19 |
A |
1381 |
1324 |
0.89 |
|
|
|
| 20 |
A |
1325 |
1270 |
1.30 |
|
|
|
| 21 |
A |
1311 |
1257 |
2.68 |
|
|
|
| 22 |
A |
1249 |
1197 |
6.65 |
|
|
|
| 23 |
A |
1188 |
1138 |
23.64 |
|
|
|
| 24 |
A |
1145 |
1097 |
17.19 |
|
|
|
| 25 |
A |
1094 |
1049 |
65.30 |
|
|
|
| 26 |
A |
1061 |
1017 |
2.21 |
|
|
|
| 27 |
A |
1005 |
963 |
20.23 |
|
|
|
| 28 |
A |
972 |
931 |
7.69 |
|
|
|
| 29 |
A |
931 |
892 |
8.70 |
|
|
|
| 30 |
A |
823 |
788 |
5.40 |
|
|
|
| 31 |
A |
770 |
738 |
0.55 |
|
|
|
| 32 |
A |
498 |
477 |
9.18 |
|
|
|
| 33 |
A |
461 |
442 |
5.72 |
|
|
|
| 34 |
A |
376 |
361 |
5.86 |
|
|
|
| 35 |
A |
270 |
259 |
103.34 |
|
|
|
| 36 |
A |
252 |
242 |
1.51 |
|
|
|
| 37 |
A |
228 |
218 |
5.48 |
|
|
|
| 38 |
A |
212 |
204 |
0.19 |
|
|
|
| 39 |
A |
114 |
109 |
0.16 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29969.8 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 28720.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.588 |
|
|
|
| 2 |
H |
0.124 |
|
|
|
| 3 |
C |
0.598 |
|
|
|
| 4 |
H |
-0.129 |
|
|
|
| 5 |
H |
-0.105 |
|
|
|
| 6 |
C |
0.508 |
|
|
|
| 7 |
H |
-0.484 |
|
|
|
| 8 |
C |
0.656 |
|
|
|
| 9 |
H |
-0.269 |
|
|
|
| 10 |
H |
-0.311 |
|
|
|
| 11 |
C |
0.422 |
|
|
|
| 12 |
H |
-0.115 |
|
|
|
| 13 |
H |
-0.090 |
|
|
|
| 14 |
H |
-0.122 |
|
|
|
| 15 |
H |
-0.096 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.164 |
-1.362 |
0.871 |
1.625 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.800 |
0.259 |
-0.701 |
| y |
0.259 |
-35.992 |
2.313 |
| z |
-0.701 |
2.313 |
-31.090 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.259 |
0.259 |
-0.701 |
| y |
0.259 |
-3.547 |
2.313 |
| z |
-0.701 |
2.313 |
3.806 |
|
| Polar |
|---|
| 3z2-r2 | 7.612 |
|---|
| x2-y2 | 2.192 |
|---|
| xy | 0.259 |
|---|
| xz | -0.701 |
|---|
| yz | 2.313 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.517 |
0.157 |
-0.081 |
| y |
0.157 |
8.250 |
0.040 |
| z |
-0.081 |
0.040 |
7.835 |
<r2> (average value of r
2) Å
2
| <r2> |
146.499 |
| (<r2>)1/2 |
12.104 |