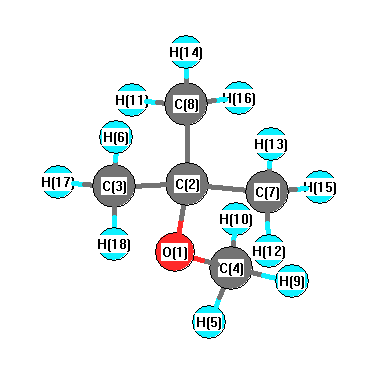
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3155 |
3024 |
53.49 |
|
|
|
| 2 |
A |
3149 |
3018 |
22.73 |
|
|
|
| 3 |
A |
3147 |
3016 |
7.29 |
|
|
|
| 4 |
A |
3144 |
3013 |
51.71 |
|
|
|
| 5 |
A |
3067 |
2939 |
5.03 |
|
|
|
| 6 |
A |
3060 |
2933 |
27.02 |
|
|
|
| 7 |
A |
3017 |
2891 |
56.97 |
|
|
|
| 8 |
A |
1506 |
1443 |
8.49 |
|
|
|
| 9 |
A |
1492 |
1430 |
4.84 |
|
|
|
| 10 |
A |
1479 |
1417 |
6.14 |
|
|
|
| 11 |
A |
1470 |
1408 |
0.80 |
|
|
|
| 12 |
A |
1447 |
1386 |
0.33 |
|
|
|
| 13 |
A |
1407 |
1348 |
15.31 |
|
|
|
| 14 |
A |
1387 |
1329 |
23.57 |
|
|
|
| 15 |
A |
1291 |
1237 |
11.65 |
|
|
|
| 16 |
A |
1230 |
1178 |
105.31 |
|
|
|
| 17 |
A |
1195 |
1145 |
2.25 |
|
|
|
| 18 |
A |
1145 |
1097 |
87.11 |
|
|
|
| 19 |
A |
1037 |
994 |
1.55 |
|
|
|
| 20 |
A |
928 |
890 |
0.21 |
|
|
|
| 21 |
A |
877 |
841 |
16.88 |
|
|
|
| 22 |
A |
744 |
713 |
5.29 |
|
|
|
| 23 |
A |
511 |
490 |
1.48 |
|
|
|
| 24 |
A |
415 |
398 |
0.86 |
|
|
|
| 25 |
A |
367 |
352 |
0.71 |
|
|
|
| 26 |
A |
288 |
276 |
0.65 |
|
|
|
| 27 |
A |
256 |
245 |
0.21 |
|
|
|
| 28 |
A |
3155 |
3023 |
33.70 |
|
|
|
| 29 |
A |
3148 |
3017 |
4.30 |
|
|
|
| 30 |
A |
3140 |
3009 |
0.00 |
|
|
|
| 31 |
A |
3080 |
2952 |
59.93 |
|
|
|
| 32 |
A |
3057 |
2930 |
12.17 |
|
|
|
| 33 |
A |
1488 |
1426 |
0.20 |
|
|
|
| 34 |
A |
1472 |
1411 |
6.39 |
|
|
|
| 35 |
A |
1465 |
1404 |
0.27 |
|
|
|
| 36 |
A |
1450 |
1390 |
0.03 |
|
|
|
| 37 |
A |
1380 |
1323 |
24.12 |
|
|
|
| 38 |
A |
1261 |
1208 |
15.75 |
|
|
|
| 39 |
A |
1161 |
1113 |
1.50 |
|
|
|
| 40 |
A |
1030 |
987 |
1.56 |
|
|
|
| 41 |
A |
945 |
906 |
0.05 |
|
|
|
| 42 |
A |
917 |
879 |
0.03 |
|
|
|
| 43 |
A |
462 |
443 |
5.23 |
|
|
|
| 44 |
A |
344 |
330 |
1.36 |
|
|
|
| 45 |
A |
277 |
266 |
1.16 |
|
|
|
| 46 |
A |
224 |
214 |
1.69 |
|
|
|
| 47 |
A |
165 |
158 |
1.24 |
|
|
|
| 48 |
A |
12 |
12 |
2.53 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35920.1 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 34422.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.611 |
|
|
|
| 2 |
C |
-1.994 |
|
|
|
| 3 |
C |
0.782 |
|
|
|
| 4 |
C |
0.877 |
|
|
|
| 5 |
H |
-0.293 |
|
|
|
| 6 |
H |
-0.039 |
|
|
|
| 7 |
C |
1.235 |
|
|
|
| 8 |
C |
1.235 |
|
|
|
| 9 |
H |
-0.148 |
|
|
|
| 10 |
H |
-0.148 |
|
|
|
| 11 |
H |
-0.076 |
|
|
|
| 12 |
H |
-0.076 |
|
|
|
| 13 |
H |
-0.109 |
|
|
|
| 14 |
H |
-0.109 |
|
|
|
| 15 |
H |
-0.165 |
|
|
|
| 16 |
H |
-0.165 |
|
|
|
| 17 |
H |
-0.098 |
|
|
|
| 18 |
H |
-0.098 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.102 |
0.572 |
0.000 |
1.242 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.598 |
-1.819 |
0.000 |
| y |
-1.819 |
-38.579 |
0.000 |
| z |
0.000 |
0.000 |
-40.040 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.288 |
-1.819 |
0.000 |
| y |
-1.819 |
1.740 |
0.000 |
| z |
0.000 |
0.000 |
-0.452 |
|
| Polar |
|---|
| 3z2-r2 | -0.904 |
|---|
| x2-y2 | -2.019 |
|---|
| xy | -1.819 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.059 |
-0.529 |
0.000 |
| y |
-0.529 |
10.794 |
0.000 |
| z |
0.000 |
0.000 |
9.970 |
<r2> (average value of r
2) Å
2
| <r2> |
179.486 |
| (<r2>)1/2 |
13.397 |