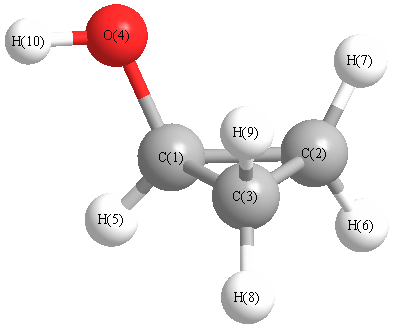
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3863 |
3702 |
31.17 |
|
|
|
| 2 |
A |
3269 |
3133 |
11.85 |
|
|
|
| 3 |
A |
3254 |
3118 |
2.25 |
|
|
|
| 4 |
A |
3170 |
3037 |
6.07 |
|
|
|
| 5 |
A |
3160 |
3028 |
15.05 |
|
|
|
| 6 |
A |
3143 |
3012 |
30.09 |
|
|
|
| 7 |
A |
1507 |
1444 |
20.88 |
|
|
|
| 8 |
A |
1441 |
1381 |
3.10 |
|
|
|
| 9 |
A |
1406 |
1347 |
7.62 |
|
|
|
| 10 |
A |
1299 |
1245 |
76.30 |
|
|
|
| 11 |
A |
1248 |
1196 |
50.98 |
|
|
|
| 12 |
A |
1191 |
1142 |
8.84 |
|
|
|
| 13 |
A |
1183 |
1134 |
0.25 |
|
|
|
| 14 |
A |
1114 |
1068 |
1.58 |
|
|
|
| 15 |
A |
1052 |
1008 |
4.83 |
|
|
|
| 16 |
A |
1041 |
998 |
19.93 |
|
|
|
| 17 |
A |
993 |
952 |
9.17 |
|
|
|
| 18 |
A |
942 |
903 |
21.96 |
|
|
|
| 19 |
A |
839 |
804 |
11.67 |
|
|
|
| 20 |
A |
818 |
784 |
6.79 |
|
|
|
| 21 |
A |
756 |
724 |
3.23 |
|
|
|
| 22 |
A |
409 |
392 |
11.15 |
|
|
|
| 23 |
A |
402 |
385 |
9.58 |
|
|
|
| 24 |
A |
301 |
288 |
106.22 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 18898.8 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 18110.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
1.113 |
|
|
|
| 2 |
C |
0.741 |
|
|
|
| 3 |
C |
0.718 |
|
|
|
| 4 |
O |
-0.570 |
|
|
|
| 5 |
H |
-0.412 |
|
|
|
| 6 |
H |
-0.440 |
|
|
|
| 7 |
H |
-0.404 |
|
|
|
| 8 |
H |
-0.438 |
|
|
|
| 9 |
H |
-0.429 |
|
|
|
| 10 |
H |
0.121 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.474 |
1.281 |
0.681 |
1.526 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.239 |
-3.189 |
0.012 |
| y |
-3.189 |
-24.491 |
-0.173 |
| z |
0.012 |
-0.173 |
-24.333 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.826 |
-3.189 |
0.012 |
| y |
-3.189 |
0.295 |
-0.173 |
| z |
0.012 |
-0.173 |
0.531 |
|
| Polar |
|---|
| 3z2-r2 | 1.063 |
|---|
| x2-y2 | -0.748 |
|---|
| xy | -3.189 |
|---|
| xz | 0.012 |
|---|
| yz | -0.173 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.529 |
-0.115 |
-0.055 |
| y |
-0.115 |
6.171 |
-0.008 |
| z |
-0.055 |
-0.008 |
5.591 |
<r2> (average value of r
2) Å
2
| <r2> |
73.424 |
| (<r2>)1/2 |
8.569 |