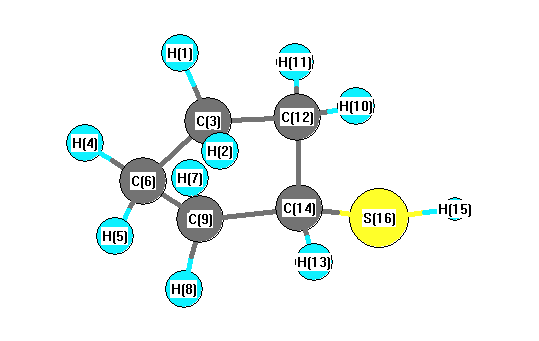
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3147 |
3016 |
64.73 |
|
|
|
| 2 |
A |
3135 |
3005 |
19.38 |
|
|
|
| 3 |
A |
3131 |
3000 |
4.95 |
|
|
|
| 4 |
A |
3117 |
2987 |
18.28 |
|
|
|
| 5 |
A |
3090 |
2961 |
40.21 |
|
|
|
| 6 |
A |
3083 |
2954 |
30.35 |
|
|
|
| 7 |
A |
3076 |
2947 |
27.33 |
|
|
|
| 8 |
A |
3071 |
2943 |
10.24 |
|
|
|
| 9 |
A |
3065 |
2937 |
11.58 |
|
|
|
| 10 |
A |
2705 |
2592 |
4.13 |
|
|
|
| 11 |
A |
1511 |
1448 |
1.41 |
|
|
|
| 12 |
A |
1487 |
1425 |
5.58 |
|
|
|
| 13 |
A |
1483 |
1421 |
4.50 |
|
|
|
| 14 |
A |
1473 |
1411 |
1.03 |
|
|
|
| 15 |
A |
1350 |
1294 |
5.54 |
|
|
|
| 16 |
A |
1338 |
1282 |
1.01 |
|
|
|
| 17 |
A |
1328 |
1273 |
1.12 |
|
|
|
| 18 |
A |
1322 |
1267 |
2.37 |
|
|
|
| 19 |
A |
1285 |
1232 |
0.26 |
|
|
|
| 20 |
A |
1269 |
1216 |
12.40 |
|
|
|
| 21 |
A |
1255 |
1202 |
1.98 |
|
|
|
| 22 |
A |
1231 |
1179 |
5.41 |
|
|
|
| 23 |
A |
1199 |
1149 |
0.39 |
|
|
|
| 24 |
A |
1166 |
1117 |
3.91 |
|
|
|
| 25 |
A |
1130 |
1083 |
0.46 |
|
|
|
| 26 |
A |
1074 |
1029 |
0.69 |
|
|
|
| 27 |
A |
993 |
952 |
0.71 |
|
|
|
| 28 |
A |
982 |
941 |
0.19 |
|
|
|
| 29 |
A |
968 |
927 |
3.02 |
|
|
|
| 30 |
A |
947 |
908 |
0.35 |
|
|
|
| 31 |
A |
911 |
873 |
1.39 |
|
|
|
| 32 |
A |
870 |
834 |
4.78 |
|
|
|
| 33 |
A |
827 |
792 |
4.56 |
|
|
|
| 34 |
A |
814 |
780 |
1.34 |
|
|
|
| 35 |
A |
750 |
718 |
1.60 |
|
|
|
| 36 |
A |
629 |
603 |
0.41 |
|
|
|
| 37 |
A |
462 |
443 |
0.24 |
|
|
|
| 38 |
A |
368 |
353 |
1.11 |
|
|
|
| 39 |
A |
249 |
239 |
2.58 |
|
|
|
| 40 |
A |
199 |
191 |
14.26 |
|
|
|
| 41 |
A |
162 |
155 |
0.69 |
|
|
|
| 42 |
A |
46 |
44 |
0.08 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30846.7 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 29560.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
-0.253 |
|
|
|
| 2 |
H |
-0.293 |
|
|
|
| 3 |
C |
0.534 |
|
|
|
| 4 |
H |
-0.226 |
|
|
|
| 5 |
H |
-0.241 |
|
|
|
| 6 |
C |
0.543 |
|
|
|
| 7 |
H |
-0.301 |
|
|
|
| 8 |
H |
-0.287 |
|
|
|
| 9 |
C |
0.625 |
|
|
|
| 10 |
H |
-0.303 |
|
|
|
| 11 |
H |
-0.277 |
|
|
|
| 12 |
C |
0.676 |
|
|
|
| 13 |
H |
-0.417 |
|
|
|
| 14 |
C |
-0.017 |
|
|
|
| 15 |
H |
0.077 |
|
|
|
| 16 |
S |
0.160 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.617 |
0.685 |
0.656 |
1.875 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-48.937 |
2.513 |
1.227 |
| y |
2.513 |
-45.159 |
1.130 |
| z |
1.227 |
1.130 |
-45.847 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.434 |
2.513 |
1.227 |
| y |
2.513 |
2.233 |
1.130 |
| z |
1.227 |
1.130 |
1.201 |
|
| Polar |
|---|
| 3z2-r2 | 2.402 |
|---|
| x2-y2 | -3.778 |
|---|
| xy | 2.513 |
|---|
| xz | 1.227 |
|---|
| yz | 1.130 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
14.314 |
0.108 |
-0.019 |
| y |
0.108 |
11.492 |
-0.022 |
| z |
-0.019 |
-0.022 |
10.123 |
<r2> (average value of r
2) Å
2
| <r2> |
230.414 |
| (<r2>)1/2 |
15.179 |