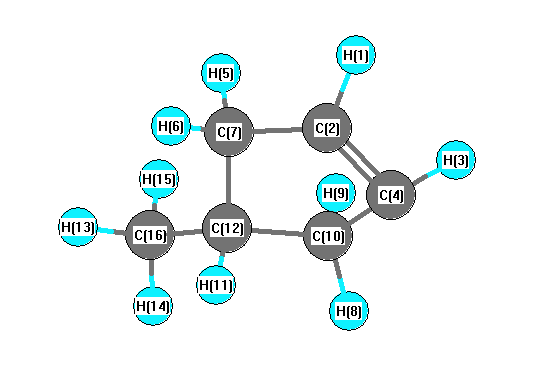
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3231 |
3096 |
25.53 |
|
|
|
| 2 |
A |
3205 |
3071 |
7.73 |
|
|
|
| 3 |
A |
3129 |
2999 |
26.80 |
|
|
|
| 4 |
A |
3125 |
2994 |
49.25 |
|
|
|
| 5 |
A |
3092 |
2963 |
30.52 |
|
|
|
| 6 |
A |
3087 |
2958 |
33.33 |
|
|
|
| 7 |
A |
3053 |
2925 |
17.69 |
|
|
|
| 8 |
A |
3045 |
2918 |
31.55 |
|
|
|
| 9 |
A |
3022 |
2896 |
38.93 |
|
|
|
| 10 |
A |
3021 |
2895 |
31.63 |
|
|
|
| 11 |
A |
1700 |
1629 |
5.79 |
|
|
|
| 12 |
A |
1483 |
1421 |
6.93 |
|
|
|
| 13 |
A |
1478 |
1417 |
1.45 |
|
|
|
| 14 |
A |
1476 |
1415 |
2.21 |
|
|
|
| 15 |
A |
1471 |
1409 |
4.30 |
|
|
|
| 16 |
A |
1397 |
1339 |
6.02 |
|
|
|
| 17 |
A |
1374 |
1317 |
0.84 |
|
|
|
| 18 |
A |
1371 |
1314 |
2.96 |
|
|
|
| 19 |
A |
1330 |
1275 |
1.98 |
|
|
|
| 20 |
A |
1299 |
1245 |
0.32 |
|
|
|
| 21 |
A |
1283 |
1229 |
3.02 |
|
|
|
| 22 |
A |
1230 |
1179 |
0.30 |
|
|
|
| 23 |
A |
1174 |
1125 |
0.00 |
|
|
|
| 24 |
A |
1147 |
1099 |
2.23 |
|
|
|
| 25 |
A |
1129 |
1082 |
0.76 |
|
|
|
| 26 |
A |
1113 |
1067 |
0.30 |
|
|
|
| 27 |
A |
1069 |
1025 |
3.95 |
|
|
|
| 28 |
A |
995 |
954 |
1.12 |
|
|
|
| 29 |
A |
995 |
953 |
11.56 |
|
|
|
| 30 |
A |
962 |
922 |
0.02 |
|
|
|
| 31 |
A |
930 |
891 |
5.11 |
|
|
|
| 32 |
A |
926 |
887 |
6.43 |
|
|
|
| 33 |
A |
905 |
867 |
0.00 |
|
|
|
| 34 |
A |
836 |
801 |
1.16 |
|
|
|
| 35 |
A |
781 |
749 |
0.70 |
|
|
|
| 36 |
A |
697 |
668 |
40.71 |
|
|
|
| 37 |
A |
577 |
553 |
4.63 |
|
|
|
| 38 |
A |
421 |
403 |
1.09 |
|
|
|
| 39 |
A |
397 |
380 |
0.07 |
|
|
|
| 40 |
A |
314 |
301 |
0.04 |
|
|
|
| 41 |
A |
250 |
240 |
0.00 |
|
|
|
| 42 |
A |
100 |
96 |
0.12 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31808.3 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 30481.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
-0.392 |
|
|
|
| 2 |
C |
0.331 |
|
|
|
| 3 |
H |
-0.392 |
|
|
|
| 4 |
C |
0.331 |
|
|
|
| 5 |
H |
-0.189 |
|
|
|
| 6 |
H |
-0.220 |
|
|
|
| 7 |
C |
0.558 |
|
|
|
| 8 |
H |
-0.220 |
|
|
|
| 9 |
H |
-0.188 |
|
|
|
| 10 |
C |
0.558 |
|
|
|
| 11 |
H |
-0.335 |
|
|
|
| 12 |
C |
-0.128 |
|
|
|
| 13 |
H |
-0.119 |
|
|
|
| 14 |
H |
-0.119 |
|
|
|
| 15 |
H |
-0.077 |
|
|
|
| 16 |
C |
0.600 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.266 |
-0.000 |
0.032 |
0.268 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.673 |
-0.000 |
0.495 |
| y |
-0.000 |
-37.297 |
0.001 |
| z |
0.495 |
0.001 |
-39.405 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.322 |
-0.000 |
0.495 |
| y |
-0.000 |
1.743 |
0.001 |
| z |
0.495 |
0.001 |
-1.420 |
|
| Polar |
|---|
| 3z2-r2 | -2.840 |
|---|
| x2-y2 | -1.377 |
|---|
| xy | -0.000 |
|---|
| xz | 0.495 |
|---|
| yz | 0.001 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.441 |
0.000 |
-0.019 |
| y |
0.000 |
11.490 |
0.000 |
| z |
-0.019 |
0.000 |
8.605 |
<r2> (average value of r
2) Å
2
| <r2> |
164.034 |
| (<r2>)1/2 |
12.808 |