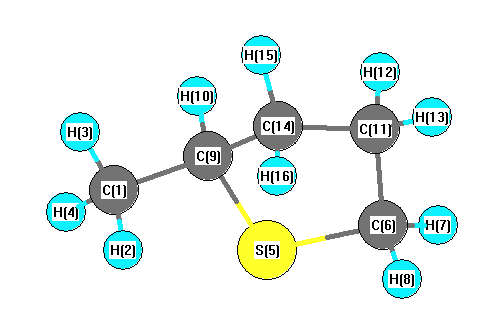
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3150 |
3018 |
19.92 |
|
|
|
| 2 |
A |
3148 |
3017 |
19.78 |
|
|
|
| 3 |
A |
3131 |
3000 |
27.39 |
|
|
|
| 4 |
A |
3122 |
2992 |
31.86 |
|
|
|
| 5 |
A |
3111 |
2981 |
26.52 |
|
|
|
| 6 |
A |
3084 |
2955 |
30.12 |
|
|
|
| 7 |
A |
3071 |
2943 |
10.83 |
|
|
|
| 8 |
A |
3066 |
2939 |
26.46 |
|
|
|
| 9 |
A |
3055 |
2927 |
40.10 |
|
|
|
| 10 |
A |
3049 |
2922 |
17.27 |
|
|
|
| 11 |
A |
1490 |
1427 |
0.69 |
|
|
|
| 12 |
A |
1477 |
1415 |
6.62 |
|
|
|
| 13 |
A |
1469 |
1408 |
5.03 |
|
|
|
| 14 |
A |
1468 |
1407 |
9.13 |
|
|
|
| 15 |
A |
1464 |
1403 |
2.22 |
|
|
|
| 16 |
A |
1395 |
1337 |
5.71 |
|
|
|
| 17 |
A |
1374 |
1317 |
1.34 |
|
|
|
| 18 |
A |
1349 |
1293 |
0.31 |
|
|
|
| 19 |
A |
1309 |
1254 |
0.19 |
|
|
|
| 20 |
A |
1288 |
1235 |
3.20 |
|
|
|
| 21 |
A |
1275 |
1222 |
19.20 |
|
|
|
| 22 |
A |
1232 |
1181 |
4.06 |
|
|
|
| 23 |
A |
1195 |
1145 |
7.49 |
|
|
|
| 24 |
A |
1163 |
1115 |
0.57 |
|
|
|
| 25 |
A |
1114 |
1068 |
0.47 |
|
|
|
| 26 |
A |
1072 |
1027 |
1.85 |
|
|
|
| 27 |
A |
1047 |
1003 |
0.73 |
|
|
|
| 28 |
A |
1034 |
991 |
4.00 |
|
|
|
| 29 |
A |
967 |
926 |
2.45 |
|
|
|
| 30 |
A |
958 |
919 |
1.49 |
|
|
|
| 31 |
A |
912 |
874 |
0.99 |
|
|
|
| 32 |
A |
862 |
826 |
0.92 |
|
|
|
| 33 |
A |
853 |
818 |
2.54 |
|
|
|
| 34 |
A |
701 |
672 |
1.93 |
|
|
|
| 35 |
A |
643 |
616 |
3.21 |
|
|
|
| 36 |
A |
529 |
507 |
0.58 |
|
|
|
| 37 |
A |
461 |
442 |
0.13 |
|
|
|
| 38 |
A |
411 |
394 |
0.57 |
|
|
|
| 39 |
A |
286 |
274 |
0.99 |
|
|
|
| 40 |
A |
253 |
242 |
0.14 |
|
|
|
| 41 |
A |
236 |
226 |
0.10 |
|
|
|
| 42 |
A |
94 |
90 |
1.38 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31183.1 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 29882.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.597 |
|
|
|
| 2 |
H |
-0.076 |
|
|
|
| 3 |
H |
-0.091 |
|
|
|
| 4 |
H |
-0.126 |
|
|
|
| 5 |
S |
0.251 |
|
|
|
| 6 |
C |
0.154 |
|
|
|
| 7 |
H |
-0.156 |
|
|
|
| 8 |
H |
-0.181 |
|
|
|
| 9 |
C |
-0.099 |
|
|
|
| 10 |
H |
-0.352 |
|
|
|
| 11 |
C |
0.600 |
|
|
|
| 12 |
H |
-0.292 |
|
|
|
| 13 |
H |
-0.176 |
|
|
|
| 14 |
C |
0.516 |
|
|
|
| 15 |
H |
-0.258 |
|
|
|
| 16 |
H |
-0.311 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.363 |
2.102 |
0.000 |
2.133 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-42.714 |
-0.293 |
1.026 |
| y |
-0.293 |
-48.543 |
-0.359 |
| z |
1.026 |
-0.359 |
-46.186 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.650 |
-0.293 |
1.026 |
| y |
-0.293 |
-4.093 |
-0.359 |
| z |
1.026 |
-0.359 |
-0.558 |
|
| Polar |
|---|
| 3z2-r2 | -1.115 |
|---|
| x2-y2 | 5.828 |
|---|
| xy | -0.293 |
|---|
| xz | 1.026 |
|---|
| yz | -0.359 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
13.469 |
-0.087 |
0.132 |
| y |
-0.087 |
12.294 |
-0.024 |
| z |
0.132 |
-0.024 |
10.066 |
<r2> (average value of r
2) Å
2
| <r2> |
203.836 |
| (<r2>)1/2 |
14.277 |