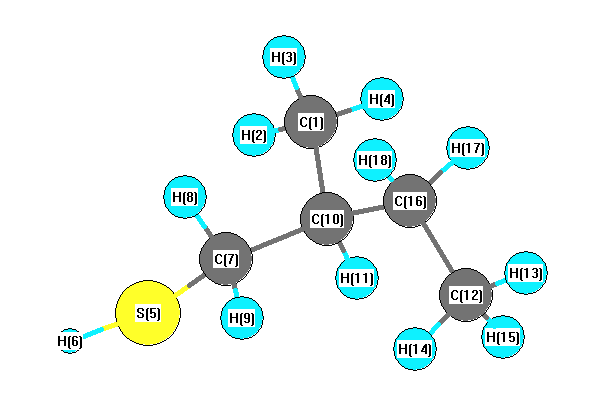
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3153 |
3022 |
11.45 |
|
|
|
| 2 |
A |
3143 |
3012 |
32.31 |
|
|
|
| 3 |
A |
3138 |
3007 |
34.53 |
|
|
|
| 4 |
A |
3133 |
3003 |
49.41 |
|
|
|
| 5 |
A |
3128 |
2998 |
17.72 |
|
|
|
| 6 |
A |
3087 |
2958 |
17.57 |
|
|
|
| 7 |
A |
3074 |
2946 |
22.84 |
|
|
|
| 8 |
A |
3055 |
2927 |
30.26 |
|
|
|
| 9 |
A |
3051 |
2924 |
39.16 |
|
|
|
| 10 |
A |
3044 |
2917 |
8.40 |
|
|
|
| 11 |
A |
3037 |
2911 |
5.54 |
|
|
|
| 12 |
A |
2711 |
2598 |
3.80 |
|
|
|
| 13 |
A |
1487 |
1425 |
10.48 |
|
|
|
| 14 |
A |
1481 |
1419 |
3.56 |
|
|
|
| 15 |
A |
1480 |
1418 |
11.18 |
|
|
|
| 16 |
A |
1477 |
1415 |
1.04 |
|
|
|
| 17 |
A |
1467 |
1406 |
2.01 |
|
|
|
| 18 |
A |
1457 |
1397 |
6.16 |
|
|
|
| 19 |
A |
1402 |
1343 |
4.15 |
|
|
|
| 20 |
A |
1400 |
1341 |
12.24 |
|
|
|
| 21 |
A |
1379 |
1321 |
1.53 |
|
|
|
| 22 |
A |
1367 |
1310 |
0.72 |
|
|
|
| 23 |
A |
1316 |
1261 |
0.88 |
|
|
|
| 24 |
A |
1295 |
1241 |
7.82 |
|
|
|
| 25 |
A |
1261 |
1209 |
4.24 |
|
|
|
| 26 |
A |
1221 |
1170 |
7.19 |
|
|
|
| 27 |
A |
1184 |
1135 |
0.53 |
|
|
|
| 28 |
A |
1150 |
1102 |
2.34 |
|
|
|
| 29 |
A |
1089 |
1044 |
0.77 |
|
|
|
| 30 |
A |
1057 |
1013 |
0.69 |
|
|
|
| 31 |
A |
1029 |
986 |
1.96 |
|
|
|
| 32 |
A |
972 |
932 |
5.13 |
|
|
|
| 33 |
A |
949 |
909 |
1.04 |
|
|
|
| 34 |
A |
896 |
859 |
3.08 |
|
|
|
| 35 |
A |
866 |
830 |
0.96 |
|
|
|
| 36 |
A |
798 |
765 |
1.52 |
|
|
|
| 37 |
A |
789 |
756 |
7.92 |
|
|
|
| 38 |
A |
725 |
695 |
0.85 |
|
|
|
| 39 |
A |
457 |
438 |
0.16 |
|
|
|
| 40 |
A |
402 |
385 |
0.07 |
|
|
|
| 41 |
A |
380 |
364 |
0.91 |
|
|
|
| 42 |
A |
250 |
240 |
0.11 |
|
|
|
| 43 |
A |
242 |
232 |
0.31 |
|
|
|
| 44 |
A |
204 |
196 |
0.05 |
|
|
|
| 45 |
A |
199 |
191 |
1.40 |
|
|
|
| 46 |
A |
147 |
141 |
11.53 |
|
|
|
| 47 |
A |
97 |
93 |
0.42 |
|
|
|
| 48 |
A |
60 |
58 |
6.51 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35093.7 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 33630.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.698 |
|
|
|
| 2 |
H |
-0.172 |
|
|
|
| 3 |
H |
-0.055 |
|
|
|
| 4 |
H |
-0.136 |
|
|
|
| 5 |
S |
0.154 |
|
|
|
| 6 |
H |
0.063 |
|
|
|
| 7 |
C |
0.499 |
|
|
|
| 8 |
H |
-0.216 |
|
|
|
| 9 |
H |
-0.304 |
|
|
|
| 10 |
C |
-0.368 |
|
|
|
| 11 |
H |
-0.483 |
|
|
|
| 12 |
C |
0.486 |
|
|
|
| 13 |
H |
-0.149 |
|
|
|
| 14 |
H |
-0.136 |
|
|
|
| 15 |
H |
-0.118 |
|
|
|
| 16 |
C |
0.810 |
|
|
|
| 17 |
H |
-0.310 |
|
|
|
| 18 |
H |
-0.262 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.513 |
-0.571 |
0.684 |
1.755 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-50.229 |
2.139 |
-1.335 |
| y |
2.139 |
-46.364 |
-1.663 |
| z |
-1.335 |
-1.663 |
-48.290 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.902 |
2.139 |
-1.335 |
| y |
2.139 |
2.895 |
-1.663 |
| z |
-1.335 |
-1.663 |
0.007 |
|
| Polar |
|---|
| 3z2-r2 | 0.014 |
|---|
| x2-y2 | -3.865 |
|---|
| xy | 2.139 |
|---|
| xz | -1.335 |
|---|
| yz | -1.663 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
14.854 |
0.177 |
0.120 |
| y |
0.177 |
12.421 |
0.052 |
| z |
0.120 |
0.052 |
10.890 |
<r2> (average value of r
2) Å
2
| <r2> |
286.727 |
| (<r2>)1/2 |
16.933 |