Jump to
S2C1
Energy calculated at HF/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -146.734930 |
| Energy at 298.15K | -146.734565 |
| HF Energy | -146.734930 |
| Nuclear repulsion energy | 46.533011 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Σg |
1229 |
1119 |
0.00 |
|
|
|
| 2 |
Σu |
1453 |
1323 |
414.07 |
|
|
|
| 3 |
Πu |
381 |
347 |
12.04 |
|
|
|
| 3 |
Πu |
381 |
347 |
12.04 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 1722.0 cm
-1
Scaled (by 0.9106) Zero Point Vibrational Energy (zpe) 1568.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/aug-cc-pVDZ
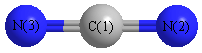
Point Group is D∞h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.000 |
| N2 |
0.000 |
0.000 |
1.234 |
| N3 |
0.000 |
0.000 |
-1.234 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
N3 |
| C1 | | 1.2339 | 1.2339 |
N2 | 1.2339 | | 2.4677 | N3 | 1.2339 | 2.4677 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N2 |
C1 |
N3 |
180.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.462 |
|
|
|
| 2 |
N |
-0.231 |
|
|
|
| 3 |
N |
-0.231 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-15.785 |
0.000 |
0.000 |
| y |
0.000 |
-15.785 |
0.000 |
| z |
0.000 |
0.000 |
-20.820 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.518 |
0.000 |
0.000 |
| y |
0.000 |
2.518 |
0.000 |
| z |
0.000 |
0.000 |
-5.036 |
|
| Polar |
|---|
| 3z2-r2 | -10.071 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.516 |
0.000 |
0.000 |
| y |
0.000 |
2.516 |
0.000 |
| z |
0.000 |
0.000 |
5.080 |
<r2> (average value of r
2) Å
2
| <r2> |
32.221 |
| (<r2>)1/2 |
5.676 |
Jump to
S1C1
Energy calculated at HF/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -146.634350 |
| Energy at 298.15K | -146.634144 |
| HF Energy | -146.634350 |
| Nuclear repulsion energy | 47.437839 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Σg |
1409 |
1283 |
0.00 |
|
|
|
| 2 |
Σu |
1522 |
1386 |
439.87 |
|
|
|
| 3 |
Πu |
631 |
575 |
18.35 |
|
|
|
| 3 |
Πu |
339 |
309 |
18.63 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 1950.4 cm
-1
Scaled (by 0.9106) Zero Point Vibrational Energy (zpe) 1776.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/aug-cc-pVDZ
Point Group is D∞h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.000 |
| N2 |
0.000 |
0.000 |
1.210 |
| N3 |
0.000 |
0.000 |
-1.210 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
N3 |
| C1 | | 1.2103 | 1.2103 |
N2 | 1.2103 | | 2.4207 | N3 | 1.2103 | 2.4207 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N2 |
C1 |
N3 |
180.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.415 |
|
|
|
| 2 |
N |
-0.208 |
|
|
|
| 3 |
N |
-0.208 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-14.700 |
0.000 |
0.000 |
| y |
0.000 |
-17.278 |
0.000 |
| z |
0.000 |
0.000 |
-20.647 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.263 |
0.000 |
0.000 |
| y |
0.000 |
0.395 |
0.000 |
| z |
0.000 |
0.000 |
-4.658 |
|
| Polar |
|---|
| 3z2-r2 | -9.315 |
|---|
| x2-y2 | 2.579 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.536 |
0.000 |
0.000 |
| y |
0.000 |
2.309 |
0.000 |
| z |
0.000 |
0.000 |
5.851 |
<r2> (average value of r
2) Å
2
| <r2> |
31.465 |
| (<r2>)1/2 |
5.609 |
