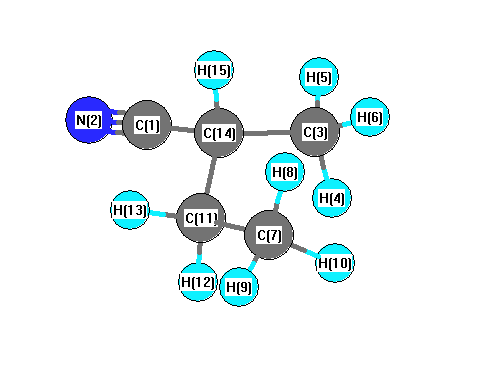
Vibrational Frequencies calculated at HF/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3271 |
2978 |
29.37 |
|
|
|
| 2 |
A |
3259 |
2967 |
35.07 |
|
|
|
| 3 |
A |
3249 |
2958 |
64.30 |
|
|
|
| 4 |
A |
3238 |
2949 |
47.33 |
|
|
|
| 5 |
A |
3219 |
2931 |
3.17 |
|
|
|
| 6 |
A |
3196 |
2911 |
7.67 |
|
|
|
| 7 |
A |
3185 |
2900 |
8.81 |
|
|
|
| 8 |
A |
3181 |
2897 |
20.38 |
|
|
|
| 9 |
A |
3168 |
2885 |
28.69 |
|
|
|
| 10 |
A |
2571 |
2341 |
21.87 |
|
|
|
| 11 |
A |
1609 |
1465 |
2.52 |
|
|
|
| 12 |
A |
1607 |
1463 |
6.97 |
|
|
|
| 13 |
A |
1596 |
1454 |
5.69 |
|
|
|
| 14 |
A |
1592 |
1450 |
7.92 |
|
|
|
| 15 |
A |
1588 |
1446 |
0.41 |
|
|
|
| 16 |
A |
1531 |
1394 |
3.20 |
|
|
|
| 17 |
A |
1526 |
1390 |
1.89 |
|
|
|
| 18 |
A |
1496 |
1362 |
1.27 |
|
|
|
| 19 |
A |
1457 |
1327 |
2.35 |
|
|
|
| 20 |
A |
1423 |
1296 |
4.67 |
|
|
|
| 21 |
A |
1382 |
1258 |
1.72 |
|
|
|
| 22 |
A |
1279 |
1165 |
2.68 |
|
|
|
| 23 |
A |
1227 |
1117 |
0.12 |
|
|
|
| 24 |
A |
1200 |
1093 |
6.69 |
|
|
|
| 25 |
A |
1114 |
1014 |
2.09 |
|
|
|
| 26 |
A |
1064 |
969 |
0.18 |
|
|
|
| 27 |
A |
1037 |
945 |
2.76 |
|
|
|
| 28 |
A |
952 |
866 |
0.54 |
|
|
|
| 29 |
A |
862 |
785 |
1.10 |
|
|
|
| 30 |
A |
814 |
741 |
2.83 |
|
|
|
| 31 |
A |
620 |
564 |
0.29 |
|
|
|
| 32 |
A |
580 |
528 |
1.75 |
|
|
|
| 33 |
A |
425 |
387 |
0.35 |
|
|
|
| 34 |
A |
351 |
319 |
1.73 |
|
|
|
| 35 |
A |
301 |
274 |
0.31 |
|
|
|
| 36 |
A |
232 |
211 |
0.07 |
|
|
|
| 37 |
A |
225 |
205 |
3.22 |
|
|
|
| 38 |
A |
181 |
165 |
7.70 |
|
|
|
| 39 |
A |
96 |
87 |
1.93 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30451.0 cm
-1
Scaled (by 0.9106) Zero Point Vibrational Energy (zpe) 27728.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.372 |
|
|
|
| 2 |
N |
-0.275 |
|
|
|
| 3 |
C |
0.712 |
|
|
|
| 4 |
H |
-0.017 |
|
|
|
| 5 |
H |
-0.081 |
|
|
|
| 6 |
H |
-0.087 |
|
|
|
| 7 |
C |
0.508 |
|
|
|
| 8 |
H |
-0.071 |
|
|
|
| 9 |
H |
-0.067 |
|
|
|
| 10 |
H |
-0.094 |
|
|
|
| 11 |
C |
0.414 |
|
|
|
| 12 |
H |
-0.154 |
|
|
|
| 13 |
H |
-0.185 |
|
|
|
| 14 |
C |
-0.118 |
|
|
|
| 15 |
H |
-0.114 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
4.221 |
1.457 |
0.560 |
4.501 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-50.330 |
-4.228 |
-0.982 |
| y |
-4.228 |
-38.096 |
-0.147 |
| z |
-0.982 |
-0.147 |
-37.299 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-12.632 |
-4.228 |
-0.982 |
| y |
-4.228 |
5.718 |
-0.147 |
| z |
-0.982 |
-0.147 |
6.914 |
|
| Polar |
|---|
| 3z2-r2 | 13.828 |
|---|
| x2-y2 | -12.234 |
|---|
| xy | -4.228 |
|---|
| xz | -0.982 |
|---|
| yz | -0.147 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
199.510 |
| (<r2>)1/2 |
14.125 |