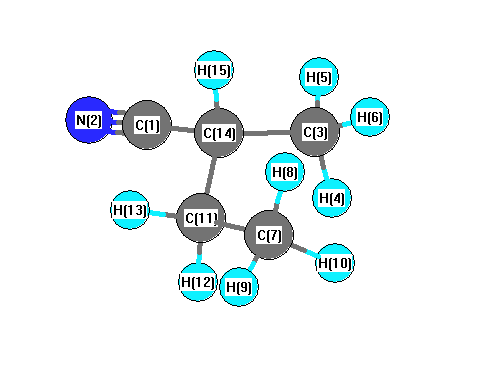
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3169 |
3037 |
14.69 |
|
|
|
| 2 |
A |
3160 |
3028 |
16.84 |
|
|
|
| 3 |
A |
3150 |
3019 |
30.73 |
|
|
|
| 4 |
A |
3141 |
3010 |
32.99 |
|
|
|
| 5 |
A |
3115 |
2986 |
3.58 |
|
|
|
| 6 |
A |
3074 |
2945 |
9.28 |
|
|
|
| 7 |
A |
3072 |
2944 |
25.10 |
|
|
|
| 8 |
A |
3059 |
2932 |
18.79 |
|
|
|
| 9 |
A |
3056 |
2929 |
1.49 |
|
|
|
| 10 |
A |
2365 |
2266 |
15.55 |
|
|
|
| 11 |
A |
1491 |
1429 |
2.60 |
|
|
|
| 12 |
A |
1487 |
1425 |
10.18 |
|
|
|
| 13 |
A |
1479 |
1418 |
7.57 |
|
|
|
| 14 |
A |
1474 |
1413 |
9.42 |
|
|
|
| 15 |
A |
1468 |
1407 |
0.35 |
|
|
|
| 16 |
A |
1401 |
1343 |
2.76 |
|
|
|
| 17 |
A |
1399 |
1341 |
6.27 |
|
|
|
| 18 |
A |
1369 |
1312 |
0.78 |
|
|
|
| 19 |
A |
1337 |
1281 |
1.47 |
|
|
|
| 20 |
A |
1308 |
1253 |
2.26 |
|
|
|
| 21 |
A |
1272 |
1219 |
1.10 |
|
|
|
| 22 |
A |
1182 |
1133 |
2.31 |
|
|
|
| 23 |
A |
1150 |
1102 |
0.22 |
|
|
|
| 24 |
A |
1112 |
1066 |
4.22 |
|
|
|
| 25 |
A |
1065 |
1020 |
2.01 |
|
|
|
| 26 |
A |
1003 |
962 |
2.02 |
|
|
|
| 27 |
A |
969 |
928 |
4.11 |
|
|
|
| 28 |
A |
910 |
872 |
0.50 |
|
|
|
| 29 |
A |
812 |
779 |
0.32 |
|
|
|
| 30 |
A |
767 |
735 |
3.46 |
|
|
|
| 31 |
A |
570 |
546 |
0.08 |
|
|
|
| 32 |
A |
540 |
517 |
1.06 |
|
|
|
| 33 |
A |
400 |
383 |
0.20 |
|
|
|
| 34 |
A |
324 |
311 |
0.76 |
|
|
|
| 35 |
A |
277 |
266 |
0.18 |
|
|
|
| 36 |
A |
220 |
211 |
0.10 |
|
|
|
| 37 |
A |
206 |
198 |
2.37 |
|
|
|
| 38 |
A |
164 |
157 |
5.93 |
|
|
|
| 39 |
A |
87 |
83 |
1.76 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 28802.1 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 27601.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.242 |
|
|
|
| 2 |
N |
-0.298 |
|
|
|
| 3 |
C |
0.676 |
|
|
|
| 4 |
H |
-0.063 |
|
|
|
| 5 |
H |
-0.127 |
|
|
|
| 6 |
H |
-0.157 |
|
|
|
| 7 |
C |
0.491 |
|
|
|
| 8 |
H |
-0.113 |
|
|
|
| 9 |
H |
-0.137 |
|
|
|
| 10 |
H |
-0.142 |
|
|
|
| 11 |
C |
0.723 |
|
|
|
| 12 |
H |
-0.262 |
|
|
|
| 13 |
H |
-0.311 |
|
|
|
| 14 |
C |
0.239 |
|
|
|
| 15 |
H |
-0.277 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
4.068 |
1.347 |
0.527 |
4.317 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-49.328 |
-3.996 |
-0.940 |
| y |
-3.996 |
-37.577 |
-0.134 |
| z |
-0.940 |
-0.134 |
-36.943 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-12.068 |
-3.996 |
-0.940 |
| y |
-3.996 |
5.558 |
-0.134 |
| z |
-0.940 |
-0.134 |
6.510 |
|
| Polar |
|---|
| 3z2-r2 | 13.019 |
|---|
| x2-y2 | -11.751 |
|---|
| xy | -3.996 |
|---|
| xz | -0.940 |
|---|
| yz | -0.134 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.866 |
0.468 |
0.226 |
| y |
0.468 |
9.349 |
0.094 |
| z |
0.226 |
0.094 |
8.168 |
<r2> (average value of r
2) Å
2
| <r2> |
198.859 |
| (<r2>)1/2 |
14.102 |