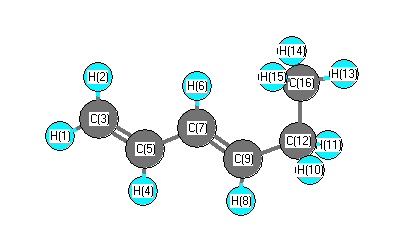
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3267 |
3131 |
12.06 |
|
|
|
| 2 |
A' |
3186 |
3053 |
17.79 |
|
|
|
| 3 |
A' |
3173 |
3041 |
2.43 |
|
|
|
| 4 |
A' |
3164 |
3032 |
10.77 |
|
|
|
| 5 |
A' |
3157 |
3026 |
17.18 |
|
|
|
| 6 |
A' |
3145 |
3014 |
31.86 |
|
|
|
| 7 |
A' |
3059 |
2931 |
21.57 |
|
|
|
| 8 |
A' |
3030 |
2903 |
32.55 |
|
|
|
| 9 |
A' |
1742 |
1669 |
29.25 |
|
|
|
| 10 |
A' |
1684 |
1613 |
8.92 |
|
|
|
| 11 |
A' |
1487 |
1425 |
5.23 |
|
|
|
| 12 |
A' |
1457 |
1396 |
7.48 |
|
|
|
| 13 |
A' |
1445 |
1385 |
4.31 |
|
|
|
| 14 |
A' |
1403 |
1345 |
7.65 |
|
|
|
| 15 |
A' |
1368 |
1311 |
10.36 |
|
|
|
| 16 |
A' |
1334 |
1279 |
4.69 |
|
|
|
| 17 |
A' |
1316 |
1261 |
0.26 |
|
|
|
| 18 |
A' |
1284 |
1230 |
0.18 |
|
|
|
| 19 |
A' |
1206 |
1156 |
4.10 |
|
|
|
| 20 |
A' |
1115 |
1068 |
5.60 |
|
|
|
| 21 |
A' |
1035 |
992 |
4.24 |
|
|
|
| 22 |
A' |
955 |
915 |
1.52 |
|
|
|
| 23 |
A' |
908 |
871 |
3.23 |
|
|
|
| 24 |
A' |
572 |
549 |
3.42 |
|
|
|
| 25 |
A' |
471 |
452 |
0.46 |
|
|
|
| 26 |
A' |
308 |
295 |
0.75 |
|
|
|
| 27 |
A' |
150 |
144 |
0.77 |
|
|
|
| 28 |
A" |
3138 |
3007 |
27.91 |
|
|
|
| 29 |
A" |
3056 |
2928 |
13.10 |
|
|
|
| 30 |
A" |
1479 |
1417 |
7.98 |
|
|
|
| 31 |
A" |
1280 |
1227 |
0.26 |
|
|
|
| 32 |
A" |
1095 |
1049 |
1.65 |
|
|
|
| 33 |
A" |
1044 |
1001 |
44.30 |
|
|
|
| 34 |
A" |
988 |
947 |
10.68 |
|
|
|
| 35 |
A" |
938 |
899 |
37.03 |
|
|
|
| 36 |
A" |
884 |
847 |
2.11 |
|
|
|
| 37 |
A" |
762 |
730 |
7.59 |
|
|
|
| 38 |
A" |
641 |
614 |
1.77 |
|
|
|
| 39 |
A" |
307 |
294 |
1.57 |
|
|
|
| 40 |
A" |
241 |
231 |
0.14 |
|
|
|
| 41 |
A" |
167 |
160 |
1.50 |
|
|
|
| 42 |
A" |
120 |
115 |
0.42 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31279.3 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 29974.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
-0.464 |
|
|
|
| 2 |
H |
-0.495 |
|
|
|
| 3 |
C |
0.300 |
|
|
|
| 4 |
H |
-0.689 |
|
|
|
| 5 |
C |
1.040 |
|
|
|
| 6 |
H |
-0.940 |
|
|
|
| 7 |
C |
1.385 |
|
|
|
| 8 |
H |
-0.742 |
|
|
|
| 9 |
C |
0.319 |
|
|
|
| 10 |
H |
-0.015 |
|
|
|
| 11 |
H |
-0.015 |
|
|
|
| 12 |
C |
0.111 |
|
|
|
| 13 |
H |
-0.262 |
|
|
|
| 14 |
H |
-0.018 |
|
|
|
| 15 |
H |
-0.018 |
|
|
|
| 16 |
C |
0.502 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.146 |
-0.839 |
0.000 |
0.851 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-36.400 |
0.306 |
0.000 |
| y |
0.306 |
-36.678 |
0.000 |
| z |
0.000 |
0.000 |
-41.687 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.783 |
0.306 |
0.000 |
| y |
0.306 |
2.365 |
0.000 |
| z |
0.000 |
0.000 |
-5.148 |
|
| Polar |
|---|
| 3z2-r2 | -10.296 |
|---|
| x2-y2 | 0.278 |
|---|
| xy | 0.306 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.902 |
1.633 |
0.000 |
| y |
1.633 |
17.984 |
0.000 |
| z |
0.000 |
0.000 |
8.313 |
<r2> (average value of r
2) Å
2
| <r2> |
249.171 |
| (<r2>)1/2 |
15.785 |