Jump to
S1C2
Energy calculated at HF/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -133.096848 |
| Energy at 298.15K | -133.102441 |
| HF Energy | -133.096849 |
| Nuclear repulsion energy | 71.246712 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3718 |
3386 |
2.15 |
|
|
|
| 2 |
A' |
3282 |
2989 |
20.55 |
|
|
|
| 3 |
A' |
3194 |
2908 |
54.03 |
|
|
|
| 4 |
A' |
3164 |
2881 |
16.62 |
|
|
|
| 5 |
A' |
1886 |
1717 |
106.87 |
|
|
|
| 6 |
A' |
1565 |
1425 |
9.43 |
|
|
|
| 7 |
A' |
1548 |
1410 |
40.15 |
|
|
|
| 8 |
A' |
1494 |
1360 |
6.67 |
|
|
|
| 9 |
A' |
1368 |
1246 |
22.87 |
|
|
|
| 10 |
A' |
1141 |
1039 |
27.18 |
|
|
|
| 11 |
A' |
980 |
892 |
5.56 |
|
|
|
| 12 |
A' |
517 |
471 |
21.22 |
|
|
|
| 13 |
A" |
3222 |
2934 |
25.97 |
|
|
|
| 14 |
A" |
1572 |
1432 |
8.06 |
|
|
|
| 15 |
A" |
1232 |
1122 |
2.42 |
|
|
|
| 16 |
A" |
1155 |
1052 |
19.69 |
|
|
|
| 17 |
A" |
736 |
670 |
56.19 |
|
|
|
| 18 |
A" |
201 |
183 |
3.23 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 15987.1 cm
-1
Scaled (by 0.9106) Zero Point Vibrational Energy (zpe) 14557.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.024 |
-0.650 |
0.000 |
| C2 |
0.000 |
0.444 |
0.000 |
| N3 |
1.234 |
0.208 |
0.000 |
| H4 |
-0.537 |
-1.622 |
0.000 |
| H5 |
-1.666 |
-0.562 |
0.880 |
| H6 |
-1.666 |
-0.562 |
-0.880 |
| H7 |
-0.393 |
1.465 |
0.000 |
| H8 |
1.770 |
1.060 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4991 | 2.4157 | 1.0871 | 1.0922 | 1.0922 | 2.2069 | 3.2762 |
C2 | 1.4991 | | 1.2564 | 2.1352 | 2.1356 | 2.1356 | 1.0934 | 1.8740 | N3 | 2.4157 | 1.2564 | | 2.5467 | 3.1263 | 3.1263 | 2.0561 | 1.0069 | H4 | 1.0871 | 2.1352 | 2.5467 | | 1.7807 | 1.7807 | 3.0900 | 3.5381 | H5 | 1.0922 | 2.1356 | 3.1263 | 1.7807 | | 1.7593 | 2.5494 | 3.8997 | H6 | 1.0922 | 2.1356 | 3.1263 | 1.7807 | 1.7593 | | 2.5494 | 3.8997 | H7 | 2.2069 | 1.0934 | 2.0561 | 3.0900 | 2.5494 | 2.5494 | | 2.2007 | H8 | 3.2762 | 1.8740 | 1.0069 | 3.5381 | 3.8997 | 3.8997 | 2.2007 | |
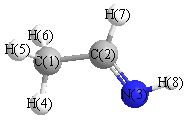 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
122.246 |
|
C1 |
C2 |
H7 |
115.817 |
| C2 |
C1 |
H4 |
110.284 |
|
C2 |
C1 |
H5 |
110.010 |
| C2 |
C1 |
H6 |
110.010 |
|
C2 |
N3 |
H8 |
111.308 |
| N3 |
C2 |
H7 |
121.936 |
|
H4 |
C1 |
H5 |
109.594 |
| H4 |
C1 |
H6 |
109.594 |
|
H5 |
C1 |
H6 |
107.297 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.175 |
|
|
|
| 2 |
C |
0.566 |
|
|
|
| 3 |
N |
-0.503 |
|
|
|
| 4 |
H |
-0.053 |
|
|
|
| 5 |
H |
0.016 |
|
|
|
| 6 |
H |
0.016 |
|
|
|
| 7 |
H |
-0.242 |
|
|
|
| 8 |
H |
0.024 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.728 |
1.459 |
0.000 |
2.262 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-20.334 |
2.492 |
0.000 |
| y |
2.492 |
-17.751 |
0.000 |
| z |
0.000 |
0.000 |
-20.188 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.364 |
2.492 |
0.000 |
| y |
2.492 |
2.509 |
0.000 |
| z |
0.000 |
0.000 |
-1.145 |
|
| Polar |
|---|
| 3z2-r2 | -2.290 |
|---|
| x2-y2 | -2.582 |
|---|
| xy | 2.492 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.258 |
0.245 |
0.000 |
| y |
0.245 |
4.705 |
0.000 |
| z |
0.000 |
0.000 |
3.925 |
<r2> (average value of r
2) Å
2
| <r2> |
50.313 |
| (<r2>)1/2 |
7.093 |
Jump to
S1C1
Energy calculated at HF/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -133.095629 |
| Energy at 298.15K | -133.101227 |
| HF Energy | -133.095629 |
| Nuclear repulsion energy | 71.255148 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3686 |
3357 |
1.93 |
|
|
|
| 2 |
A' |
3261 |
2969 |
56.48 |
|
|
|
| 3 |
A' |
3238 |
2948 |
8.53 |
|
|
|
| 4 |
A' |
3165 |
2882 |
13.38 |
|
|
|
| 5 |
A' |
1888 |
1719 |
107.54 |
|
|
|
| 6 |
A' |
1573 |
1432 |
27.73 |
|
|
|
| 7 |
A' |
1533 |
1396 |
17.53 |
|
|
|
| 8 |
A' |
1501 |
1367 |
1.76 |
|
|
|
| 9 |
A' |
1378 |
1255 |
61.47 |
|
|
|
| 10 |
A' |
1143 |
1041 |
21.12 |
|
|
|
| 11 |
A' |
970 |
884 |
1.56 |
|
|
|
| 12 |
A' |
524 |
477 |
8.78 |
|
|
|
| 13 |
A" |
3229 |
2941 |
21.02 |
|
|
|
| 14 |
A" |
1567 |
1427 |
8.51 |
|
|
|
| 15 |
A" |
1240 |
1129 |
44.93 |
|
|
|
| 16 |
A" |
1160 |
1057 |
18.77 |
|
|
|
| 17 |
A" |
753 |
686 |
13.41 |
|
|
|
| 18 |
A" |
197 |
179 |
0.10 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16003.2 cm
-1
Scaled (by 0.9106) Zero Point Vibrational Energy (zpe) 14572.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.010 |
-0.631 |
0.000 |
| C2 |
0.000 |
0.483 |
0.000 |
| N3 |
1.252 |
0.384 |
0.000 |
| H4 |
-0.528 |
-1.608 |
0.000 |
| H5 |
-1.652 |
-0.551 |
0.879 |
| H6 |
-1.652 |
-0.551 |
-0.879 |
| H7 |
-0.414 |
1.491 |
0.000 |
| H8 |
1.544 |
-0.582 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5039 | 2.4788 | 1.0894 | 1.0918 | 1.0918 | 2.2042 | 2.5547 |
C2 | 1.5039 | | 1.2557 | 2.1569 | 2.1382 | 2.1382 | 1.0896 | 1.8762 | N3 | 2.4788 | 1.2557 | | 2.6709 | 3.1747 | 3.1747 | 2.0004 | 1.0092 | H4 | 1.0894 | 2.1569 | 2.6709 | | 1.7761 | 1.7761 | 3.1012 | 2.3122 | H5 | 1.0918 | 2.1382 | 3.1747 | 1.7761 | | 1.7585 | 2.5446 | 3.3153 | H6 | 1.0918 | 2.1382 | 3.1747 | 1.7761 | 1.7585 | | 2.5446 | 3.3153 | H7 | 2.2042 | 1.0896 | 2.0004 | 3.1012 | 2.5446 | 2.5446 | | 2.8521 | H8 | 2.5547 | 1.8762 | 1.0092 | 2.3122 | 3.3153 | 3.3153 | 2.8521 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
127.635 |
|
C1 |
C2 |
H7 |
115.473 |
| C2 |
C1 |
H4 |
111.554 |
|
C2 |
C1 |
H5 |
109.910 |
| C2 |
C1 |
H6 |
109.910 |
|
C2 |
N3 |
H8 |
111.402 |
| N3 |
C2 |
H7 |
116.892 |
|
H4 |
C1 |
H5 |
109.039 |
| H4 |
C1 |
H6 |
109.039 |
|
H5 |
C1 |
H6 |
107.285 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.329 |
|
|
|
| 2 |
C |
0.440 |
|
|
|
| 3 |
N |
-0.433 |
|
|
|
| 4 |
H |
-0.146 |
|
|
|
| 5 |
H |
0.018 |
|
|
|
| 6 |
H |
0.018 |
|
|
|
| 7 |
H |
-0.230 |
|
|
|
| 8 |
H |
0.003 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.273 |
-1.483 |
0.000 |
2.714 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-21.712 |
-3.511 |
0.000 |
| y |
-3.511 |
-19.123 |
0.000 |
| z |
0.000 |
0.000 |
-20.132 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.084 |
-3.511 |
0.000 |
| y |
-3.511 |
1.799 |
0.000 |
| z |
0.000 |
0.000 |
0.285 |
|
| Polar |
|---|
| 3z2-r2 | 0.570 |
|---|
| x2-y2 | -2.589 |
|---|
| xy | -3.511 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.180 |
0.072 |
0.000 |
| y |
0.072 |
4.693 |
0.000 |
| z |
0.000 |
0.000 |
3.903 |
<r2> (average value of r
2) Å
2
| <r2> |
50.197 |
| (<r2>)1/2 |
7.085 |
