Jump to
S1C2
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at mPW1PW91/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -133.935786 |
| Energy at 298.15K | -133.941245 |
| HF Energy | -133.935786 |
| Nuclear repulsion energy | 70.850874 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3447 |
3303 |
4.42 |
|
|
|
| 2 |
A' |
3154 |
3022 |
22.58 |
|
|
|
| 3 |
A' |
3095 |
2966 |
26.19 |
|
|
|
| 4 |
A' |
3048 |
2921 |
7.92 |
|
|
|
| 5 |
A' |
1752 |
1678 |
78.99 |
|
|
|
| 6 |
A' |
1451 |
1391 |
31.19 |
|
|
|
| 7 |
A' |
1424 |
1365 |
18.54 |
|
|
|
| 8 |
A' |
1372 |
1315 |
7.97 |
|
|
|
| 9 |
A' |
1277 |
1224 |
51.77 |
|
|
|
| 10 |
A' |
1070 |
1025 |
17.51 |
|
|
|
| 11 |
A' |
919 |
881 |
4.90 |
|
|
|
| 12 |
A' |
489 |
469 |
7.03 |
|
|
|
| 13 |
A" |
3124 |
2993 |
8.69 |
|
|
|
| 14 |
A" |
1445 |
1385 |
9.83 |
|
|
|
| 15 |
A" |
1144 |
1096 |
46.55 |
|
|
|
| 16 |
A" |
1061 |
1017 |
9.34 |
|
|
|
| 17 |
A" |
690 |
661 |
7.26 |
|
|
|
| 18 |
A" |
183 |
176 |
0.27 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 15072.4 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 14443.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.019 |
-0.617 |
0.000 |
| C2 |
0.000 |
0.480 |
0.000 |
| N3 |
1.266 |
0.372 |
0.000 |
| H4 |
-0.546 |
-1.607 |
0.000 |
| H5 |
-1.669 |
-0.530 |
0.882 |
| H6 |
-1.669 |
-0.530 |
-0.882 |
| H7 |
-0.404 |
1.503 |
0.000 |
| H8 |
1.539 |
-0.616 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4974 | 2.4899 | 1.0967 | 1.0993 | 1.0993 | 2.2078 | 2.5577 |
C2 | 1.4974 | | 1.2708 | 2.1573 | 2.1414 | 2.1414 | 1.0998 | 1.8896 | N3 | 2.4899 | 1.2708 | | 2.6834 | 3.1950 | 3.1950 | 2.0170 | 1.0253 | H4 | 1.0967 | 2.1573 | 2.6834 | | 1.7881 | 1.7881 | 3.1133 | 2.3084 | H5 | 1.0993 | 2.1414 | 3.1950 | 1.7881 | | 1.7641 | 2.5525 | 3.3281 | H6 | 1.0993 | 2.1414 | 3.1950 | 1.7881 | 1.7641 | | 2.5525 | 3.3281 | H7 | 2.2078 | 1.0998 | 2.0170 | 3.1133 | 2.5525 | 2.5525 | | 2.8751 | H8 | 2.5577 | 1.8896 | 1.0253 | 2.3084 | 3.3281 | 3.3281 | 2.8751 | |
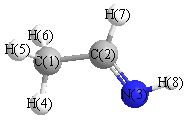 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
127.988 |
|
C1 |
C2 |
H7 |
115.591 |
| C2 |
C1 |
H4 |
111.602 |
|
C2 |
C1 |
H5 |
110.163 |
| C2 |
C1 |
H6 |
110.163 |
|
C2 |
N3 |
H8 |
110.312 |
| N3 |
C2 |
H7 |
116.422 |
|
H4 |
C1 |
H5 |
109.029 |
| H4 |
C1 |
H6 |
109.029 |
|
H5 |
C1 |
H6 |
106.720 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.384 |
|
|
|
| 2 |
C |
0.523 |
|
|
|
| 3 |
N |
-0.340 |
|
|
|
| 4 |
H |
-0.194 |
|
|
|
| 5 |
H |
-0.030 |
|
|
|
| 6 |
H |
-0.030 |
|
|
|
| 7 |
H |
-0.304 |
|
|
|
| 8 |
H |
-0.010 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.182 |
-1.428 |
0.000 |
2.608 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-21.551 |
-3.258 |
0.000 |
| y |
-3.258 |
-18.997 |
0.000 |
| z |
0.000 |
0.000 |
-19.916 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.094 |
-3.258 |
0.000 |
| y |
-3.258 |
1.736 |
0.000 |
| z |
0.000 |
0.000 |
0.358 |
|
| Polar |
|---|
| 3z2-r2 | 0.716 |
|---|
| x2-y2 | -2.554 |
|---|
| xy | -3.258 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.552 |
0.170 |
0.000 |
| y |
0.170 |
5.107 |
0.000 |
| z |
0.000 |
0.000 |
4.066 |
<r2> (average value of r
2) Å
2
| <r2> |
50.417 |
| (<r2>)1/2 |
7.100 |
