Jump to
S1C2
Vibrational Frequencies calculated at MP4/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3405 |
3326 |
|
|
|
|
| 2 |
A' |
3172 |
3099 |
|
|
|
|
| 3 |
A' |
3058 |
2988 |
|
|
|
|
| 4 |
A' |
3042 |
2971 |
|
|
|
|
| 5 |
A' |
1705 |
1666 |
|
|
|
|
| 6 |
A' |
1522 |
1487 |
|
|
|
|
| 7 |
A' |
1462 |
1428 |
|
|
|
|
| 8 |
A' |
1427 |
1394 |
|
|
|
|
| 9 |
A' |
1303 |
1273 |
|
|
|
|
| 10 |
A' |
1083 |
1058 |
|
|
|
|
| 11 |
A' |
949 |
927 |
|
|
|
|
| 12 |
A' |
490 |
478 |
|
|
|
|
| 13 |
A" |
3125 |
3053 |
|
|
|
|
| 14 |
A" |
1524 |
1489 |
|
|
|
|
| 15 |
A" |
1125 |
1099 |
|
|
|
|
| 16 |
A" |
1099 |
1074 |
|
|
|
|
| 17 |
A" |
688 |
672 |
|
|
|
|
| 18 |
A" |
182 |
178 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 15179.3 cm
-1
Scaled (by 0.9769) Zero Point Vibrational Energy (zpe) 14828.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP4/aug-cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP4/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -133.610064 |
| Energy at 298.15K | -133.615486 |
| HF Energy | -133.092373 |
| Nuclear repulsion energy | 70.011121 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3362 |
3284 |
|
|
|
|
| 2 |
A' |
3144 |
3071 |
|
|
|
|
| 3 |
A' |
3114 |
3042 |
|
|
|
|
| 4 |
A' |
3050 |
2980 |
|
|
|
|
| 5 |
A' |
1699 |
1660 |
|
|
|
|
| 6 |
A' |
1528 |
1492 |
|
|
|
|
| 7 |
A' |
1455 |
1422 |
|
|
|
|
| 8 |
A' |
1439 |
1405 |
|
|
|
|
| 9 |
A' |
1307 |
1277 |
|
|
|
|
| 10 |
A' |
1087 |
1062 |
|
|
|
|
| 11 |
A' |
928 |
906 |
|
|
|
|
| 12 |
A' |
494 |
482 |
|
|
|
|
| 13 |
A" |
3127 |
3055 |
|
|
|
|
| 14 |
A" |
1518 |
1482 |
|
|
|
|
| 15 |
A" |
1154 |
1127 |
|
|
|
|
| 16 |
A" |
1095 |
1070 |
|
|
|
|
| 17 |
A" |
686 |
670 |
|
|
|
|
| 18 |
A" |
161 |
157 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 15172.4 cm
-1
Scaled (by 0.9769) Zero Point Vibrational Energy (zpe) 14821.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP4/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.037 |
-0.617 |
0.000 |
| C2 |
0.000 |
0.489 |
0.000 |
| N3 |
1.288 |
0.365 |
0.000 |
| H4 |
-0.559 |
-1.612 |
0.000 |
| H5 |
-1.687 |
-0.528 |
0.891 |
| H6 |
-1.687 |
-0.528 |
-0.891 |
| H7 |
-0.391 |
1.522 |
0.000 |
| H8 |
1.534 |
-0.639 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5161 | 2.5240 | 1.1048 | 1.1063 | 1.1063 | 2.2345 | 2.5718 |
C2 | 1.5161 | | 1.2938 | 2.1741 | 2.1621 | 2.1621 | 1.1048 | 1.9042 | N3 | 2.5240 | 1.2938 | | 2.7053 | 3.2315 | 3.2315 | 2.0388 | 1.0338 | H4 | 1.1048 | 2.1741 | 2.7053 | | 1.8010 | 1.8010 | 3.1390 | 2.3080 | H5 | 1.1063 | 2.1621 | 3.2315 | 1.8010 | | 1.7818 | 2.5842 | 3.3443 | H6 | 1.1063 | 2.1621 | 3.2315 | 1.8010 | 1.7818 | | 2.5842 | 3.3443 | H7 | 2.2345 | 1.1048 | 2.0388 | 3.1390 | 2.5842 | 2.5842 | | 2.8942 | H8 | 2.5718 | 1.9042 | 1.0338 | 2.3080 | 3.3443 | 3.3443 | 2.8942 | |
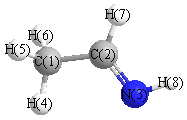 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
127.688 |
|
C1 |
C2 |
H7 |
116.105 |
| C2 |
C1 |
H4 |
111.131 |
|
C2 |
C1 |
H5 |
110.094 |
| C2 |
C1 |
H6 |
110.094 |
|
C2 |
N3 |
H8 |
109.281 |
| N3 |
C2 |
H7 |
116.208 |
|
H4 |
C1 |
H5 |
109.077 |
| H4 |
C1 |
H6 |
109.077 |
|
H5 |
C1 |
H6 |
107.278 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
