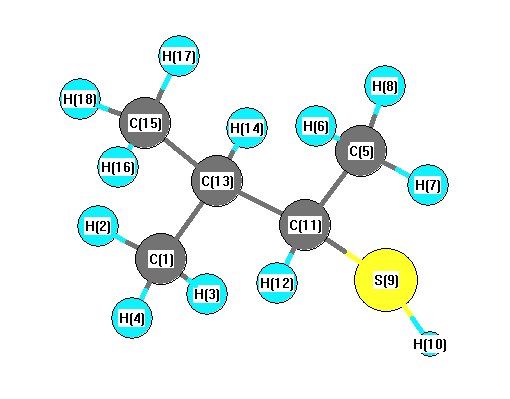
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3153 |
3022 |
25.27 |
|
|
|
| 2 |
A |
3150 |
3018 |
28.13 |
|
|
|
| 3 |
A |
3146 |
3015 |
22.14 |
|
|
|
| 4 |
A |
3139 |
3008 |
26.22 |
|
|
|
| 5 |
A |
3132 |
3002 |
54.02 |
|
|
|
| 6 |
A |
3126 |
2996 |
1.47 |
|
|
|
| 7 |
A |
3070 |
2942 |
24.50 |
|
|
|
| 8 |
A |
3058 |
2930 |
19.50 |
|
|
|
| 9 |
A |
3053 |
2926 |
25.90 |
|
|
|
| 10 |
A |
3049 |
2922 |
17.97 |
|
|
|
| 11 |
A |
3041 |
2914 |
2.87 |
|
|
|
| 12 |
A |
2709 |
2596 |
5.10 |
|
|
|
| 13 |
A |
1501 |
1438 |
3.90 |
|
|
|
| 14 |
A |
1486 |
1424 |
3.54 |
|
|
|
| 15 |
A |
1480 |
1418 |
22.22 |
|
|
|
| 16 |
A |
1471 |
1410 |
2.13 |
|
|
|
| 17 |
A |
1468 |
1407 |
4.41 |
|
|
|
| 18 |
A |
1463 |
1402 |
1.35 |
|
|
|
| 19 |
A |
1407 |
1349 |
7.73 |
|
|
|
| 20 |
A |
1395 |
1337 |
11.37 |
|
|
|
| 21 |
A |
1390 |
1332 |
3.60 |
|
|
|
| 22 |
A |
1373 |
1316 |
1.31 |
|
|
|
| 23 |
A |
1339 |
1283 |
0.55 |
|
|
|
| 24 |
A |
1307 |
1252 |
3.26 |
|
|
|
| 25 |
A |
1239 |
1188 |
22.83 |
|
|
|
| 26 |
A |
1201 |
1151 |
1.98 |
|
|
|
| 27 |
A |
1175 |
1126 |
6.24 |
|
|
|
| 28 |
A |
1160 |
1112 |
1.87 |
|
|
|
| 29 |
A |
1095 |
1050 |
1.26 |
|
|
|
| 30 |
A |
1040 |
996 |
12.07 |
|
|
|
| 31 |
A |
979 |
938 |
2.61 |
|
|
|
| 32 |
A |
969 |
929 |
0.91 |
|
|
|
| 33 |
A |
938 |
899 |
1.96 |
|
|
|
| 34 |
A |
922 |
884 |
0.99 |
|
|
|
| 35 |
A |
889 |
852 |
2.00 |
|
|
|
| 36 |
A |
800 |
767 |
3.07 |
|
|
|
| 37 |
A |
695 |
666 |
2.39 |
|
|
|
| 38 |
A |
480 |
460 |
0.39 |
|
|
|
| 39 |
A |
407 |
390 |
0.43 |
|
|
|
| 40 |
A |
376 |
360 |
0.12 |
|
|
|
| 41 |
A |
361 |
345 |
0.14 |
|
|
|
| 42 |
A |
332 |
319 |
1.11 |
|
|
|
| 43 |
A |
252 |
241 |
0.93 |
|
|
|
| 44 |
A |
235 |
226 |
1.93 |
|
|
|
| 45 |
A |
227 |
218 |
0.17 |
|
|
|
| 46 |
A |
196 |
188 |
1.21 |
|
|
|
| 47 |
A |
163 |
156 |
11.86 |
|
|
|
| 48 |
A |
60 |
58 |
1.97 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35048.6 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 33587.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.752 |
|
|
|
| 2 |
H |
-0.122 |
|
|
|
| 3 |
H |
-0.162 |
|
|
|
| 4 |
H |
-0.068 |
|
|
|
| 5 |
C |
0.624 |
|
|
|
| 6 |
H |
-0.141 |
|
|
|
| 7 |
H |
-0.219 |
|
|
|
| 8 |
H |
-0.077 |
|
|
|
| 9 |
S |
0.095 |
|
|
|
| 10 |
H |
0.048 |
|
|
|
| 11 |
C |
0.071 |
|
|
|
| 12 |
H |
-0.394 |
|
|
|
| 13 |
C |
-0.415 |
|
|
|
| 14 |
H |
-0.458 |
|
|
|
| 15 |
C |
0.854 |
|
|
|
| 16 |
H |
-0.057 |
|
|
|
| 17 |
H |
-0.161 |
|
|
|
| 18 |
H |
-0.166 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.013 |
1.308 |
0.554 |
1.744 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-47.458 |
2.615 |
1.708 |
| y |
2.615 |
-48.408 |
0.855 |
| z |
1.708 |
0.855 |
-48.306 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.899 |
2.615 |
1.708 |
| y |
2.615 |
-0.526 |
0.855 |
| z |
1.708 |
0.855 |
-0.373 |
|
| Polar |
|---|
| 3z2-r2 | -0.745 |
|---|
| x2-y2 | 0.950 |
|---|
| xy | 2.615 |
|---|
| xz | 1.708 |
|---|
| yz | 0.855 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
14.352 |
-0.313 |
-0.052 |
| y |
-0.313 |
13.000 |
-0.024 |
| z |
-0.052 |
-0.024 |
10.740 |
<r2> (average value of r
2) Å
2
| <r2> |
249.479 |
| (<r2>)1/2 |
15.795 |