Jump to
S1C2
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at mPW1PW91/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -193.439486 |
| Energy at 298.15K | |
| HF Energy | -193.439486 |
| Nuclear repulsion energy | 148.625447 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3280 |
3143 |
0.02 |
|
|
|
| 2 |
A1 |
3272 |
3135 |
6.16 |
|
|
|
| 3 |
A1 |
3238 |
3103 |
0.90 |
|
|
|
| 4 |
A1 |
1585 |
1519 |
0.18 |
|
|
|
| 5 |
A1 |
1465 |
1404 |
3.29 |
|
|
|
| 6 |
A1 |
1149 |
1101 |
0.03 |
|
|
|
| 7 |
A1 |
1081 |
1035 |
7.78 |
|
|
|
| 8 |
A1 |
1043 |
1000 |
13.77 |
|
|
|
| 9 |
A1 |
837 |
803 |
0.32 |
|
|
|
| 10 |
A2 |
892 |
854 |
0.00 |
|
|
|
| 11 |
A2 |
808 |
775 |
0.00 |
|
|
|
| 12 |
A2 |
522 |
500 |
0.00 |
|
|
|
| 13 |
B1 |
863 |
827 |
7.68 |
|
|
|
| 14 |
B1 |
713 |
684 |
0.93 |
|
|
|
| 15 |
B1 |
675 |
647 |
114.35 |
|
|
|
| 16 |
B1 |
483 |
463 |
11.11 |
|
|
|
| 17 |
B2 |
3254 |
3119 |
12.67 |
|
|
|
| 18 |
B2 |
3239 |
3104 |
6.74 |
|
|
|
| 19 |
B2 |
1389 |
1331 |
40.41 |
|
|
|
| 20 |
B2 |
1274 |
1221 |
0.01 |
|
|
|
| 21 |
B2 |
1208 |
1157 |
1.41 |
|
|
|
| 22 |
B2 |
972 |
932 |
0.01 |
|
|
|
| 23 |
B2 |
919 |
881 |
3.75 |
|
|
|
| 24 |
B2 |
32i |
31i |
4.40 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17063.7 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 16352.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
1.225 |
| C2 |
0.000 |
1.124 |
0.392 |
| C3 |
0.000 |
0.680 |
-1.005 |
| C4 |
0.000 |
-0.680 |
-1.005 |
| C5 |
0.000 |
-1.124 |
0.392 |
| H6 |
0.000 |
0.000 |
2.310 |
| H7 |
0.000 |
2.163 |
0.717 |
| H8 |
0.000 |
1.337 |
-1.870 |
| H9 |
0.000 |
-1.337 |
-1.870 |
| H10 |
0.000 |
-2.163 |
0.717 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.3989 | 2.3311 | 2.3311 | 1.3989 | 1.0851 | 2.2220 | 3.3713 | 3.3713 | 2.2220 |
C2 | 1.3989 | | 1.4663 | 2.2821 | 2.2486 | 2.2228 | 1.0883 | 2.2726 | 3.3431 | 3.3034 | C3 | 2.3311 | 1.4663 | | 1.3600 | 2.2821 | 3.3839 | 2.2725 | 1.0863 | 2.1945 | 3.3238 | C4 | 2.3311 | 2.2821 | 1.3600 | | 1.4663 | 3.3839 | 3.3238 | 2.1945 | 1.0863 | 2.2725 | C5 | 1.3989 | 2.2486 | 2.2821 | 1.4663 | | 2.2228 | 3.3034 | 3.3431 | 2.2726 | 1.0883 | H6 | 1.0851 | 2.2228 | 3.3839 | 3.3839 | 2.2228 | | 2.6865 | 4.3886 | 4.3886 | 2.6865 | H7 | 2.2220 | 1.0883 | 2.2725 | 3.3238 | 3.3034 | 2.6865 | | 2.7158 | 4.3522 | 4.3263 | H8 | 3.3713 | 2.2726 | 1.0863 | 2.1945 | 3.3431 | 4.3886 | 2.7158 | | 2.6735 | 4.3522 | H9 | 3.3713 | 3.3431 | 2.1945 | 1.0863 | 2.2726 | 4.3886 | 4.3522 | 2.6735 | | 2.7158 | H10 | 2.2220 | 3.3034 | 3.3238 | 2.2725 | 1.0883 | 2.6865 | 4.3263 | 4.3522 | 2.7158 | |
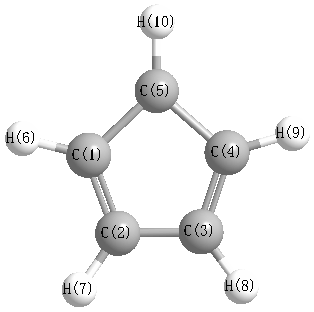 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
108.874 |
|
C1 |
C2 |
H7 |
126.146 |
| C1 |
C5 |
C4 |
108.874 |
|
C1 |
C5 |
H10 |
126.146 |
| C2 |
C1 |
C5 |
106.973 |
|
C2 |
C1 |
H6 |
126.514 |
| C2 |
C3 |
C4 |
107.639 |
|
C2 |
C3 |
H8 |
125.161 |
| C3 |
C2 |
H7 |
124.980 |
|
C3 |
C4 |
C5 |
107.639 |
| C3 |
C4 |
H9 |
127.200 |
|
C4 |
C3 |
H8 |
127.200 |
| C4 |
C5 |
H10 |
124.980 |
|
C5 |
C1 |
H6 |
126.514 |
| C5 |
C4 |
H9 |
125.161 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.409 |
|
|
|
| 2 |
C |
0.379 |
|
|
|
| 3 |
C |
0.385 |
|
|
|
| 4 |
C |
0.385 |
|
|
|
| 5 |
C |
0.379 |
|
|
|
| 6 |
H |
-0.395 |
|
|
|
| 7 |
H |
-0.381 |
|
|
|
| 8 |
H |
-0.390 |
|
|
|
| 9 |
H |
-0.390 |
|
|
|
| 10 |
H |
-0.381 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.037 |
0.037 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.494 |
0.000 |
0.000 |
| y |
0.000 |
-25.629 |
0.000 |
| z |
0.000 |
0.000 |
-27.777 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.791 |
0.000 |
0.000 |
| y |
0.000 |
5.006 |
0.000 |
| z |
0.000 |
0.000 |
1.785 |
|
| Polar |
|---|
| 3z2-r2 | 3.570 |
|---|
| x2-y2 | -7.864 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.809 |
0.000 |
0.000 |
| y |
0.000 |
9.971 |
0.000 |
| z |
0.000 |
0.000 |
9.365 |
<r2> (average value of r
2) Å
2
| <r2> |
88.067 |
| (<r2>)1/2 |
9.384 |
