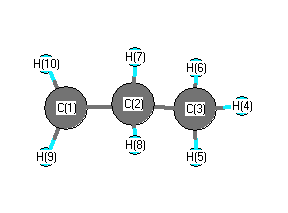
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3279 |
3142 |
13.63 |
|
|
|
| 2 |
A |
3167 |
3035 |
16.99 |
|
|
|
| 3 |
A |
3142 |
3011 |
28.80 |
|
|
|
| 4 |
A |
3134 |
3004 |
33.06 |
|
|
|
| 5 |
A |
3056 |
2928 |
27.61 |
|
|
|
| 6 |
A |
3039 |
2912 |
21.20 |
|
|
|
| 7 |
A |
2981 |
2857 |
31.62 |
|
|
|
| 8 |
A |
1483 |
1422 |
2.94 |
|
|
|
| 9 |
A |
1475 |
1413 |
7.01 |
|
|
|
| 10 |
A |
1457 |
1396 |
2.49 |
|
|
|
| 11 |
A |
1446 |
1385 |
3.72 |
|
|
|
| 12 |
A |
1396 |
1338 |
3.52 |
|
|
|
| 13 |
A |
1349 |
1293 |
1.95 |
|
|
|
| 14 |
A |
1257 |
1205 |
0.06 |
|
|
|
| 15 |
A |
1168 |
1119 |
0.10 |
|
|
|
| 16 |
A |
1110 |
1064 |
0.28 |
|
|
|
| 17 |
A |
1038 |
995 |
1.05 |
|
|
|
| 18 |
A |
926 |
887 |
1.44 |
|
|
|
| 19 |
A |
891 |
854 |
0.21 |
|
|
|
| 20 |
A |
753 |
722 |
0.56 |
|
|
|
| 21 |
A |
485 |
465 |
50.96 |
|
|
|
| 22 |
A |
374 |
358 |
1.25 |
|
|
|
| 23 |
A |
247 |
236 |
0.04 |
|
|
|
| 24 |
A |
66 |
63 |
1.50 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19358.1 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 18550.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.491 |
|
|
|
| 2 |
C |
0.451 |
|
|
|
| 3 |
C |
0.426 |
|
|
|
| 4 |
H |
-0.152 |
|
|
|
| 5 |
H |
-0.105 |
|
|
|
| 6 |
H |
-0.112 |
|
|
|
| 7 |
H |
-0.149 |
|
|
|
| 8 |
H |
-0.162 |
|
|
|
| 9 |
H |
-0.340 |
|
|
|
| 10 |
H |
-0.348 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.214 |
0.242 |
0.083 |
0.334 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-21.327 |
0.167 |
-0.052 |
| y |
0.167 |
-20.654 |
-0.323 |
| z |
-0.052 |
-0.323 |
-21.768 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.116 |
0.167 |
-0.052 |
| y |
0.167 |
0.893 |
-0.323 |
| z |
-0.052 |
-0.323 |
-0.778 |
|
| Polar |
|---|
| 3z2-r2 | -1.556 |
|---|
| x2-y2 | -0.673 |
|---|
| xy | 0.167 |
|---|
| xz | -0.052 |
|---|
| yz | -0.323 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.684 |
-0.133 |
-0.034 |
| y |
-0.133 |
5.916 |
0.042 |
| z |
-0.034 |
0.042 |
5.434 |
<r2> (average value of r
2) Å
2
| <r2> |
59.447 |
| (<r2>)1/2 |
7.710 |