Jump to
S1C2
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at mPW1PW91/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -235.817176 |
| Energy at 298.15K | -235.830586 |
| HF Energy | -235.817176 |
| Nuclear repulsion energy | 249.671580 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3131 |
3000 |
114.97 |
|
|
|
| 2 |
A' |
3123 |
2992 |
22.83 |
|
|
|
| 3 |
A' |
3119 |
2989 |
25.07 |
|
|
|
| 4 |
A' |
3084 |
2955 |
18.80 |
|
|
|
| 5 |
A' |
3072 |
2944 |
11.25 |
|
|
|
| 6 |
A' |
3065 |
2937 |
5.78 |
|
|
|
| 7 |
A' |
3042 |
2915 |
33.12 |
|
|
|
| 8 |
A' |
3040 |
2913 |
32.63 |
|
|
|
| 9 |
A' |
1490 |
1428 |
5.12 |
|
|
|
| 10 |
A' |
1479 |
1417 |
2.84 |
|
|
|
| 11 |
A' |
1476 |
1414 |
6.60 |
|
|
|
| 12 |
A' |
1401 |
1343 |
3.11 |
|
|
|
| 13 |
A' |
1398 |
1340 |
10.14 |
|
|
|
| 14 |
A' |
1373 |
1316 |
0.35 |
|
|
|
| 15 |
A' |
1360 |
1303 |
7.81 |
|
|
|
| 16 |
A' |
1273 |
1220 |
2.95 |
|
|
|
| 17 |
A' |
1250 |
1198 |
0.50 |
|
|
|
| 18 |
A' |
1199 |
1149 |
0.17 |
|
|
|
| 19 |
A' |
1153 |
1105 |
0.28 |
|
|
|
| 20 |
A' |
1057 |
1013 |
0.46 |
|
|
|
| 21 |
A' |
958 |
918 |
0.92 |
|
|
|
| 22 |
A' |
948 |
909 |
0.62 |
|
|
|
| 23 |
A' |
873 |
837 |
0.94 |
|
|
|
| 24 |
A' |
814 |
780 |
1.05 |
|
|
|
| 25 |
A' |
651 |
624 |
1.61 |
|
|
|
| 26 |
A' |
431 |
413 |
0.07 |
|
|
|
| 27 |
A' |
327 |
313 |
0.10 |
|
|
|
| 28 |
A' |
95 |
91 |
0.01 |
|
|
|
| 29 |
A" |
3126 |
2995 |
37.42 |
|
|
|
| 30 |
A" |
3123 |
2993 |
11.75 |
|
|
|
| 31 |
A" |
3122 |
2992 |
23.36 |
|
|
|
| 32 |
A" |
3063 |
2935 |
82.61 |
|
|
|
| 33 |
A" |
1477 |
1416 |
3.58 |
|
|
|
| 34 |
A" |
1475 |
1413 |
0.18 |
|
|
|
| 35 |
A" |
1460 |
1399 |
7.10 |
|
|
|
| 36 |
A" |
1287 |
1233 |
0.25 |
|
|
|
| 37 |
A" |
1264 |
1211 |
0.00 |
|
|
|
| 38 |
A" |
1224 |
1173 |
0.07 |
|
|
|
| 39 |
A" |
1142 |
1095 |
0.43 |
|
|
|
| 40 |
A" |
1128 |
1081 |
0.03 |
|
|
|
| 41 |
A" |
991 |
950 |
3.09 |
|
|
|
| 42 |
A" |
922 |
883 |
0.00 |
|
|
|
| 43 |
A" |
911 |
873 |
0.02 |
|
|
|
| 44 |
A" |
805 |
772 |
0.13 |
|
|
|
| 45 |
A" |
375 |
360 |
0.00 |
|
|
|
| 46 |
A" |
261 |
250 |
0.07 |
|
|
|
| 47 |
A" |
253 |
243 |
0.00 |
|
|
|
| 48 |
A" |
232 |
222 |
0.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36659.9 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 35131.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.252 |
0.083 |
1.080 |
| C2 |
0.252 |
0.083 |
-1.080 |
| C3 |
-0.658 |
0.717 |
0.000 |
| C4 |
0.844 |
-0.859 |
0.000 |
| C5 |
-0.846 |
2.222 |
0.000 |
| C6 |
0.252 |
-2.259 |
0.000 |
| H7 |
0.994 |
0.801 |
1.452 |
| H8 |
-0.239 |
-0.391 |
1.940 |
| H9 |
0.994 |
0.801 |
-1.452 |
| H10 |
-0.239 |
-0.391 |
-1.940 |
| H11 |
-1.640 |
0.226 |
0.000 |
| H12 |
1.940 |
-0.927 |
0.000 |
| H13 |
0.125 |
2.735 |
0.000 |
| H14 |
-0.846 |
-2.227 |
0.000 |
| H15 |
-1.403 |
2.551 |
-0.888 |
| H16 |
-1.403 |
2.551 |
0.888 |
| H17 |
0.571 |
-2.821 |
-0.889 |
| H18 |
0.571 |
-2.821 |
0.889 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
| C1 | | 2.1598 | 1.5481 | 1.5507 | 2.6357 | 2.5791 | 1.0979 | 1.0979 | 2.7348 | 3.0962 | 2.1834 | 2.2439 | 2.8665 | 2.7760 | 3.5644 | 2.9780 | 3.5223 | 2.9270 |
C2 | 2.1598 | | 1.5481 | 1.5507 | 2.6357 | 2.5791 | 2.7348 | 3.0962 | 1.0979 | 1.0979 | 2.1834 | 2.2439 | 2.8665 | 2.7760 | 2.9780 | 3.5644 | 2.9270 | 3.5223 | C3 | 1.5481 | 1.5481 | | 2.1777 | 1.5159 | 3.1128 | 2.2015 | 2.2735 | 2.2015 | 2.2735 | 1.0983 | 3.0745 | 2.1642 | 2.9503 | 2.1695 | 2.1695 | 3.8493 | 3.8493 | C4 | 1.5507 | 1.5507 | 2.1777 | | 3.5143 | 1.5202 | 2.2108 | 2.2709 | 2.2108 | 2.2709 | 2.7114 | 1.0976 | 3.6656 | 2.1744 | 4.1798 | 4.1798 | 2.1704 | 2.1704 | C5 | 2.6357 | 2.6357 | 1.5159 | 3.5143 | | 4.6136 | 2.7418 | 3.3104 | 2.7418 | 3.3104 | 2.1474 | 4.2046 | 1.0989 | 4.4484 | 1.0987 | 1.0987 | 5.3124 | 5.3124 | C6 | 2.5791 | 2.5791 | 3.1128 | 1.5202 | 4.6136 | | 3.4676 | 2.7380 | 3.4676 | 2.7380 | 3.1240 | 2.1501 | 4.9961 | 1.0988 | 5.1643 | 5.1643 | 1.0985 | 1.0985 | H7 | 1.0979 | 2.7348 | 2.2015 | 2.2108 | 2.7418 | 3.4676 | | 1.7828 | 2.9049 | 3.8014 | 3.0629 | 2.4472 | 2.5705 | 3.8293 | 3.7803 | 3.0217 | 4.3328 | 3.6891 | H8 | 1.0979 | 3.0962 | 2.2735 | 2.2709 | 3.3104 | 2.7380 | 1.7828 | | 3.8014 | 3.8805 | 2.4718 | 2.9662 | 3.6971 | 2.7394 | 4.2439 | 3.3345 | 3.8161 | 2.7685 | H9 | 2.7348 | 1.0979 | 2.2015 | 2.2108 | 2.7418 | 3.4676 | 2.9049 | 3.8014 | | 1.7828 | 3.0629 | 2.4472 | 2.5705 | 3.8293 | 3.0217 | 3.7803 | 3.6891 | 4.3328 | H10 | 3.0962 | 1.0979 | 2.2735 | 2.2709 | 3.3104 | 2.7380 | 3.8014 | 3.8805 | 1.7828 | | 2.4718 | 2.9662 | 3.6971 | 2.7394 | 3.3345 | 4.2439 | 2.7685 | 3.8161 | H11 | 2.1834 | 2.1834 | 1.0983 | 2.7114 | 2.1474 | 3.1240 | 3.0629 | 2.4718 | 3.0629 | 2.4718 | | 3.7613 | 3.0677 | 2.5784 | 2.5000 | 2.5000 | 3.8683 | 3.8683 | H12 | 2.2439 | 2.2439 | 3.0745 | 1.0976 | 4.2046 | 2.1501 | 2.4472 | 2.9662 | 2.4472 | 2.9662 | 3.7613 | | 4.0873 | 3.0742 | 4.9054 | 4.9054 | 2.4994 | 2.4994 | H13 | 2.8665 | 2.8665 | 2.1642 | 3.6656 | 1.0989 | 4.9961 | 2.5705 | 3.6971 | 2.5705 | 3.6971 | 3.0677 | 4.0873 | | 5.0561 | 1.7770 | 1.7770 | 5.6439 | 5.6439 | H14 | 2.7760 | 2.7760 | 2.9503 | 2.1744 | 4.4484 | 1.0988 | 3.8293 | 2.7394 | 3.8293 | 2.7394 | 2.5784 | 3.0742 | 5.0561 | | 4.8916 | 4.8916 | 1.7753 | 1.7753 | H15 | 3.5644 | 2.9780 | 2.1695 | 4.1798 | 1.0987 | 5.1643 | 3.7803 | 4.2439 | 3.0217 | 3.3345 | 2.5000 | 4.9054 | 1.7770 | 4.8916 | | 1.7762 | 5.7230 | 5.9925 | H16 | 2.9780 | 3.5644 | 2.1695 | 4.1798 | 1.0987 | 5.1643 | 3.0217 | 3.3345 | 3.7803 | 4.2439 | 2.5000 | 4.9054 | 1.7770 | 4.8916 | 1.7762 | | 5.9925 | 5.7230 | H17 | 3.5223 | 2.9270 | 3.8493 | 2.1704 | 5.3124 | 1.0985 | 4.3328 | 3.8161 | 3.6891 | 2.7685 | 3.8683 | 2.4994 | 5.6439 | 1.7753 | 5.7230 | 5.9925 | | 1.7776 | H18 | 2.9270 | 3.5223 | 3.8493 | 2.1704 | 5.3124 | 1.0985 | 3.6891 | 2.7685 | 4.3328 | 3.8161 | 3.8683 | 2.4994 | 5.6439 | 1.7753 | 5.9925 | 5.7230 | 1.7776 | |
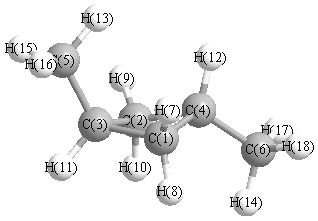 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C3 |
C2 |
88.461 |
|
C1 |
C3 |
C5 |
118.677 |
| C1 |
C3 |
H11 |
110.021 |
|
C1 |
C4 |
C2 |
88.278 |
| C1 |
C4 |
C6 |
114.248 |
|
C1 |
C4 |
H12 |
114.768 |
| C2 |
C3 |
C5 |
118.677 |
|
C2 |
C3 |
H11 |
110.021 |
| C2 |
C4 |
C6 |
114.248 |
|
C2 |
C4 |
H12 |
114.768 |
| C3 |
C1 |
C4 |
89.300 |
|
C3 |
C1 |
H7 |
111.478 |
| C3 |
C1 |
H8 |
117.443 |
|
C3 |
C2 |
C4 |
89.300 |
| C3 |
C2 |
H9 |
111.478 |
|
C3 |
C2 |
H10 |
117.443 |
| C3 |
C5 |
H13 |
110.715 |
|
C3 |
C5 |
H15 |
111.146 |
| C3 |
C5 |
H16 |
111.146 |
|
C4 |
C1 |
H7 |
112.035 |
| C4 |
C1 |
H8 |
117.024 |
|
C4 |
C2 |
H9 |
112.035 |
| C4 |
C2 |
H10 |
117.024 |
|
C4 |
C6 |
H14 |
111.231 |
| C4 |
C6 |
H17 |
110.929 |
|
C4 |
C6 |
H18 |
110.929 |
| C5 |
C3 |
H11 |
109.415 |
|
C6 |
C4 |
H12 |
109.375 |
| H7 |
C1 |
H8 |
108.565 |
|
H9 |
C2 |
H10 |
108.565 |
| H13 |
C5 |
H15 |
107.917 |
|
H13 |
C5 |
H16 |
107.917 |
| H14 |
C6 |
H17 |
107.794 |
|
H14 |
C6 |
H18 |
107.794 |
| H15 |
C5 |
H16 |
107.861 |
|
H17 |
C6 |
H18 |
108.018 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
1.208 |
|
|
|
| 2 |
C |
1.208 |
|
|
|
| 3 |
C |
-0.087 |
|
|
|
| 4 |
C |
0.024 |
|
|
|
| 5 |
C |
0.378 |
|
|
|
| 6 |
C |
0.417 |
|
|
|
| 7 |
H |
-0.370 |
|
|
|
| 8 |
H |
-0.405 |
|
|
|
| 9 |
H |
-0.370 |
|
|
|
| 10 |
H |
-0.405 |
|
|
|
| 11 |
H |
-0.517 |
|
|
|
| 12 |
H |
-0.552 |
|
|
|
| 13 |
H |
-0.122 |
|
|
|
| 14 |
H |
-0.117 |
|
|
|
| 15 |
H |
-0.046 |
|
|
|
| 16 |
H |
-0.046 |
|
|
|
| 17 |
H |
-0.098 |
|
|
|
| 18 |
H |
-0.098 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.015 |
-0.016 |
0.000 |
0.022 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.589 |
0.338 |
0.000 |
| y |
0.338 |
-40.915 |
0.000 |
| z |
0.000 |
0.000 |
-40.581 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.159 |
0.338 |
0.000 |
| y |
0.338 |
-0.830 |
0.000 |
| z |
0.000 |
0.000 |
-0.329 |
|
| Polar |
|---|
| 3z2-r2 | -0.659 |
|---|
| x2-y2 | 1.326 |
|---|
| xy | 0.338 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.916 |
-0.750 |
0.000 |
| y |
-0.750 |
12.127 |
0.000 |
| z |
0.000 |
0.000 |
10.474 |
<r2> (average value of r
2) Å
2
| <r2> |
191.810 |
| (<r2>)1/2 |
13.850 |
