Jump to
S1C2
S1C3
Energy calculated at HF/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -99.790350 |
| Energy at 298.15K | -99.789457 |
| HF Energy | -99.790350 |
| Nuclear repulsion energy | 27.806383 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at HF/aug-cc-pVDZ
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| Li1 |
0.000 |
0.000 |
-2.085 |
| C2 |
0.000 |
0.000 |
-0.138 |
| N3 |
0.000 |
0.000 |
1.012 |
Atom - Atom Distances (Å)
| |
Li1 |
C2 |
N3 |
| Li1 | | 1.9472 | 3.0972 |
C2 | 1.9472 | | 1.1500 | N3 | 3.0972 | 1.1500 | |
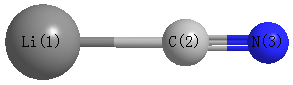 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Li1 |
C2 |
N3 |
180.000 |
|
Li1 |
N3 |
C2 |
0.000 |
| C2 |
Li1 |
N3 |
0.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
Li |
0.195 |
|
|
|
| 2 |
C |
0.302 |
|
|
|
| 3 |
N |
-0.497 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-9.688 |
9.688 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-14.437 |
0.000 |
0.000 |
| y |
0.000 |
-14.437 |
0.000 |
| z |
0.000 |
0.000 |
1.607 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-8.022 |
0.000 |
0.000 |
| y |
0.000 |
-8.022 |
0.000 |
| z |
0.000 |
0.000 |
16.045 |
|
| Polar |
|---|
| 3z2-r2 | 32.090 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.798 |
0.000 |
0.000 |
| y |
0.000 |
2.798 |
0.000 |
| z |
0.000 |
0.000 |
4.243 |
<r2> (average value of r
2) Å
2
| <r2> |
26.004 |
| (<r2>)1/2 |
5.099 |
Jump to
S1C1
S1C3
Energy calculated at HF/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -99.800321 |
| Energy at 298.15K | -99.799506 |
| HF Energy | -99.800321 |
| Nuclear repulsion energy | 28.556410 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at HF/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| Li1 |
-0.006 |
-1.887 |
0.000 |
| C2 |
0.003 |
1.062 |
0.000 |
| N3 |
0.000 |
-0.101 |
0.000 |
Atom - Atom Distances (Å)
| |
Li1 |
C2 |
N3 |
| Li1 | | 2.9491 | 1.7856 |
C2 | 2.9491 | | 1.1634 | N3 | 1.7856 | 1.1634 | |
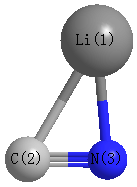 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Li1 |
C2 |
N3 |
0.028 |
|
Li1 |
N3 |
C2 |
179.954 |
| C2 |
Li1 |
N3 |
0.018 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
Li |
0.439 |
|
|
|
| 2 |
C |
-0.355 |
|
|
|
| 3 |
N |
-0.084 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.030 |
-8.981 |
0.000 |
8.981 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-14.392 |
0.044 |
0.000 |
| y |
0.044 |
-1.930 |
0.000 |
| z |
0.000 |
0.000 |
-14.392 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.231 |
0.044 |
0.000 |
| y |
0.044 |
12.462 |
0.000 |
| z |
0.000 |
0.000 |
-6.231 |
|
| Polar |
|---|
| 3z2-r2 | -12.462 |
|---|
| x2-y2 | -12.462 |
|---|
| xy | 0.044 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.829 |
0.004 |
0.000 |
| y |
0.004 |
4.219 |
0.000 |
| z |
0.000 |
0.000 |
2.829 |
<r2> (average value of r
2) Å
2
| <r2> |
23.917 |
| (<r2>)1/2 |
4.890 |
Jump to
S1C1
S1C2
Energy calculated at HF/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -99.800321 |
| Energy at 298.15K | -99.799254 |
| HF Energy | -99.800321 |
| Nuclear repulsion energy | 28.559753 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at HF/aug-cc-pVDZ
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| Li1 |
0.000 |
0.000 |
1.887 |
| C2 |
0.000 |
0.000 |
-1.062 |
| N3 |
0.000 |
0.000 |
0.102 |
Atom - Atom Distances (Å)
| |
Li1 |
C2 |
N3 |
| Li1 | | 2.9484 | 1.7849 |
C2 | 2.9484 | | 1.1634 | N3 | 1.7849 | 1.1634 | |
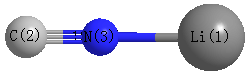 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Li1 |
C2 |
N3 |
0.000 |
|
Li1 |
N3 |
C2 |
180.000 |
| C2 |
Li1 |
N3 |
0.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
Li |
0.439 |
|
|
|
| 2 |
C |
-0.356 |
|
|
|
| 3 |
N |
-0.083 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
8.978 |
8.978 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-14.392 |
0.000 |
0.000 |
| y |
0.000 |
-14.392 |
0.000 |
| z |
0.000 |
0.000 |
-1.939 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.226 |
0.000 |
0.000 |
| y |
0.000 |
-6.226 |
0.000 |
| z |
0.000 |
0.000 |
12.452 |
|
| Polar |
|---|
| 3z2-r2 | 24.905 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.829 |
0.000 |
0.000 |
| y |
0.000 |
2.829 |
0.000 |
| z |
0.000 |
0.000 |
4.219 |
<r2> (average value of r
2) Å
2
| <r2> |
23.910 |
| (<r2>)1/2 |
4.890 |
