Jump to
S1C2
S2C1
S2C2
Energy calculated at MP4/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -39.046594 |
| Energy at 298.15K | -39.046408 |
| HF Energy | -38.927962 |
| Nuclear repulsion energy | 6.074924 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at MP4/aug-cc-pVDZ
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.109 |
| H2 |
0.000 |
1.002 |
-0.327 |
| H3 |
0.000 |
-1.002 |
-0.327 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
H3 |
| C1 | | 1.0928 | 1.0928 |
H2 | 1.0928 | | 2.0037 | H3 | 1.0928 | 2.0037 | |
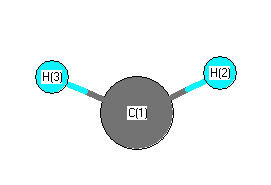 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| H2 |
C1 |
H3 |
132.910 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S2C1
S2C2
Energy calculated at MP4/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -39.035919 |
| Energy at 298.15K | |
| HF Energy | -38.916054 |
| Nuclear repulsion energy | 6.119351 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at MP4/aug-cc-pVDZ
Point Group is D∞h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.000 |
| H2 |
0.000 |
0.000 |
1.080 |
| H3 |
0.000 |
0.000 |
-1.080 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
H3 |
| C1 | | 1.0804 | 1.0804 |
H2 | 1.0804 | | 2.1608 | H3 | 1.0804 | 2.1608 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| H2 |
C1 |
H3 |
180.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
S2C2
Energy calculated at MP4/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -39.027539 |
| Energy at 298.15K | -39.027375 |
| HF Energy | -38.883775 |
| Nuclear repulsion energy | 5.940271 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at MP4/aug-cc-pVDZ
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.180 |
| H2 |
0.000 |
0.868 |
-0.539 |
| H3 |
0.000 |
-0.868 |
-0.539 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
H3 |
| C1 | | 1.1268 | 1.1268 |
H2 | 1.1268 | | 1.7359 | H3 | 1.1268 | 1.7359 | |
More geometry information
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
S2C1
Energy calculated at MP4/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -39.027539 |
| Energy at 298.15K | -39.027375 |
| HF Energy | -38.883775 |
| Nuclear repulsion energy | 5.940271 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4/6-31G*
Geometric Data calculated at MP4/aug-cc-pVDZ
Point Group is C2v
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
