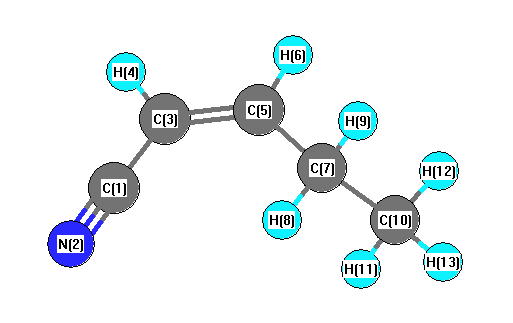
Vibrational Frequencies calculated at HF/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3366 |
3065 |
3.59 |
|
|
|
| 2 |
A |
3314 |
3017 |
11.16 |
|
|
|
| 3 |
A |
3254 |
2963 |
31.49 |
|
|
|
| 4 |
A |
3249 |
2958 |
39.05 |
|
|
|
| 5 |
A |
3229 |
2941 |
12.91 |
|
|
|
| 6 |
A |
3173 |
2890 |
15.76 |
|
|
|
| 7 |
A |
3169 |
2885 |
37.94 |
|
|
|
| 8 |
A |
2567 |
2338 |
49.79 |
|
|
|
| 9 |
A |
1837 |
1673 |
26.39 |
|
|
|
| 10 |
A |
1603 |
1459 |
6.82 |
|
|
|
| 11 |
A |
1593 |
1451 |
5.27 |
|
|
|
| 12 |
A |
1584 |
1442 |
3.85 |
|
|
|
| 13 |
A |
1530 |
1393 |
1.28 |
|
|
|
| 14 |
A |
1519 |
1383 |
1.50 |
|
|
|
| 15 |
A |
1444 |
1315 |
5.84 |
|
|
|
| 16 |
A |
1391 |
1266 |
0.20 |
|
|
|
| 17 |
A |
1344 |
1224 |
1.36 |
|
|
|
| 18 |
A |
1227 |
1117 |
0.47 |
|
|
|
| 19 |
A |
1184 |
1078 |
3.84 |
|
|
|
| 20 |
A |
1100 |
1002 |
1.81 |
|
|
|
| 21 |
A |
1084 |
987 |
2.42 |
|
|
|
| 22 |
A |
1016 |
925 |
6.04 |
|
|
|
| 23 |
A |
910 |
829 |
3.24 |
|
|
|
| 24 |
A |
859 |
782 |
11.03 |
|
|
|
| 25 |
A |
827 |
753 |
36.74 |
|
|
|
| 26 |
A |
716 |
652 |
1.70 |
|
|
|
| 27 |
A |
616 |
561 |
0.59 |
|
|
|
| 28 |
A |
433 |
395 |
0.80 |
|
|
|
| 29 |
A |
398 |
363 |
0.02 |
|
|
|
| 30 |
A |
263 |
239 |
0.83 |
|
|
|
| 31 |
A |
234 |
213 |
2.92 |
|
|
|
| 32 |
A |
160 |
146 |
5.81 |
|
|
|
| 33 |
A |
58 |
53 |
1.36 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25124.7 cm
-1
Scaled (by 0.9106) Zero Point Vibrational Energy (zpe) 22878.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.550 |
|
|
|
| 2 |
N |
-0.292 |
|
|
|
| 3 |
C |
0.948 |
|
|
|
| 4 |
H |
-0.368 |
|
|
|
| 5 |
C |
0.595 |
|
|
|
| 6 |
H |
-0.448 |
|
|
|
| 7 |
C |
0.073 |
|
|
|
| 8 |
H |
-0.083 |
|
|
|
| 9 |
H |
0.033 |
|
|
|
| 10 |
C |
0.301 |
|
|
|
| 11 |
H |
-0.067 |
|
|
|
| 12 |
H |
-0.062 |
|
|
|
| 13 |
H |
-0.080 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
3.767 |
2.712 |
0.081 |
4.642 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-44.089 |
-6.018 |
1.160 |
| y |
-6.018 |
-36.705 |
-0.739 |
| z |
1.160 |
-0.739 |
-37.900 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.786 |
-6.018 |
1.160 |
| y |
-6.018 |
4.289 |
-0.739 |
| z |
1.160 |
-0.739 |
2.497 |
|
| Polar |
|---|
| 3z2-r2 | 4.993 |
|---|
| x2-y2 | -7.383 |
|---|
| xy | -6.018 |
|---|
| xz | 1.160 |
|---|
| yz | -0.739 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
201.620 |
| (<r2>)1/2 |
14.199 |