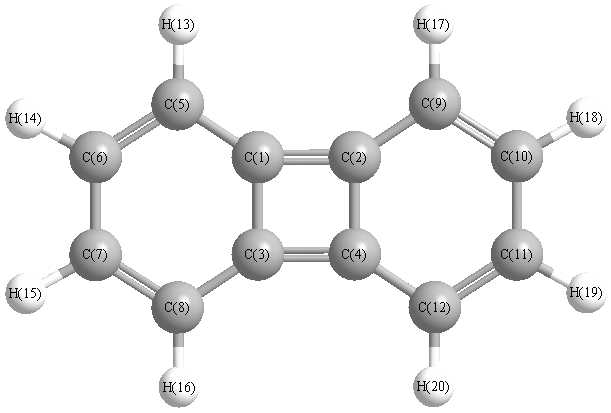
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3230 |
3095 |
0.00 |
|
|
|
| 2 |
Ag |
3214 |
3080 |
0.00 |
|
|
|
| 3 |
Ag |
1740 |
1668 |
0.00 |
|
|
|
| 4 |
Ag |
1541 |
1477 |
0.00 |
|
|
|
| 5 |
Ag |
1437 |
1377 |
0.00 |
|
|
|
| 6 |
Ag |
1191 |
1141 |
0.00 |
|
|
|
| 7 |
Ag |
1159 |
1111 |
0.00 |
|
|
|
| 8 |
Ag |
1025 |
983 |
0.00 |
|
|
|
| 9 |
Ag |
786 |
753 |
0.00 |
|
|
|
| 10 |
Ag |
403 |
386 |
0.00 |
|
|
|
| 11 |
Au |
985 |
943 |
0.00 |
|
|
|
| 12 |
Au |
887 |
850 |
0.00 |
|
|
|
| 13 |
Au |
801 |
767 |
0.00 |
|
|
|
| 14 |
Au |
590 |
565 |
0.00 |
|
|
|
| 15 |
Au |
146 |
140 |
0.00 |
|
|
|
| 16 |
B1g |
985 |
944 |
0.00 |
|
|
|
| 17 |
B1g |
877 |
840 |
0.00 |
|
|
|
| 18 |
B1g |
723 |
693 |
0.00 |
|
|
|
| 19 |
B1g |
462 |
442 |
0.00 |
|
|
|
| 20 |
B1u |
3229 |
3095 |
31.72 |
|
|
|
| 21 |
B1u |
3213 |
3079 |
10.36 |
|
|
|
| 22 |
B1u |
1660 |
1591 |
0.03 |
|
|
|
| 23 |
B1u |
1465 |
1404 |
33.24 |
|
|
|
| 24 |
B1u |
1337 |
1281 |
34.09 |
|
|
|
| 25 |
B1u |
1178 |
1129 |
9.22 |
|
|
|
| 26 |
B1u |
1053 |
1009 |
0.02 |
|
|
|
| 27 |
B1u |
1008 |
966 |
26.65 |
|
|
|
| 28 |
B1u |
623 |
597 |
3.85 |
|
|
|
| 29 |
B2g |
931 |
892 |
0.00 |
|
|
|
| 30 |
B2g |
756 |
724 |
0.00 |
|
|
|
| 31 |
B2g |
428 |
410 |
0.00 |
|
|
|
| 32 |
B2g |
318 |
305 |
0.00 |
|
|
|
| 33 |
B2u |
3223 |
3089 |
37.41 |
|
|
|
| 34 |
B2u |
3202 |
3068 |
0.12 |
|
|
|
| 35 |
B2u |
1663 |
1594 |
1.61 |
|
|
|
| 36 |
B2u |
1482 |
1421 |
4.70 |
|
|
|
| 37 |
B2u |
1291 |
1237 |
5.01 |
|
|
|
| 38 |
B2u |
1152 |
1104 |
13.94 |
|
|
|
| 39 |
B2u |
1074 |
1030 |
0.41 |
|
|
|
| 40 |
B2u |
735 |
705 |
0.37 |
|
|
|
| 41 |
B2u |
214 |
205 |
0.80 |
|
|
|
| 42 |
B3g |
3223 |
3088 |
0.00 |
|
|
|
| 43 |
B3g |
3201 |
3068 |
0.00 |
|
|
|
| 44 |
B3g |
1670 |
1600 |
0.00 |
|
|
|
| 45 |
B3g |
1483 |
1422 |
0.00 |
|
|
|
| 46 |
B3g |
1303 |
1248 |
0.00 |
|
|
|
| 47 |
B3g |
1117 |
1071 |
0.00 |
|
|
|
| 48 |
B3g |
997 |
955 |
0.00 |
|
|
|
| 49 |
B3g |
608 |
583 |
0.00 |
|
|
|
| 50 |
B3g |
568 |
544 |
0.00 |
|
|
|
| 51 |
B3u |
929 |
891 |
6.67 |
|
|
|
| 52 |
B3u |
745 |
714 |
146.88 |
|
|
|
| 53 |
B3u |
376 |
361 |
3.37 |
|
|
|
| 54 |
B3u |
107 |
102 |
3.11 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 34870.9 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 33416.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.186 |
|
|
|
| 2 |
C |
0.186 |
|
|
|
| 3 |
C |
0.186 |
|
|
|
| 4 |
C |
0.186 |
|
|
|
| 5 |
C |
0.716 |
|
|
|
| 6 |
C |
0.238 |
|
|
|
| 7 |
C |
0.238 |
|
|
|
| 8 |
C |
0.716 |
|
|
|
| 9 |
C |
0.716 |
|
|
|
| 10 |
C |
0.238 |
|
|
|
| 11 |
C |
0.238 |
|
|
|
| 12 |
C |
0.716 |
|
|
|
| 13 |
H |
-0.537 |
|
|
|
| 14 |
H |
-0.602 |
|
|
|
| 15 |
H |
-0.602 |
|
|
|
| 16 |
H |
-0.537 |
|
|
|
| 17 |
H |
-0.537 |
|
|
|
| 18 |
H |
-0.602 |
|
|
|
| 19 |
H |
-0.602 |
|
|
|
| 20 |
H |
-0.537 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-75.170 |
0.000 |
0.000 |
| y |
0.000 |
-60.228 |
0.000 |
| z |
0.000 |
0.000 |
-60.504 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-14.804 |
0.000 |
0.000 |
| y |
0.000 |
7.609 |
0.000 |
| z |
0.000 |
0.000 |
7.195 |
|
| Polar |
|---|
| 3z2-r2 | 14.389 |
|---|
| x2-y2 | -14.942 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.105 |
0.000 |
0.000 |
| y |
0.000 |
20.828 |
0.000 |
| z |
0.000 |
0.000 |
33.829 |
<r2> (average value of r
2) Å
2
| <r2> |
561.550 |
| (<r2>)1/2 |
23.697 |