Jump to
S1C2
S2C1
S2C2
Energy calculated at HF/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -130.696432 |
| Energy at 298.15K | -130.695808 |
| HF Energy | -130.696432 |
| Nuclear repulsion energy | 47.100362 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3475 |
3165 |
18.70 |
|
|
|
| 2 |
A' |
1663 |
1514 |
149.90 |
|
|
|
| 3 |
A' |
1203 |
1096 |
2.96 |
|
|
|
| 4 |
A' |
646 |
588 |
19.05 |
|
|
|
| 5 |
A' |
376 |
342 |
0.25 |
|
|
|
| 6 |
A" |
401 |
366 |
0.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 3882.3 cm
-1
Scaled (by 0.9106) Zero Point Vibrational Energy (zpe) 3535.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.249 |
-1.212 |
0.000 |
| C2 |
0.000 |
0.092 |
0.000 |
| N3 |
-0.361 |
1.237 |
0.000 |
| H4 |
1.033 |
-1.941 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
| C1 | | 1.3278 | 2.5235 | 1.0706 |
C2 | 1.3278 | | 1.2001 | 2.2806 | N3 | 2.5235 | 1.2001 | | 3.4700 | H4 | 1.0706 | 2.2806 | 3.4700 | |
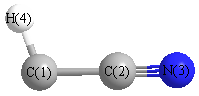 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
173.305 |
|
C2 |
C1 |
H4 |
143.724 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S2C1
S2C2
Energy calculated at HF/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -130.693916 |
| Energy at 298.15K | |
| HF Energy | -130.693916 |
| Nuclear repulsion energy | 47.200309 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Σ |
3546 |
3229 |
59.88 |
|
|
|
| 2 |
Σ |
1591 |
1448 |
133.94 |
|
|
|
| 3 |
Σ |
1251 |
1139 |
0.21 |
|
|
|
| 4 |
Π |
400 |
365 |
0.21 |
|
|
|
| 4 |
Π |
400 |
365 |
0.21 |
|
|
|
| 5 |
Π |
470i |
428i |
44.42 |
|
|
|
| 5 |
Π |
470i |
428i |
44.42 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 3124.3 cm
-1
Scaled (by 0.9106) Zero Point Vibrational Energy (zpe) 2845.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/aug-cc-pVDZ
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-1.214 |
| C2 |
0.000 |
0.000 |
0.080 |
| N3 |
0.000 |
0.000 |
1.297 |
| H4 |
0.000 |
0.000 |
-2.278 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
| C1 | | 1.2934 | 2.5110 | 1.0640 |
C2 | 1.2934 | | 1.2176 | 2.3574 | N3 | 2.5110 | 1.2176 | | 3.5750 | H4 | 1.0640 | 2.3574 | 3.5750 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
180.000 |
|
C2 |
C1 |
H4 |
180.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.724 |
|
|
|
| 2 |
C |
-0.068 |
|
|
|
| 3 |
N |
-0.390 |
|
|
|
| 4 |
H |
-0.266 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-3.213 |
3.213 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-17.591 |
0.000 |
0.000 |
| y |
0.000 |
-17.591 |
0.000 |
| z |
0.000 |
0.000 |
-15.712 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.940 |
0.000 |
0.000 |
| y |
0.000 |
-0.940 |
0.000 |
| z |
0.000 |
0.000 |
1.880 |
|
| Polar |
|---|
| 3z2-r2 | 3.760 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.938 |
0.000 |
0.000 |
| y |
0.000 |
2.938 |
0.000 |
| z |
0.000 |
0.000 |
6.333 |
<r2> (average value of r
2) Å
2
| <r2> |
36.442 |
| (<r2>)1/2 |
6.037 |
Jump to
S1C1
S1C2
S2C2
Energy calculated at HF/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -130.638163 |
| Energy at 298.15K | -130.637686 |
| HF Energy | -130.638163 |
| Nuclear repulsion energy | 47.178611 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3231 |
2942 |
4.93 |
|
|
|
| 2 |
A' |
2343 |
2133 |
39.70 |
|
|
|
| 3 |
A' |
1134 |
1033 |
67.16 |
|
|
|
| 4 |
A' |
1025 |
934 |
24.71 |
|
|
|
| 5 |
A' |
478 |
435 |
26.65 |
|
|
|
| 6 |
A" |
341 |
310 |
15.57 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4276.1 cm
-1
Scaled (by 0.9106) Zero Point Vibrational Energy (zpe) 3893.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.245 |
-1.277 |
0.000 |
| C2 |
0.000 |
0.112 |
0.000 |
| N3 |
0.108 |
1.256 |
0.000 |
| H4 |
0.718 |
-1.799 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
| C1 | | 1.4105 | 2.5576 | 1.0957 |
C2 | 1.4105 | | 1.1491 | 2.0414 | N3 | 2.5576 | 1.1491 | | 3.1154 | H4 | 1.0957 | 2.0414 | 3.1154 | |
More geometry information
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.200 |
|
|
|
| 2 |
C |
0.030 |
|
|
|
| 3 |
N |
-0.254 |
|
|
|
| 4 |
H |
0.025 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.378 |
-2.176 |
0.000 |
2.576 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-18.714 |
-3.420 |
0.000 |
| y |
-3.420 |
-20.415 |
0.000 |
| z |
0.000 |
0.000 |
-16.253 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.380 |
-3.420 |
0.000 |
| y |
-3.420 |
-2.931 |
0.000 |
| z |
0.000 |
0.000 |
3.311 |
|
| Polar |
|---|
| 3z2-r2 | 6.622 |
|---|
| x2-y2 | 1.701 |
|---|
| xy | -3.420 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.207 |
0.282 |
0.000 |
| y |
0.282 |
6.274 |
0.000 |
| z |
0.000 |
0.000 |
3.292 |
<r2> (average value of r
2) Å
2
| <r2> |
36.629 |
| (<r2>)1/2 |
6.052 |
Jump to
S1C1
S1C2
S2C1
Energy calculated at HF/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -130.638163 |
| Energy at 298.15K | -130.637686 |
| HF Energy | -130.638163 |
| Nuclear repulsion energy | 47.178611 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/aug-cc-pVDZ
Geometric Data calculated at HF/aug-cc-pVDZ
Point Group is Cs
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
