Jump to
S1C2
S2C1
S2C2
Energy calculated at mPW1PW91/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -131.400654 |
| Energy at 298.15K | -131.399972 |
| HF Energy | -131.400654 |
| Nuclear repulsion energy | 47.220976 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3398 |
3257 |
35.15 |
|
|
|
| 2 |
A' |
1813 |
1737 |
42.28 |
|
|
|
| 3 |
A' |
1232 |
1181 |
8.06 |
|
|
|
| 4 |
A' |
485 |
465 |
26.83 |
|
|
|
| 5 |
A' |
390 |
374 |
13.46 |
|
|
|
| 6 |
A" |
431 |
413 |
0.58 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 3874.7 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 3713.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.080 |
-1.224 |
0.000 |
| C2 |
0.000 |
0.089 |
0.000 |
| N3 |
-0.162 |
1.278 |
0.000 |
| H4 |
0.654 |
-2.134 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
| C1 | | 1.3149 | 2.5137 | 1.0759 |
C2 | 1.3149 | | 1.2005 | 2.3166 | N3 | 2.5137 | 1.2005 | | 3.5081 | H4 | 1.0759 | 2.3166 | 3.5081 | |
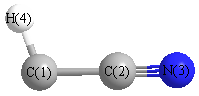 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
175.734 |
|
C2 |
C1 |
H4 |
151.222 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S2C1
S2C2
Energy calculated at mPW1PW91/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -131.399838 |
| Energy at 298.15K | |
| HF Energy | -131.399838 |
| Nuclear repulsion energy | 47.286686 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Σ |
3459 |
3315 |
66.33 |
|
|
|
| 2 |
Σ |
1742 |
1669 |
45.13 |
|
|
|
| 3 |
Σ |
1275 |
1222 |
4.70 |
|
|
|
| 4 |
Π |
433 |
415 |
0.04 |
|
|
|
| 4 |
Π |
433 |
415 |
0.04 |
|
|
|
| 5 |
Π |
349i |
334i |
45.40 |
|
|
|
| 5 |
Π |
349i |
334i |
45.40 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 3322.0 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 3183.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-1.212 |
| C2 |
0.000 |
0.000 |
0.084 |
| N3 |
0.000 |
0.000 |
1.293 |
| H4 |
0.000 |
0.000 |
-2.283 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
| C1 | | 1.2958 | 2.5056 | 1.0706 |
C2 | 1.2958 | | 1.2098 | 2.3664 | N3 | 2.5056 | 1.2098 | | 3.5762 | H4 | 1.0706 | 2.3664 | 3.5762 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
180.000 |
|
C2 |
C1 |
H4 |
180.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.651 |
|
|
|
| 2 |
C |
0.089 |
|
|
|
| 3 |
N |
-0.385 |
|
|
|
| 4 |
H |
-0.354 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-3.336 |
3.336 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-17.432 |
0.000 |
0.000 |
| y |
0.000 |
-17.432 |
0.000 |
| z |
0.000 |
0.000 |
-15.584 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.924 |
0.000 |
0.000 |
| y |
0.000 |
-0.924 |
0.000 |
| z |
0.000 |
0.000 |
1.848 |
|
| Polar |
|---|
| 3z2-r2 | 3.696 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.035 |
0.000 |
0.000 |
| y |
0.000 |
3.035 |
0.000 |
| z |
0.000 |
0.000 |
7.106 |
<r2> (average value of r
2) Å
2
| <r2> |
36.283 |
| (<r2>)1/2 |
6.024 |
Jump to
S1C1
S1C2
S2C2
Energy calculated at mPW1PW91/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -131.370223 |
| Energy at 298.15K | -131.369653 |
| HF Energy | -131.370223 |
| Nuclear repulsion energy | 46.910555 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3091 |
2962 |
4.27 |
|
|
|
| 2 |
A' |
2120 |
2031 |
24.59 |
|
|
|
| 3 |
A' |
1093 |
1048 |
39.78 |
|
|
|
| 4 |
A' |
953 |
914 |
42.43 |
|
|
|
| 5 |
A' |
450 |
431 |
23.60 |
|
|
|
| 6 |
A" |
298 |
285 |
10.22 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4002.4 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 3835.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.068 |
-1.285 |
0.000 |
| C2 |
0.000 |
0.093 |
0.000 |
| N3 |
-0.218 |
1.254 |
0.000 |
| H4 |
1.121 |
-1.623 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
| C1 | | 1.3797 | 2.5549 | 1.1062 |
C2 | 1.3797 | | 1.1811 | 2.0493 | N3 | 2.5549 | 1.1811 | | 3.1728 | H4 | 1.1062 | 2.0493 | 3.1728 | |
More geometry information
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.074 |
|
|
|
| 2 |
C |
0.254 |
|
|
|
| 3 |
N |
-0.267 |
|
|
|
| 4 |
H |
-0.061 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.719 |
-1.467 |
0.000 |
2.260 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-17.169 |
-2.463 |
0.000 |
| y |
-2.463 |
-21.146 |
0.000 |
| z |
0.000 |
0.000 |
-16.083 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.446 |
-2.463 |
0.000 |
| y |
-2.463 |
-4.520 |
0.000 |
| z |
0.000 |
0.000 |
3.074 |
|
| Polar |
|---|
| 3z2-r2 | 6.148 |
|---|
| x2-y2 | 3.978 |
|---|
| xy | -2.463 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.476 |
-0.509 |
0.000 |
| y |
-0.509 |
7.068 |
0.000 |
| z |
0.000 |
0.000 |
3.310 |
<r2> (average value of r
2) Å
2
| <r2> |
36.540 |
| (<r2>)1/2 |
6.045 |
Jump to
S1C1
S1C2
S2C1
Energy calculated at mPW1PW91/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -131.370223 |
| Energy at 298.15K | -131.369653 |
| HF Energy | -131.370223 |
| Nuclear repulsion energy | 46.910555 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Point Group is Cs
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
