Jump to
S1C2
S2C1
S2C2
Energy calculated at MP4/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -131.079337 |
| Energy at 298.15K | -131.078858 |
| HF Energy | -130.694822 |
| Nuclear repulsion energy | 46.910527 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3359 |
3282 |
|
|
|
|
| 2 |
A' |
2033 |
1986 |
|
|
|
|
| 3 |
A' |
1130 |
1104 |
|
|
|
|
| 4 |
A' |
745 |
727 |
|
|
|
|
| 5 |
A' |
426 |
416 |
|
|
|
|
| 6 |
A" |
486 |
475 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4089.6 cm
-1
Scaled (by 0.9769) Zero Point Vibrational Energy (zpe) 3995.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP4/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.120 |
-1.255 |
0.000 |
| C2 |
0.000 |
0.110 |
0.000 |
| N3 |
0.028 |
1.286 |
0.000 |
| H4 |
0.525 |
-2.130 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
| C1 | | 1.3700 | 2.5448 | 1.0875 |
C2 | 1.3700 | | 1.1760 | 2.3011 | N3 | 2.5448 | 1.1760 | | 3.4521 | H4 | 1.0875 | 2.3011 | 3.4521 | |
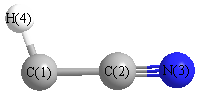 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
176.331 |
|
C2 |
C1 |
H4 |
138.613 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S2C1
S2C2
Energy calculated at MP4/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -131.075501 |
| Energy at 298.15K | |
| HF Energy | -130.693233 |
| Nuclear repulsion energy | 47.105929 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Σ |
3474 |
3394 |
|
|
|
|
| 2 |
Σ |
1639 |
1601 |
|
|
|
|
| 3 |
Σ |
1264 |
1235 |
|
|
|
|
| 4 |
Π |
478 |
467 |
|
|
|
|
| 4 |
Π |
478 |
467 |
|
|
|
|
| 5 |
Π |
521i |
509i |
|
|
|
|
| 5 |
Π |
521i |
509i |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 3145.4 cm
-1
Scaled (by 0.9769) Zero Point Vibrational Energy (zpe) 3072.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP4/aug-cc-pVDZ
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-1.222 |
| C2 |
0.000 |
0.000 |
0.094 |
| N3 |
0.000 |
0.000 |
1.295 |
| H4 |
0.000 |
0.000 |
-2.300 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
| C1 | | 1.3163 | 2.5177 | 1.0773 |
C2 | 1.3163 | | 1.2013 | 2.3936 | N3 | 2.5177 | 1.2013 | | 3.5950 | H4 | 1.0773 | 2.3936 | 3.5950 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
180.000 |
|
C2 |
C1 |
H4 |
180.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at MP4/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.599 |
|
|
|
| 2 |
C |
-0.047 |
|
|
|
| 3 |
N |
-0.358 |
|
|
|
| 4 |
H |
-0.193 |
|
|
|
Electric dipole moments
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
36.656 |
| (<r2>)1/2 |
6.054 |
Jump to
S1C1
S1C2
S2C2
Energy calculated at MP4/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -131.075159 |
| Energy at 298.15K | -131.074535 |
| HF Energy | -130.631406 |
| Nuclear repulsion energy | 45.958558 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3040 |
2970 |
|
|
|
|
| 2 |
A' |
2089 |
2041 |
|
|
|
|
| 3 |
A' |
1058 |
1034 |
|
|
|
|
| 4 |
A' |
973 |
951 |
|
|
|
|
| 5 |
A' |
390 |
381 |
|
|
|
|
| 6 |
A" |
310 |
302 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 3929.9 cm
-1
Scaled (by 0.9769) Zero Point Vibrational Energy (zpe) 3839.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP4/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.143 |
-1.314 |
0.000 |
| C2 |
0.000 |
0.098 |
0.000 |
| N3 |
-0.299 |
1.263 |
0.000 |
| H4 |
1.236 |
-1.543 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
| C1 | | 1.4184 | 2.6138 | 1.1172 |
C2 | 1.4184 | | 1.2028 | 2.0541 | N3 | 2.6138 | 1.2028 | | 3.1980 | H4 | 1.1172 | 2.0541 | 3.1980 | |
More geometry information
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
S2C1
Energy calculated at MP4/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -131.075159 |
| Energy at 298.15K | -131.074535 |
| HF Energy | -130.631406 |
| Nuclear repulsion energy | 45.958558 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4/6-31G*
Geometric Data calculated at MP4/aug-cc-pVDZ
Point Group is Cs
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
