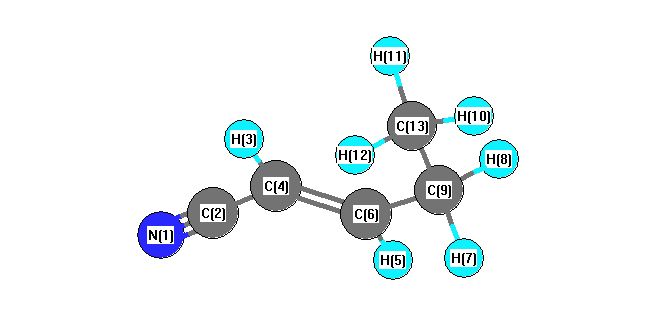
Vibrational Frequencies calculated at HF/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3376 |
3074 |
3.11 |
|
|
|
| 2 |
A' |
3324 |
3027 |
6.35 |
|
|
|
| 3 |
A' |
3247 |
2957 |
40.89 |
|
|
|
| 4 |
A' |
3173 |
2889 |
17.15 |
|
|
|
| 5 |
A' |
3163 |
2880 |
28.91 |
|
|
|
| 6 |
A' |
2569 |
2339 |
67.59 |
|
|
|
| 7 |
A' |
1846 |
1681 |
33.16 |
|
|
|
| 8 |
A' |
1604 |
1460 |
2.81 |
|
|
|
| 9 |
A' |
1575 |
1434 |
5.58 |
|
|
|
| 10 |
A' |
1533 |
1396 |
5.68 |
|
|
|
| 11 |
A' |
1488 |
1355 |
11.38 |
|
|
|
| 12 |
A' |
1434 |
1306 |
4.35 |
|
|
|
| 13 |
A' |
1398 |
1273 |
2.33 |
|
|
|
| 14 |
A' |
1190 |
1084 |
1.69 |
|
|
|
| 15 |
A' |
1124 |
1023 |
3.68 |
|
|
|
| 16 |
A' |
1045 |
951 |
2.85 |
|
|
|
| 17 |
A' |
948 |
863 |
2.40 |
|
|
|
| 18 |
A' |
615 |
560 |
0.58 |
|
|
|
| 19 |
A' |
550 |
500 |
0.36 |
|
|
|
| 20 |
A' |
276 |
252 |
2.07 |
|
|
|
| 21 |
A' |
161 |
147 |
5.46 |
|
|
|
| 22 |
A" |
3243 |
2953 |
42.59 |
|
|
|
| 23 |
A" |
3189 |
2904 |
10.53 |
|
|
|
| 24 |
A" |
1597 |
1454 |
7.18 |
|
|
|
| 25 |
A" |
1390 |
1265 |
0.58 |
|
|
|
| 26 |
A" |
1203 |
1096 |
3.53 |
|
|
|
| 27 |
A" |
1089 |
992 |
35.96 |
|
|
|
| 28 |
A" |
930 |
847 |
11.08 |
|
|
|
| 29 |
A" |
796 |
725 |
0.02 |
|
|
|
| 30 |
A" |
536 |
488 |
6.17 |
|
|
|
| 31 |
A" |
301 |
274 |
0.02 |
|
|
|
| 32 |
A" |
193 |
176 |
0.01 |
|
|
|
| 33 |
A" |
132 |
120 |
4.06 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25117.3 cm
-1
Scaled (by 0.9106) Zero Point Vibrational Energy (zpe) 22871.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.153 |
|
|
|
| 2 |
C |
-1.228 |
|
|
|
| 3 |
H |
-0.595 |
|
|
|
| 4 |
C |
1.635 |
|
|
|
| 5 |
H |
-0.499 |
|
|
|
| 6 |
C |
0.179 |
|
|
|
| 7 |
H |
0.064 |
|
|
|
| 8 |
H |
0.064 |
|
|
|
| 9 |
C |
0.077 |
|
|
|
| 10 |
H |
-0.139 |
|
|
|
| 11 |
H |
-0.003 |
|
|
|
| 12 |
H |
-0.003 |
|
|
|
| 13 |
C |
0.602 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.426 |
-4.722 |
0.000 |
5.309 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.312 |
7.972 |
0.000 |
| y |
7.972 |
-46.674 |
0.000 |
| z |
0.000 |
0.000 |
-38.022 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.036 |
7.972 |
0.000 |
| y |
7.972 |
-8.007 |
0.000 |
| z |
0.000 |
0.000 |
4.972 |
|
| Polar |
|---|
| 3z2-r2 | 9.943 |
|---|
| x2-y2 | 7.362 |
|---|
| xy | 7.972 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
227.776 |
| (<r2>)1/2 |
15.092 |