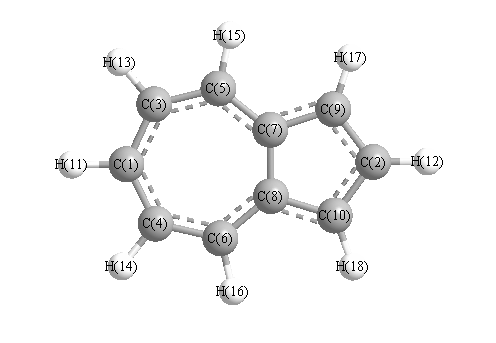
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3261 |
3125 |
7.57 |
|
|
|
| 2 |
A1 |
3232 |
3097 |
5.77 |
|
|
|
| 3 |
A1 |
3209 |
3075 |
16.52 |
|
|
|
| 4 |
A1 |
3182 |
3049 |
5.79 |
|
|
|
| 5 |
A1 |
3172 |
3040 |
0.10 |
|
|
|
| 6 |
A1 |
1671 |
1601 |
72.51 |
|
|
|
| 7 |
A1 |
1595 |
1529 |
7.39 |
|
|
|
| 8 |
A1 |
1496 |
1434 |
40.24 |
|
|
|
| 9 |
A1 |
1435 |
1375 |
79.45 |
|
|
|
| 10 |
A1 |
1311 |
1256 |
1.26 |
|
|
|
| 11 |
A1 |
1244 |
1192 |
0.12 |
|
|
|
| 12 |
A1 |
1083 |
1038 |
8.24 |
|
|
|
| 13 |
A1 |
973 |
933 |
3.37 |
|
|
|
| 14 |
A1 |
906 |
869 |
2.62 |
|
|
|
| 15 |
A1 |
833 |
798 |
6.03 |
|
|
|
| 16 |
A1 |
690 |
661 |
0.91 |
|
|
|
| 17 |
A1 |
408 |
391 |
1.57 |
|
|
|
| 18 |
A2 |
999 |
957 |
0.00 |
|
|
|
| 19 |
A2 |
883 |
846 |
0.00 |
|
|
|
| 20 |
A2 |
803 |
769 |
0.00 |
|
|
|
| 21 |
A2 |
736 |
705 |
0.00 |
|
|
|
| 22 |
A2 |
431 |
413 |
0.00 |
|
|
|
| 23 |
A2 |
163 |
156 |
0.00 |
|
|
|
| 24 |
B1 |
1020 |
977 |
0.00 |
|
|
|
| 25 |
B1 |
980 |
940 |
5.57 |
|
|
|
| 26 |
B1 |
927 |
889 |
0.05 |
|
|
|
| 27 |
B1 |
785 |
752 |
122.22 |
|
|
|
| 28 |
B1 |
741 |
710 |
1.62 |
|
|
|
| 29 |
B1 |
615 |
589 |
1.00 |
|
|
|
| 30 |
B1 |
575 |
551 |
11.77 |
|
|
|
| 31 |
B1 |
323 |
309 |
12.04 |
|
|
|
| 32 |
B1 |
170 |
163 |
2.13 |
|
|
|
| 33 |
B2 |
3252 |
3116 |
9.20 |
|
|
|
| 34 |
B2 |
3201 |
3067 |
25.68 |
|
|
|
| 35 |
B2 |
3174 |
3042 |
10.17 |
|
|
|
| 36 |
B2 |
1665 |
1595 |
6.64 |
|
|
|
| 37 |
B2 |
1561 |
1496 |
7.74 |
|
|
|
| 38 |
B2 |
1485 |
1423 |
7.23 |
|
|
|
| 39 |
B2 |
1425 |
1366 |
0.69 |
|
|
|
| 40 |
B2 |
1337 |
1281 |
0.13 |
|
|
|
| 41 |
B2 |
1315 |
1260 |
4.11 |
|
|
|
| 42 |
B2 |
1239 |
1187 |
8.82 |
|
|
|
| 43 |
B2 |
1186 |
1136 |
0.05 |
|
|
|
| 44 |
B2 |
1067 |
1023 |
0.06 |
|
|
|
| 45 |
B2 |
1029 |
986 |
13.26 |
|
|
|
| 46 |
B2 |
745 |
714 |
0.25 |
|
|
|
| 47 |
B2 |
497 |
476 |
1.79 |
|
|
|
| 48 |
B2 |
335 |
321 |
0.92 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 32178.9 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 30837.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.363 |
|
|
|
| 2 |
C |
0.065 |
|
|
|
| 3 |
C |
0.358 |
|
|
|
| 4 |
C |
0.358 |
|
|
|
| 5 |
C |
0.472 |
|
|
|
| 6 |
C |
0.472 |
|
|
|
| 7 |
C |
1.485 |
|
|
|
| 8 |
C |
1.485 |
|
|
|
| 9 |
C |
0.124 |
|
|
|
| 10 |
C |
0.124 |
|
|
|
| 11 |
H |
-0.643 |
|
|
|
| 12 |
H |
-0.462 |
|
|
|
| 13 |
H |
-0.688 |
|
|
|
| 14 |
H |
-0.688 |
|
|
|
| 15 |
H |
-0.722 |
|
|
|
| 16 |
H |
-0.722 |
|
|
|
| 17 |
H |
-0.690 |
|
|
|
| 18 |
H |
-0.690 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-1.077 |
1.077 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-64.375 |
0.000 |
0.000 |
| y |
0.000 |
-50.904 |
0.000 |
| z |
0.000 |
0.000 |
-51.831 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-13.007 |
0.000 |
0.000 |
| y |
0.000 |
7.198 |
0.000 |
| z |
0.000 |
0.000 |
5.809 |
|
| Polar |
|---|
| 3z2-r2 | 11.618 |
|---|
| x2-y2 | -13.470 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.907 |
0.000 |
0.000 |
| y |
0.000 |
19.420 |
0.000 |
| z |
0.000 |
0.000 |
28.932 |
<r2> (average value of r
2) Å
2
| <r2> |
366.090 |
| (<r2>)1/2 |
19.133 |