return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3C(OH)=NH (Ethaninidic acid)

using model chemistry: MP4/aug-cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP4/aug-cc-pVDZ
 hartrees
Energy at 0K-208.709947
Energy at 298.15K-208.716258
HF Energy-207.990460
Nuclear repulsion energy120.322307
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3671 3586        
2 A' 3485 3405        
3 A' 3173 3100        
4 A' 3075 3004        
5 A' 1749 1708        
6 A' 1533 1498        
7 A' 1481 1447        
8 A' 1426 1393        
9 A' 1289 1260        
10 A' 1133 1107        
11 A' 1037 1013        
12 A' 881 860        
13 A' 548 535        
14 A' 422 413        
15 A" 3154 3082        
16 A" 1517 1482        
17 A" 1088 1063        
18 A" 863 843        
19 A" 631 616        
20 A" 515 503        
21 A" 124 121        

Unscaled Zero Point Vibrational Energy (zpe) 16396.3 cm-1
Scaled (by 0.9769) Zero Point Vibrational Energy (zpe) 16017.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP4/aug-cc-pVDZ
ABC
0.35416 0.30391 0.16882

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP4/aug-cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.131 0.000
C2 0.964 -1.035 0.000
N3 0.236 1.399 0.000
O4 -1.302 -0.305 0.000
H5 2.006 -0.677 0.000
H6 0.788 -1.661 0.892
H7 0.788 -1.661 -0.892
H8 1.249 1.563 0.000
H9 -1.848 0.502 0.000

Atom - Atom Distances (Å)
  C1 C2 N3 O4 H5 H6 H7 H8 H9
C11.51261.29001.37332.16252.15112.15111.90041.8847
C21.51262.54032.38091.10131.10421.10422.61343.2044
N31.29002.54032.29572.72833.23493.23491.02612.2688
O41.37332.38092.29573.32902.64652.64653.16200.9740
H52.16251.10132.72833.32901.80191.80192.36494.0300
H62.15111.10423.23492.64651.80191.78453.37693.5244
H72.15111.10423.23492.64651.80191.78453.37693.5244
H81.90042.61341.02613.16202.36493.37693.37693.2735
H91.88473.20442.26880.97404.03003.52443.52443.2735

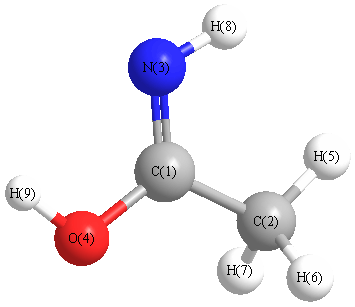
picture of Ethaninidic acid state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 H5 110.660 C1 C2 H6 109.594
C1 C2 H7 109.594 C1 N3 H8 109.745
C1 O4 H9 105.564 C2 C1 N3 129.862
C2 C1 O4 111.096 N3 C1 O4 119.042
H5 C2 H6 109.568 H5 C2 H7 109.568
H6 C2 H7 107.806
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability