Jump to
S1C2
Energy calculated at HF/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -343.092755 |
| Energy at 298.15K | -343.110357 |
| Nuclear repulsion energy | 426.670472 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1' |
3203 |
2917 |
0.00 |
|
|
|
| 2 |
A1' |
1635 |
1489 |
0.00 |
|
|
|
| 3 |
A1' |
1491 |
1357 |
0.00 |
|
|
|
| 4 |
A1' |
1062 |
967 |
0.00 |
|
|
|
| 5 |
A1' |
875 |
797 |
0.00 |
|
|
|
| 6 |
A1' |
651 |
593 |
0.00 |
|
|
|
| 7 |
A1" |
3207 |
2921 |
0.00 |
|
|
|
| 8 |
A1" |
1360 |
1238 |
0.00 |
|
|
|
| 9 |
A1" |
1106 |
1007 |
0.00 |
|
|
|
| 10 |
A1" |
46 |
42 |
0.00 |
|
|
|
| 11 |
A2' |
3233 |
2944 |
0.00 |
|
|
|
| 12 |
A2' |
1278 |
1164 |
0.00 |
|
|
|
| 13 |
A2' |
865 |
788 |
0.00 |
|
|
|
| 14 |
A2" |
3183 |
2898 |
121.60 |
|
|
|
| 15 |
A2" |
1619 |
1475 |
6.07 |
|
|
|
| 16 |
A2" |
1499 |
1365 |
7.72 |
|
|
|
| 17 |
A2" |
1070 |
974 |
31.46 |
|
|
|
| 18 |
A2" |
847 |
772 |
67.37 |
|
|
|
| 19 |
E' |
3245 |
2955 |
140.07 |
|
|
|
| 19 |
E' |
3245 |
2955 |
140.07 |
|
|
|
| 20 |
E' |
3190 |
2905 |
136.43 |
|
|
|
| 20 |
E' |
3190 |
2905 |
136.43 |
|
|
|
| 21 |
E' |
1615 |
1470 |
5.30 |
|
|
|
| 21 |
E' |
1615 |
1470 |
5.30 |
|
|
|
| 22 |
E' |
1470 |
1339 |
18.78 |
|
|
|
| 22 |
E' |
1470 |
1339 |
18.78 |
|
|
|
| 23 |
E' |
1436 |
1307 |
1.43 |
|
|
|
| 23 |
E' |
1436 |
1307 |
1.43 |
|
|
|
| 24 |
E' |
1191 |
1085 |
50.20 |
|
|
|
| 24 |
E' |
1191 |
1085 |
50.20 |
|
|
|
| 25 |
E' |
971 |
884 |
7.78 |
|
|
|
| 25 |
E' |
971 |
884 |
7.78 |
|
|
|
| 26 |
E' |
894 |
815 |
4.69 |
|
|
|
| 26 |
E' |
894 |
815 |
4.69 |
|
|
|
| 27 |
E' |
460 |
419 |
0.09 |
|
|
|
| 27 |
E' |
460 |
419 |
0.09 |
|
|
|
| 28 |
E" |
3215 |
2927 |
0.00 |
|
|
|
| 28 |
E" |
3215 |
2927 |
0.00 |
|
|
|
| 29 |
E" |
3175 |
2892 |
0.00 |
|
|
|
| 29 |
E" |
3175 |
2892 |
0.00 |
|
|
|
| 30 |
E" |
1601 |
1458 |
0.00 |
|
|
|
| 30 |
E" |
1601 |
1458 |
0.00 |
|
|
|
| 31 |
E" |
1462 |
1331 |
0.00 |
|
|
|
| 31 |
E" |
1462 |
1331 |
0.00 |
|
|
|
| 32 |
E" |
1422 |
1295 |
0.00 |
|
|
|
| 32 |
E" |
1422 |
1295 |
0.00 |
|
|
|
| 33 |
E" |
1317 |
1199 |
0.00 |
|
|
|
| 33 |
E" |
1317 |
1199 |
0.00 |
|
|
|
| 34 |
E" |
1133 |
1031 |
0.00 |
|
|
|
| 34 |
E" |
1133 |
1031 |
0.00 |
|
|
|
| 35 |
E" |
636 |
579 |
0.00 |
|
|
|
| 35 |
E" |
636 |
579 |
0.00 |
|
|
|
| 36 |
E" |
356 |
324 |
0.00 |
|
|
|
| 36 |
E" |
356 |
324 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 42902.7 cm
-1
Scaled (by 0.9106) Zero Point Vibrational Energy (zpe) 39067.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/aug-cc-pVDZ
Point Group is D3h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
0.000 |
0.000 |
1.276 |
| N2 |
0.000 |
0.000 |
-1.276 |
| C3 |
0.000 |
1.374 |
0.778 |
| C4 |
1.190 |
-0.687 |
0.778 |
| C5 |
-1.190 |
-0.687 |
0.778 |
| C6 |
0.000 |
1.374 |
-0.778 |
| C7 |
-1.190 |
-0.687 |
-0.778 |
| C8 |
1.190 |
-0.687 |
-0.778 |
| H9 |
0.877 |
1.885 |
1.175 |
| H10 |
-0.877 |
1.885 |
1.175 |
| H11 |
1.194 |
-1.702 |
1.175 |
| H12 |
2.071 |
-0.183 |
1.175 |
| H13 |
-2.071 |
-0.183 |
1.175 |
| H14 |
-1.194 |
-1.702 |
1.175 |
| H15 |
-0.877 |
1.885 |
-1.175 |
| H16 |
0.877 |
1.885 |
-1.175 |
| H17 |
-1.194 |
-1.702 |
-1.175 |
| H18 |
-2.071 |
-0.183 |
-1.175 |
| H19 |
2.071 |
-0.183 |
-1.175 |
| H20 |
1.194 |
-1.702 |
-1.175 |
Atom - Atom Distances (Å)
| |
N1 |
N2 |
C3 |
C4 |
C5 |
C6 |
C7 |
C8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
H19 |
H20 |
| N1 | | 2.5518 | 1.4616 | 1.4616 | 1.4616 | 2.4711 | 2.4711 | 2.4711 | 2.0813 | 2.0813 | 2.0813 | 2.0813 | 2.0813 | 2.0813 | 3.2136 | 3.2136 | 3.2136 | 3.2136 | 3.2136 | 3.2136 |
N2 | 2.5518 | | 2.4711 | 2.4711 | 2.4711 | 1.4616 | 1.4616 | 1.4616 | 3.2136 | 3.2136 | 3.2136 | 3.2136 | 3.2136 | 3.2136 | 2.0813 | 2.0813 | 2.0813 | 2.0813 | 2.0813 | 2.0813 | C3 | 1.4616 | 2.4711 | | 2.3800 | 2.3800 | 1.5558 | 2.8434 | 2.8434 | 1.0898 | 1.0898 | 3.3234 | 2.6210 | 2.6210 | 3.3234 | 2.2006 | 2.2006 | 3.8341 | 3.2442 | 3.2442 | 3.8341 | C4 | 1.4616 | 2.4711 | 2.3800 | | 2.3800 | 2.8434 | 2.8434 | 1.5558 | 2.6210 | 3.3234 | 1.0898 | 1.0898 | 3.3234 | 2.6210 | 3.8341 | 3.2442 | 3.2442 | 3.8341 | 2.2006 | 2.2006 | C5 | 1.4616 | 2.4711 | 2.3800 | 2.3800 | | 2.8434 | 1.5558 | 2.8434 | 3.3234 | 2.6210 | 2.6210 | 3.3234 | 1.0898 | 1.0898 | 3.2442 | 3.8341 | 2.2006 | 2.2006 | 3.8341 | 3.2442 | C6 | 2.4711 | 1.4616 | 1.5558 | 2.8434 | 2.8434 | | 2.3800 | 2.3800 | 2.2006 | 2.2006 | 3.8341 | 3.2442 | 3.2442 | 3.8341 | 1.0898 | 1.0898 | 3.3234 | 2.6210 | 2.6210 | 3.3234 | C7 | 2.4711 | 1.4616 | 2.8434 | 2.8434 | 1.5558 | 2.3800 | | 2.3800 | 3.8341 | 3.2442 | 3.2442 | 3.8341 | 2.2006 | 2.2006 | 2.6210 | 3.3234 | 1.0898 | 1.0898 | 3.3234 | 2.6210 | C8 | 2.4711 | 1.4616 | 2.8434 | 1.5558 | 2.8434 | 2.3800 | 2.3800 | | 3.2442 | 3.8341 | 2.2006 | 2.2006 | 3.8341 | 3.2442 | 3.3234 | 2.6210 | 2.6210 | 3.3234 | 1.0898 | 1.0898 | H9 | 2.0813 | 3.2136 | 1.0898 | 2.6210 | 3.3234 | 2.2006 | 3.8341 | 3.2442 | | 1.7542 | 3.6007 | 2.3875 | 3.6007 | 4.1417 | 2.9321 | 2.3495 | 4.7617 | 4.2995 | 3.3496 | 4.2995 | H10 | 2.0813 | 3.2136 | 1.0898 | 3.3234 | 2.6210 | 2.2006 | 3.2442 | 3.8341 | 1.7542 | | 4.1417 | 3.6007 | 2.3875 | 3.6007 | 2.3495 | 2.9321 | 4.2995 | 3.3496 | 4.2995 | 4.7617 | H11 | 2.0813 | 3.2136 | 3.3234 | 1.0898 | 2.6210 | 3.8341 | 3.2442 | 2.2006 | 3.6007 | 4.1417 | | 1.7542 | 3.6007 | 2.3875 | 4.7617 | 4.2995 | 3.3496 | 4.2995 | 2.9321 | 2.3495 | H12 | 2.0813 | 3.2136 | 2.6210 | 1.0898 | 3.3234 | 3.2442 | 3.8341 | 2.2006 | 2.3875 | 3.6007 | 1.7542 | | 4.1417 | 3.6007 | 4.2995 | 3.3496 | 4.2995 | 4.7617 | 2.3495 | 2.9321 | H13 | 2.0813 | 3.2136 | 2.6210 | 3.3234 | 1.0898 | 3.2442 | 2.2006 | 3.8341 | 3.6007 | 2.3875 | 3.6007 | 4.1417 | | 1.7542 | 3.3496 | 4.2995 | 2.9321 | 2.3495 | 4.7617 | 4.2995 | H14 | 2.0813 | 3.2136 | 3.3234 | 2.6210 | 1.0898 | 3.8341 | 2.2006 | 3.2442 | 4.1417 | 3.6007 | 2.3875 | 3.6007 | 1.7542 | | 4.2995 | 4.7617 | 2.3495 | 2.9321 | 4.2995 | 3.3496 | H15 | 3.2136 | 2.0813 | 2.2006 | 3.8341 | 3.2442 | 1.0898 | 2.6210 | 3.3234 | 2.9321 | 2.3495 | 4.7617 | 4.2995 | 3.3496 | 4.2995 | | 1.7542 | 3.6007 | 2.3875 | 3.6007 | 4.1417 | H16 | 3.2136 | 2.0813 | 2.2006 | 3.2442 | 3.8341 | 1.0898 | 3.3234 | 2.6210 | 2.3495 | 2.9321 | 4.2995 | 3.3496 | 4.2995 | 4.7617 | 1.7542 | | 4.1417 | 3.6007 | 2.3875 | 3.6007 | H17 | 3.2136 | 2.0813 | 3.8341 | 3.2442 | 2.2006 | 3.3234 | 1.0898 | 2.6210 | 4.7617 | 4.2995 | 3.3496 | 4.2995 | 2.9321 | 2.3495 | 3.6007 | 4.1417 | | 1.7542 | 3.6007 | 2.3875 | H18 | 3.2136 | 2.0813 | 3.2442 | 3.8341 | 2.2006 | 2.6210 | 1.0898 | 3.3234 | 4.2995 | 3.3496 | 4.2995 | 4.7617 | 2.3495 | 2.9321 | 2.3875 | 3.6007 | 1.7542 | | 4.1417 | 3.6007 | H19 | 3.2136 | 2.0813 | 3.2442 | 2.2006 | 3.8341 | 2.6210 | 3.3234 | 1.0898 | 3.3496 | 4.2995 | 2.9321 | 2.3495 | 4.7617 | 4.2995 | 3.6007 | 2.3875 | 3.6007 | 4.1417 | | 1.7542 | H20 | 3.2136 | 2.0813 | 3.8341 | 2.2006 | 3.2442 | 3.3234 | 2.6210 | 1.0898 | 4.2995 | 4.7617 | 2.3495 | 2.9321 | 4.2995 | 3.3496 | 4.1417 | 3.6007 | 2.3875 | 3.6007 | 1.7542 | |
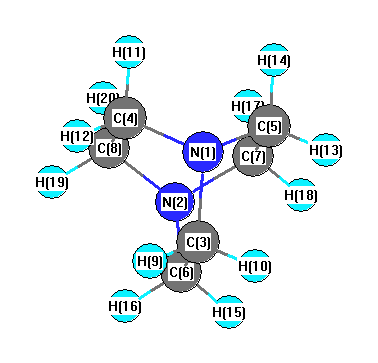 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
C3 |
C6 |
109.922 |
|
N1 |
C3 |
H9 |
108.452 |
| N1 |
C3 |
H10 |
108.452 |
|
N1 |
C4 |
C8 |
109.922 |
| N1 |
C4 |
H11 |
108.452 |
|
N1 |
C4 |
H12 |
108.452 |
| N1 |
C5 |
C7 |
109.922 |
|
N1 |
C5 |
H13 |
108.452 |
| N1 |
C5 |
H14 |
108.452 |
|
N2 |
C6 |
C3 |
109.922 |
| N2 |
C6 |
H15 |
108.452 |
|
N2 |
C6 |
H16 |
108.452 |
| N2 |
C7 |
C5 |
109.922 |
|
N2 |
C7 |
H17 |
108.452 |
| N2 |
C7 |
H18 |
108.452 |
|
N2 |
C8 |
C4 |
109.922 |
| N2 |
C8 |
H19 |
108.452 |
|
N2 |
C8 |
H20 |
108.452 |
| C3 |
N1 |
C4 |
109.016 |
|
C3 |
N1 |
C5 |
109.016 |
| C3 |
C6 |
H15 |
111.356 |
|
C3 |
C6 |
H16 |
111.356 |
| C4 |
N1 |
C5 |
109.016 |
|
C4 |
C8 |
H19 |
111.356 |
| C4 |
C8 |
H20 |
111.356 |
|
C5 |
C6 |
H15 |
101.661 |
| C5 |
C6 |
H16 |
151.140 |
|
C6 |
N2 |
C7 |
109.016 |
| C6 |
N2 |
C8 |
109.016 |
|
C6 |
C3 |
H9 |
111.356 |
| C6 |
C3 |
H10 |
111.356 |
|
C7 |
N2 |
C8 |
109.016 |
| C7 |
C5 |
H13 |
111.356 |
|
C7 |
C5 |
H14 |
111.356 |
| C8 |
C4 |
H11 |
111.356 |
|
C8 |
C4 |
H12 |
111.356 |
| H9 |
C3 |
H10 |
107.190 |
|
H11 |
C4 |
H12 |
107.190 |
| H13 |
C5 |
H14 |
107.190 |
|
H15 |
C6 |
H16 |
107.190 |
| H17 |
C7 |
H18 |
107.190 |
|
H19 |
C8 |
H20 |
107.190 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.756 |
|
|
|
| 2 |
N |
-0.756 |
|
|
|
| 3 |
C |
0.664 |
|
|
|
| 4 |
C |
0.664 |
|
|
|
| 5 |
C |
0.664 |
|
|
|
| 6 |
C |
0.664 |
|
|
|
| 7 |
C |
0.664 |
|
|
|
| 8 |
C |
0.664 |
|
|
|
| 9 |
H |
-0.206 |
|
|
|
| 10 |
H |
-0.206 |
|
|
|
| 11 |
H |
-0.206 |
|
|
|
| 12 |
H |
-0.206 |
|
|
|
| 13 |
H |
-0.206 |
|
|
|
| 14 |
H |
-0.206 |
|
|
|
| 15 |
H |
-0.206 |
|
|
|
| 16 |
H |
-0.206 |
|
|
|
| 17 |
H |
-0.206 |
|
|
|
| 18 |
H |
-0.206 |
|
|
|
| 19 |
H |
-0.206 |
|
|
|
| 20 |
H |
-0.206 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-47.230 |
0.000 |
0.000 |
| y |
0.000 |
-47.230 |
0.000 |
| z |
0.000 |
0.000 |
-58.866 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.818 |
0.000 |
0.000 |
| y |
0.000 |
5.818 |
0.000 |
| z |
0.000 |
0.000 |
-11.636 |
|
| Polar |
|---|
| 3z2-r2 | -23.272 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.615 |
0.000 |
0.000 |
| y |
0.000 |
11.615 |
0.000 |
| z |
0.000 |
0.000 |
10.939 |
<r2> (average value of r
2) Å
2
| <r2> |
212.890 |
| (<r2>)1/2 |
14.591 |
Jump to
S1C1
Energy calculated at HF/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -343.092755 |
| Energy at 298.15K | -343.110357 |
| HF Energy | -343.092755 |
| Nuclear repulsion energy | 426.667453 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3207 |
2921 |
0.00 |
|
|
|
| 2 |
A1 |
3203 |
2917 |
0.00 |
|
|
|
| 3 |
A1 |
1635 |
1489 |
0.00 |
|
|
|
| 4 |
A1 |
1491 |
1357 |
0.00 |
|
|
|
| 5 |
A1 |
1360 |
1238 |
0.00 |
|
|
|
| 6 |
A1 |
1106 |
1007 |
0.00 |
|
|
|
| 7 |
A1 |
1062 |
967 |
0.00 |
|
|
|
| 8 |
A1 |
875 |
797 |
0.00 |
|
|
|
| 9 |
A1 |
651 |
593 |
0.00 |
|
|
|
| 10 |
A1 |
46 |
42 |
0.00 |
|
|
|
| 11 |
A2 |
3233 |
2944 |
0.00 |
|
|
|
| 12 |
A2 |
3183 |
2898 |
121.59 |
|
|
|
| 13 |
A2 |
1619 |
1475 |
6.06 |
|
|
|
| 14 |
A2 |
1499 |
1365 |
7.73 |
|
|
|
| 15 |
A2 |
1278 |
1164 |
0.00 |
|
|
|
| 16 |
A2 |
1070 |
974 |
31.47 |
|
|
|
| 17 |
A2 |
865 |
788 |
0.00 |
|
|
|
| 18 |
A2 |
847 |
771 |
67.34 |
|
|
|
| 19 |
E |
3245 |
2955 |
140.06 |
|
|
|
| 19 |
E |
3245 |
2955 |
140.06 |
|
|
|
| 20 |
E |
3215 |
2927 |
0.00 |
|
|
|
| 20 |
E |
3215 |
2927 |
0.00 |
|
|
|
| 21 |
E |
3190 |
2905 |
136.42 |
|
|
|
| 21 |
E |
3190 |
2905 |
136.42 |
|
|
|
| 22 |
E |
3175 |
2892 |
0.00 |
|
|
|
| 22 |
E |
3175 |
2892 |
0.00 |
|
|
|
| 23 |
E |
1615 |
1470 |
5.30 |
|
|
|
| 23 |
E |
1615 |
1470 |
5.30 |
|
|
|
| 24 |
E |
1601 |
1458 |
0.00 |
|
|
|
| 24 |
E |
1601 |
1458 |
0.00 |
|
|
|
| 25 |
E |
1470 |
1339 |
18.76 |
|
|
|
| 25 |
E |
1470 |
1339 |
18.76 |
|
|
|
| 26 |
E |
1462 |
1331 |
0.00 |
|
|
|
| 26 |
E |
1462 |
1331 |
0.00 |
|
|
|
| 27 |
E |
1436 |
1307 |
1.43 |
|
|
|
| 27 |
E |
1436 |
1307 |
1.43 |
|
|
|
| 28 |
E |
1423 |
1295 |
0.00 |
|
|
|
| 28 |
E |
1423 |
1295 |
0.00 |
|
|
|
| 29 |
E |
1317 |
1199 |
0.00 |
|
|
|
| 29 |
E |
1317 |
1199 |
0.00 |
|
|
|
| 30 |
E |
1191 |
1085 |
50.18 |
|
|
|
| 30 |
E |
1191 |
1085 |
50.18 |
|
|
|
| 31 |
E |
1133 |
1031 |
0.00 |
|
|
|
| 31 |
E |
1133 |
1031 |
0.00 |
|
|
|
| 32 |
E |
971 |
884 |
7.79 |
|
|
|
| 32 |
E |
971 |
884 |
7.79 |
|
|
|
| 33 |
E |
894 |
815 |
4.70 |
|
|
|
| 33 |
E |
894 |
815 |
4.70 |
|
|
|
| 34 |
E |
636 |
579 |
0.00 |
|
|
|
| 34 |
E |
636 |
579 |
0.00 |
|
|
|
| 35 |
E |
460 |
419 |
0.09 |
|
|
|
| 35 |
E |
460 |
419 |
0.09 |
|
|
|
| 36 |
E |
356 |
324 |
0.00 |
|
|
|
| 36 |
E |
356 |
324 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 42902.5 cm
-1
Scaled (by 0.9106) Zero Point Vibrational Energy (zpe) 39067.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/aug-cc-pVDZ
Point Group is D3
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
0.000 |
0.000 |
1.276 |
| N2 |
0.000 |
0.000 |
-1.276 |
| C3 |
0.000 |
1.374 |
0.778 |
| C4 |
1.190 |
-0.687 |
0.778 |
| C5 |
-1.190 |
-0.687 |
0.778 |
| C6 |
-0.000 |
1.374 |
-0.778 |
| C7 |
-1.190 |
-0.687 |
-0.778 |
| C8 |
1.190 |
-0.687 |
-0.778 |
| H9 |
0.878 |
1.884 |
1.174 |
| H10 |
-0.876 |
1.885 |
1.175 |
| H11 |
1.193 |
-1.702 |
1.174 |
| H12 |
2.071 |
-0.183 |
1.175 |
| H13 |
-2.071 |
-0.182 |
1.174 |
| H14 |
-1.194 |
-1.702 |
1.175 |
| H15 |
-0.878 |
1.884 |
-1.174 |
| H16 |
0.876 |
1.885 |
-1.175 |
| H17 |
-1.193 |
-1.702 |
-1.174 |
| H18 |
-2.071 |
-0.183 |
-1.175 |
| H19 |
2.071 |
-0.182 |
-1.174 |
| H20 |
1.194 |
-1.702 |
-1.175 |
Atom - Atom Distances (Å)
| |
N1 |
N2 |
C3 |
C4 |
C5 |
C6 |
C7 |
C8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
H19 |
H20 |
| N1 | | 2.5518 | 1.4616 | 1.4616 | 1.4616 | 2.4711 | 2.4711 | 2.4711 | 2.0813 | 2.0814 | 2.0813 | 2.0814 | 2.0813 | 2.0814 | 3.2134 | 3.2140 | 3.2134 | 3.2140 | 3.2134 | 3.2140 |
N2 | 2.5518 | | 2.4711 | 2.4711 | 2.4711 | 1.4616 | 1.4616 | 1.4616 | 3.2134 | 3.2140 | 3.2134 | 3.2140 | 3.2134 | 3.2140 | 2.0813 | 2.0814 | 2.0813 | 2.0814 | 2.0813 | 2.0814 | C3 | 1.4616 | 2.4711 | | 2.3800 | 2.3800 | 1.5557 | 2.8436 | 2.8432 | 1.0898 | 1.0898 | 3.3234 | 2.6213 | 2.6208 | 3.3234 | 2.2006 | 2.2007 | 3.8341 | 3.2449 | 3.2436 | 3.8341 | C4 | 1.4616 | 2.4711 | 2.3800 | | 2.3800 | 2.8436 | 2.8432 | 1.5557 | 2.6208 | 3.3234 | 1.0898 | 1.0898 | 3.3234 | 2.6213 | 3.8341 | 3.2449 | 3.2436 | 3.8341 | 2.2006 | 2.2007 | C5 | 1.4616 | 2.4711 | 2.3800 | 2.3800 | | 2.8432 | 1.5557 | 2.8436 | 3.3234 | 2.6213 | 2.6208 | 3.3234 | 1.0898 | 1.0898 | 3.2436 | 3.8341 | 2.2006 | 2.2007 | 3.8341 | 3.2449 | C6 | 2.4711 | 1.4616 | 1.5557 | 2.8436 | 2.8432 | | 2.3800 | 2.3800 | 2.2006 | 2.2007 | 3.8341 | 3.2449 | 3.2436 | 3.8341 | 1.0898 | 1.0898 | 3.3234 | 2.6213 | 2.6208 | 3.3234 | C7 | 2.4711 | 1.4616 | 2.8436 | 2.8432 | 1.5557 | 2.3800 | | 2.3800 | 3.8341 | 3.2449 | 3.2436 | 3.8341 | 2.2006 | 2.2007 | 2.6208 | 3.3234 | 1.0898 | 1.0898 | 3.3234 | 2.6213 | C8 | 2.4711 | 1.4616 | 2.8432 | 1.5557 | 2.8436 | 2.3800 | 2.3800 | | 3.2436 | 3.8341 | 2.2006 | 2.2007 | 3.8341 | 3.2449 | 3.3234 | 2.6213 | 2.6208 | 3.3234 | 1.0898 | 1.0898 | H9 | 2.0813 | 3.2134 | 1.0898 | 2.6208 | 3.3234 | 2.2006 | 3.8341 | 3.2436 | | 1.7542 | 3.6006 | 2.3875 | 3.6006 | 4.1416 | 2.9324 | 2.3496 | 4.7613 | 4.3001 | 3.3484 | 4.2989 | H10 | 2.0814 | 3.2140 | 1.0898 | 3.3234 | 2.6213 | 2.2007 | 3.2449 | 3.8341 | 1.7542 | | 4.1416 | 3.6008 | 2.3875 | 3.6008 | 2.3496 | 2.9320 | 4.3001 | 3.3510 | 4.2989 | 4.7620 | H11 | 2.0813 | 3.2134 | 3.3234 | 1.0898 | 2.6208 | 3.8341 | 3.2436 | 2.2006 | 3.6006 | 4.1416 | | 1.7542 | 3.6006 | 2.3875 | 4.7613 | 4.3001 | 3.3484 | 4.2989 | 2.9324 | 2.3496 | H12 | 2.0814 | 3.2140 | 2.6213 | 1.0898 | 3.3234 | 3.2449 | 3.8341 | 2.2007 | 2.3875 | 3.6008 | 1.7542 | | 4.1416 | 3.6008 | 4.3001 | 3.3510 | 4.2989 | 4.7620 | 2.3496 | 2.9320 | H13 | 2.0813 | 3.2134 | 2.6208 | 3.3234 | 1.0898 | 3.2436 | 2.2006 | 3.8341 | 3.6006 | 2.3875 | 3.6006 | 4.1416 | | 1.7542 | 3.3484 | 4.2989 | 2.9324 | 2.3496 | 4.7613 | 4.3001 | H14 | 2.0814 | 3.2140 | 3.3234 | 2.6213 | 1.0898 | 3.8341 | 2.2007 | 3.2449 | 4.1416 | 3.6008 | 2.3875 | 3.6008 | 1.7542 | | 4.2989 | 4.7620 | 2.3496 | 2.9320 | 4.3001 | 3.3510 | H15 | 3.2134 | 2.0813 | 2.2006 | 3.8341 | 3.2436 | 1.0898 | 2.6208 | 3.3234 | 2.9324 | 2.3496 | 4.7613 | 4.3001 | 3.3484 | 4.2989 | | 1.7542 | 3.6006 | 2.3875 | 3.6006 | 4.1416 | H16 | 3.2140 | 2.0814 | 2.2007 | 3.2449 | 3.8341 | 1.0898 | 3.3234 | 2.6213 | 2.3496 | 2.9320 | 4.3001 | 3.3510 | 4.2989 | 4.7620 | 1.7542 | | 4.1416 | 3.6008 | 2.3875 | 3.6008 | H17 | 3.2134 | 2.0813 | 3.8341 | 3.2436 | 2.2006 | 3.3234 | 1.0898 | 2.6208 | 4.7613 | 4.3001 | 3.3484 | 4.2989 | 2.9324 | 2.3496 | 3.6006 | 4.1416 | | 1.7542 | 3.6006 | 2.3875 | H18 | 3.2140 | 2.0814 | 3.2449 | 3.8341 | 2.2007 | 2.6213 | 1.0898 | 3.3234 | 4.3001 | 3.3510 | 4.2989 | 4.7620 | 2.3496 | 2.9320 | 2.3875 | 3.6008 | 1.7542 | | 4.1416 | 3.6008 | H19 | 3.2134 | 2.0813 | 3.2436 | 2.2006 | 3.8341 | 2.6208 | 3.3234 | 1.0898 | 3.3484 | 4.2989 | 2.9324 | 2.3496 | 4.7613 | 4.3001 | 3.6006 | 2.3875 | 3.6006 | 4.1416 | | 1.7542 | H20 | 3.2140 | 2.0814 | 3.8341 | 2.2007 | 3.2449 | 3.3234 | 2.6213 | 1.0898 | 4.2989 | 4.7620 | 2.3496 | 2.9320 | 4.3001 | 3.3510 | 4.1416 | 3.6008 | 2.3875 | 3.6008 | 1.7542 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
C3 |
C6 |
109.924 |
|
N1 |
C3 |
H9 |
108.444 |
| N1 |
C3 |
H10 |
108.451 |
|
N1 |
C4 |
C8 |
109.924 |
| N1 |
C4 |
H11 |
108.444 |
|
N1 |
C4 |
H12 |
108.450 |
| N1 |
C5 |
C7 |
109.924 |
|
N1 |
C5 |
H13 |
108.444 |
| N1 |
C5 |
H14 |
108.450 |
|
N2 |
C6 |
C3 |
109.924 |
| N2 |
C6 |
H15 |
108.444 |
|
N2 |
C6 |
H16 |
108.451 |
| N2 |
C7 |
C5 |
109.924 |
|
N2 |
C7 |
H17 |
108.444 |
| N2 |
C7 |
H18 |
108.450 |
|
N2 |
C8 |
C4 |
109.924 |
| N2 |
C8 |
H19 |
108.444 |
|
N2 |
C8 |
H20 |
108.450 |
| C3 |
N1 |
C4 |
109.015 |
|
C3 |
N1 |
C5 |
109.015 |
| C3 |
C6 |
H15 |
111.358 |
|
C3 |
C6 |
H16 |
111.363 |
| C4 |
N1 |
C5 |
109.015 |
|
C4 |
C8 |
H19 |
111.358 |
| C4 |
C8 |
H20 |
111.363 |
|
C5 |
C6 |
H15 |
101.631 |
| C5 |
C6 |
H16 |
151.171 |
|
C6 |
N2 |
C7 |
109.015 |
| C6 |
N2 |
C8 |
109.015 |
|
C6 |
C3 |
H9 |
111.358 |
| C6 |
C3 |
H10 |
111.363 |
|
C7 |
N2 |
C8 |
109.015 |
| C7 |
C5 |
H13 |
111.358 |
|
C7 |
C5 |
H14 |
111.363 |
| C8 |
C4 |
H11 |
111.358 |
|
C8 |
C4 |
H12 |
111.363 |
| H9 |
C3 |
H10 |
107.188 |
|
H11 |
C4 |
H12 |
107.188 |
| H13 |
C5 |
H14 |
107.188 |
|
H15 |
C6 |
H16 |
107.188 |
| H17 |
C7 |
H18 |
107.188 |
|
H19 |
C8 |
H20 |
107.188 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.757 |
|
|
|
| 2 |
N |
-0.757 |
|
|
|
| 3 |
C |
0.664 |
|
|
|
| 4 |
C |
0.664 |
|
|
|
| 5 |
C |
0.664 |
|
|
|
| 6 |
C |
0.664 |
|
|
|
| 7 |
C |
0.664 |
|
|
|
| 8 |
C |
0.664 |
|
|
|
| 9 |
H |
-0.206 |
|
|
|
| 10 |
H |
-0.207 |
|
|
|
| 11 |
H |
-0.206 |
|
|
|
| 12 |
H |
-0.207 |
|
|
|
| 13 |
H |
-0.206 |
|
|
|
| 14 |
H |
-0.207 |
|
|
|
| 15 |
H |
-0.206 |
|
|
|
| 16 |
H |
-0.207 |
|
|
|
| 17 |
H |
-0.206 |
|
|
|
| 18 |
H |
-0.207 |
|
|
|
| 19 |
H |
-0.206 |
|
|
|
| 20 |
H |
-0.207 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-47.231 |
0.000 |
0.000 |
| y |
0.000 |
-47.231 |
0.000 |
| z |
0.000 |
0.000 |
-58.866 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.817 |
0.000 |
0.000 |
| y |
0.000 |
5.817 |
0.000 |
| z |
0.000 |
0.000 |
-11.635 |
|
| Polar |
|---|
| 3z2-r2 | -23.270 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.615 |
0.000 |
0.000 |
| y |
0.000 |
11.615 |
0.000 |
| z |
0.000 |
0.000 |
10.939 |
<r2> (average value of r
2) Å
2
| <r2> |
212.893 |
| (<r2>)1/2 |
14.591 |
