Jump to
S1C2
Vibrational Frequencies calculated at CCSD=FULL/aug-cc-pVDZ
Geometric Data calculated at CCSD=FULL/aug-cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCSD=FULL/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -344.417938 |
| Energy at 298.15K | |
| HF Energy | -343.089742 |
| Nuclear repulsion energy | 423.086649 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD=FULL/aug-cc-pVDZ
Geometric Data calculated at CCSD=FULL/aug-cc-pVDZ
Point Group is D3
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
0.000 |
0.000 |
1.300 |
| N2 |
0.000 |
0.000 |
-1.300 |
| C3 |
0.074 |
1.381 |
0.779 |
| C4 |
1.159 |
-0.754 |
0.779 |
| C5 |
-1.233 |
-0.627 |
0.779 |
| C6 |
-0.074 |
1.381 |
-0.779 |
| C7 |
-1.159 |
-0.754 |
-0.779 |
| C8 |
1.233 |
-0.627 |
-0.779 |
| H9 |
1.044 |
1.804 |
1.088 |
| H10 |
-0.721 |
1.978 |
1.258 |
| H11 |
1.040 |
-1.807 |
1.088 |
| H12 |
2.073 |
-0.365 |
1.258 |
| H13 |
-2.085 |
0.002 |
1.088 |
| H14 |
-1.353 |
-1.613 |
1.258 |
| H15 |
-1.044 |
1.804 |
-1.088 |
| H16 |
0.721 |
1.978 |
-1.258 |
| H17 |
-1.040 |
-1.807 |
-1.088 |
| H18 |
-2.073 |
-0.365 |
-1.258 |
| H19 |
2.085 |
0.002 |
-1.088 |
| H20 |
1.353 |
-1.613 |
-1.258 |
Atom - Atom Distances (Å)
| |
N1 |
N2 |
C3 |
C4 |
C5 |
C6 |
C7 |
C8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
H19 |
H20 |
| N1 | | 2.5998 | 1.4782 | 1.4782 | 1.4782 | 2.4966 | 2.4966 | 2.4966 | 2.0954 | 2.1056 | 2.0954 | 2.1056 | 2.0954 | 2.1056 | 3.1701 | 3.3131 | 3.1701 | 3.3131 | 3.1701 | 3.3131 |
N2 | 2.5998 | | 2.4966 | 2.4966 | 2.4966 | 1.4782 | 1.4782 | 1.4782 | 3.1701 | 3.3131 | 3.1701 | 3.3131 | 3.1701 | 3.3131 | 2.0954 | 2.1056 | 2.0954 | 2.1056 | 2.0954 | 2.1056 | C3 | 1.4782 | 2.4966 | | 2.3959 | 2.3959 | 1.5640 | 2.9165 | 2.7932 | 1.1033 | 1.1032 | 3.3456 | 2.6978 | 2.5798 | 3.3512 | 2.2168 | 2.2189 | 3.8586 | 3.4361 | 3.0711 | 3.8408 | C4 | 1.4782 | 2.4966 | 2.3959 | | 2.3959 | 2.9165 | 2.7932 | 1.5640 | 2.5798 | 3.3512 | 1.1033 | 1.1032 | 3.3456 | 2.6978 | 3.8586 | 3.4361 | 3.0711 | 3.8408 | 2.2168 | 2.2189 | C5 | 1.4782 | 2.4966 | 2.3959 | 2.3959 | | 2.7932 | 1.5640 | 2.9165 | 3.3456 | 2.6978 | 2.5798 | 3.3512 | 1.1033 | 1.1032 | 3.0711 | 3.8408 | 2.2168 | 2.2189 | 3.8586 | 3.4361 | C6 | 2.4966 | 1.4782 | 1.5640 | 2.9165 | 2.7932 | | 2.3959 | 2.3959 | 2.2168 | 2.2189 | 3.8586 | 3.4361 | 3.0711 | 3.8408 | 1.1033 | 1.1032 | 3.3456 | 2.6978 | 2.5798 | 3.3512 | C7 | 2.4966 | 1.4782 | 2.9165 | 2.7932 | 1.5640 | 2.3959 | | 2.3959 | 3.8586 | 3.4361 | 3.0711 | 3.8408 | 2.2168 | 2.2189 | 2.5798 | 3.3512 | 1.1033 | 1.1032 | 3.3456 | 2.6978 | C8 | 2.4966 | 1.4782 | 2.7932 | 1.5640 | 2.9165 | 2.3959 | 2.3959 | | 3.0711 | 3.8408 | 2.2168 | 2.2189 | 3.8586 | 3.4361 | 3.3456 | 2.6978 | 2.5798 | 3.3512 | 1.1033 | 1.1032 | H9 | 2.0954 | 3.1701 | 1.1033 | 2.5798 | 3.3456 | 2.2168 | 3.8586 | 3.0711 | | 1.7817 | 3.6107 | 2.4067 | 3.6107 | 4.1776 | 3.0169 | 2.3754 | 4.7034 | 4.4645 | 3.0112 | 4.1570 | H10 | 2.1056 | 3.3131 | 1.1032 | 3.3512 | 2.6978 | 2.2189 | 3.4361 | 3.8408 | 1.7817 | | 4.1776 | 3.6463 | 2.4067 | 3.6463 | 2.3754 | 2.9002 | 4.4645 | 3.6949 | 4.1570 | 4.8506 | H11 | 2.0954 | 3.1701 | 3.3456 | 1.1033 | 2.5798 | 3.8586 | 3.0711 | 2.2168 | 3.6107 | 4.1776 | | 1.7817 | 3.6107 | 2.4067 | 4.7034 | 4.4645 | 3.0112 | 4.1570 | 3.0169 | 2.3754 | H12 | 2.1056 | 3.3131 | 2.6978 | 1.1032 | 3.3512 | 3.4361 | 3.8408 | 2.2189 | 2.4067 | 3.6463 | 1.7817 | | 4.1776 | 3.6463 | 4.4645 | 3.6949 | 4.1570 | 4.8506 | 2.3754 | 2.9002 | H13 | 2.0954 | 3.1701 | 2.5798 | 3.3456 | 1.1033 | 3.0711 | 2.2168 | 3.8586 | 3.6107 | 2.4067 | 3.6107 | 4.1776 | | 1.7817 | 3.0112 | 4.1570 | 3.0169 | 2.3754 | 4.7034 | 4.4645 | H14 | 2.1056 | 3.3131 | 3.3512 | 2.6978 | 1.1032 | 3.8408 | 2.2189 | 3.4361 | 4.1776 | 3.6463 | 2.4067 | 3.6463 | 1.7817 | | 4.1570 | 4.8506 | 2.3754 | 2.9002 | 4.4645 | 3.6949 | H15 | 3.1701 | 2.0954 | 2.2168 | 3.8586 | 3.0711 | 1.1033 | 2.5798 | 3.3456 | 3.0169 | 2.3754 | 4.7034 | 4.4645 | 3.0112 | 4.1570 | | 1.7817 | 3.6107 | 2.4067 | 3.6107 | 4.1776 | H16 | 3.3131 | 2.1056 | 2.2189 | 3.4361 | 3.8408 | 1.1032 | 3.3512 | 2.6978 | 2.3754 | 2.9002 | 4.4645 | 3.6949 | 4.1570 | 4.8506 | 1.7817 | | 4.1776 | 3.6463 | 2.4067 | 3.6463 | H17 | 3.1701 | 2.0954 | 3.8586 | 3.0711 | 2.2168 | 3.3456 | 1.1033 | 2.5798 | 4.7034 | 4.4645 | 3.0112 | 4.1570 | 3.0169 | 2.3754 | 3.6107 | 4.1776 | | 1.7817 | 3.6107 | 2.4067 | H18 | 3.3131 | 2.1056 | 3.4361 | 3.8408 | 2.2189 | 2.6978 | 1.1032 | 3.3512 | 4.4645 | 3.6949 | 4.1570 | 4.8506 | 2.3754 | 2.9002 | 2.4067 | 3.6463 | 1.7817 | | 4.1776 | 3.6463 | H19 | 3.1701 | 2.0954 | 3.0711 | 2.2168 | 3.8586 | 2.5798 | 3.3456 | 1.1033 | 3.0112 | 4.1570 | 3.0169 | 2.3754 | 4.7034 | 4.4645 | 3.6107 | 2.4067 | 3.6107 | 4.1776 | | 1.7817 | H20 | 3.3131 | 2.1056 | 3.8408 | 2.2189 | 3.4361 | 3.3512 | 2.6978 | 1.1032 | 4.1570 | 4.8506 | 2.3754 | 2.9002 | 4.4645 | 3.6949 | 4.1776 | 3.6463 | 2.4067 | 3.6463 | 1.7817 | |
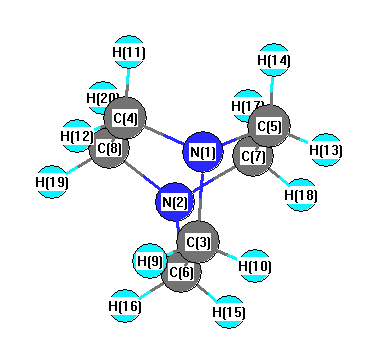 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
C3 |
C6 |
110.270 |
|
N1 |
C3 |
H9 |
107.630 |
| N1 |
C3 |
H10 |
108.431 |
|
N1 |
C4 |
C8 |
110.270 |
| N1 |
C4 |
H11 |
107.630 |
|
N1 |
C4 |
H12 |
108.431 |
| N1 |
C5 |
C7 |
110.270 |
|
N1 |
C5 |
H13 |
107.630 |
| N1 |
C5 |
H14 |
108.431 |
|
N2 |
C6 |
C3 |
110.270 |
| N2 |
C6 |
H15 |
107.630 |
|
N2 |
C6 |
H16 |
108.431 |
| N2 |
C7 |
C5 |
110.270 |
|
N2 |
C7 |
H17 |
107.630 |
| N2 |
C7 |
H18 |
108.431 |
|
N2 |
C8 |
C4 |
110.270 |
| N2 |
C8 |
H19 |
107.630 |
|
N2 |
C8 |
H20 |
108.431 |
| C3 |
N1 |
C4 |
108.265 |
|
C3 |
N1 |
C5 |
108.265 |
| C3 |
C6 |
H15 |
111.247 |
|
C3 |
C6 |
H16 |
111.426 |
| C4 |
N1 |
C5 |
108.265 |
|
C4 |
C8 |
H19 |
111.247 |
| C4 |
C8 |
H20 |
111.426 |
|
C5 |
C6 |
H15 |
93.836 |
| C5 |
C6 |
H16 |
158.461 |
|
C6 |
N2 |
C7 |
108.265 |
| C6 |
N2 |
C8 |
108.265 |
|
C6 |
C3 |
H9 |
111.247 |
| C6 |
C3 |
H10 |
111.426 |
|
C7 |
N2 |
C8 |
108.265 |
| C7 |
C5 |
H13 |
111.247 |
|
C7 |
C5 |
H14 |
111.426 |
| C8 |
C4 |
H11 |
111.247 |
|
C8 |
C4 |
H12 |
111.426 |
| H9 |
C3 |
H10 |
107.698 |
|
H11 |
C4 |
H12 |
107.698 |
| H13 |
C5 |
H14 |
107.698 |
|
H15 |
C6 |
H16 |
107.698 |
| H17 |
C7 |
H18 |
107.698 |
|
H19 |
C8 |
H20 |
107.698 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
