return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H6S3 (1,3,5-Trithiane)

using model chemistry: MP4/aug-cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C3V 1A1
Energy calculated at MP4/aug-cc-pVDZ
 hartrees
Energy at 0K-1310.665416
Energy at 298.15K-1310.673139
HF Energy-1309.688895
Nuclear repulsion energy447.378865
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A1 3148 3075        
2 A1 3063 2992        
3 A1 1484 1450        
4 A1 950 928        
5 A1 674 659        
6 A1 410 401        
7 A1 304 297        
8 A2 1258 1229        
9 A2 1162 1135        
10 A2 785 767        
11 E 3150 3077        
11 E 3150 3077        
12 E 3068 2998        
12 E 3068 2997        
13 E 1467 1433        
13 E 1466 1432        
14 E 1313 1283        
14 E 1313 1283        
15 E 1246 1217        
15 E 1245 1216        
16 E 825 806        
16 E 825 806        
17 E 767 750        
17 E 767 749        
18 E 683 668        
18 E 683 667        
19 E 288 281        
19 E 287 281        
20 E 188 183        
20 E 185 181        

Unscaled Zero Point Vibrational Energy (zpe) 19611.0 cm-1
Scaled (by 0.9769) Zero Point Vibrational Energy (zpe) 19158.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP4/aug-cc-pVDZ
ABC
0.07185 0.07185 0.03924

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP4/aug-cc-pVDZ

Point Group is C3v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 1.594 0.419
C2 1.381 -0.797 0.419
C3 -1.381 -0.797 0.419
S4 1.557 0.899 -0.263
S5 0.000 -1.798 -0.263
S6 -1.557 0.899 -0.263
H7 0.000 1.483 1.520
H8 1.284 -0.741 1.520
H9 -1.284 -0.741 1.520
H10 0.000 2.670 0.171
H11 2.312 -1.335 0.171
H12 -2.312 -1.335 0.171

Atom - Atom Distances (Å)
  C1 C2 C3 S4 S5 S6 H7 H8 H9 H10 H11 H12
C12.76142.76141.83623.45961.83621.10672.88372.88371.10353.73983.7398
C22.76142.76141.83621.83623.45962.88371.10672.88373.73981.10353.7398
C32.76142.76143.45961.83621.83622.88372.88371.10673.73983.73981.1035
S41.83621.83623.45963.11343.11342.43762.43763.73322.39732.39734.4881
S53.45961.83621.83623.11343.11343.73322.43762.43764.48812.39732.3973
S61.83623.45961.83623.11343.11342.43763.73322.43762.39734.48812.3973
H71.10672.88372.88372.43763.73322.43762.56792.56791.79703.88623.8862
H82.88371.10672.88372.43762.43763.73322.56792.56793.88621.79703.8862
H92.88372.88371.10673.73322.43762.43762.56792.56793.88623.88621.7970
H101.10353.73983.73982.39734.48812.39731.79703.88623.88624.62384.6238
H113.73981.10353.73982.39732.39734.48813.88621.79703.88624.62384.6238
H123.73983.73981.10354.48812.39732.39733.88623.88621.79704.62384.6238

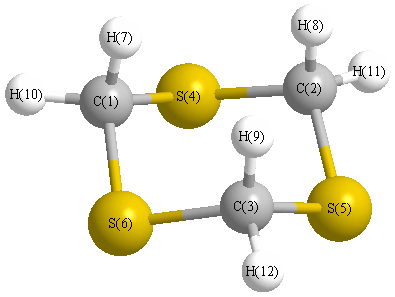
picture of 1,3,5-Trithiane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 S4 C2 97.517 C1 S6 C3 97.517
C2 S5 C3 97.517 S4 C1 S6 115.942
S4 C1 H7 109.332 S4 C1 H10 106.597
S4 C2 S5 115.942 S4 C2 H8 109.332
S4 C2 H11 106.597 S5 C2 H8 109.332
S5 C2 H11 106.597 S5 C3 S6 115.942
S5 C3 H9 109.332 S5 C3 H12 106.597
S6 C1 H7 109.332 S6 C1 H10 106.597
S6 C3 H9 109.332 S6 C3 H12 106.597
H7 C1 H10 108.789 H8 C2 H11 108.789
H9 C3 H12 108.789
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability