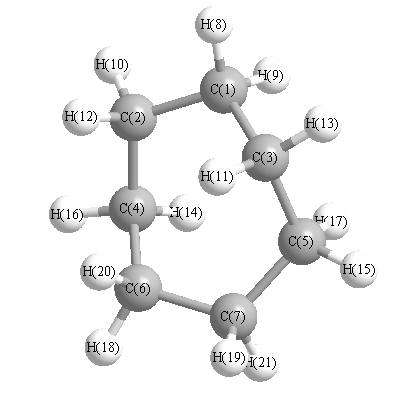
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3095 |
2966 |
5.51 |
|
|
|
| 2 |
A |
3087 |
2958 |
33.24 |
|
|
|
| 3 |
A |
3080 |
2952 |
76.11 |
|
|
|
| 4 |
A |
3052 |
2925 |
26.73 |
|
|
|
| 5 |
A |
3042 |
2915 |
3.23 |
|
|
|
| 6 |
A |
3040 |
2913 |
9.06 |
|
|
|
| 7 |
A |
3031 |
2905 |
19.83 |
|
|
|
| 8 |
A |
1497 |
1434 |
0.72 |
|
|
|
| 9 |
A |
1479 |
1417 |
7.24 |
|
|
|
| 10 |
A |
1469 |
1408 |
0.16 |
|
|
|
| 11 |
A |
1466 |
1405 |
0.59 |
|
|
|
| 12 |
A |
1395 |
1337 |
0.20 |
|
|
|
| 13 |
A |
1389 |
1331 |
0.49 |
|
|
|
| 14 |
A |
1367 |
1310 |
0.55 |
|
|
|
| 15 |
A |
1309 |
1254 |
0.00 |
|
|
|
| 16 |
A |
1298 |
1244 |
1.84 |
|
|
|
| 17 |
A |
1228 |
1177 |
0.62 |
|
|
|
| 18 |
A |
1215 |
1164 |
0.09 |
|
|
|
| 19 |
A |
1141 |
1093 |
0.10 |
|
|
|
| 20 |
A |
1049 |
1005 |
0.29 |
|
|
|
| 21 |
A |
1039 |
995 |
0.00 |
|
|
|
| 22 |
A |
960 |
920 |
1.50 |
|
|
|
| 23 |
A |
856 |
820 |
0.62 |
|
|
|
| 24 |
A |
843 |
808 |
1.64 |
|
|
|
| 25 |
A |
750 |
719 |
0.02 |
|
|
|
| 26 |
A |
517 |
495 |
0.20 |
|
|
|
| 27 |
A |
393 |
377 |
0.00 |
|
|
|
| 28 |
A |
306 |
293 |
0.00 |
|
|
|
| 29 |
A |
170 |
163 |
0.00 |
|
|
|
| 30 |
B |
3103 |
2973 |
114.59 |
|
|
|
| 31 |
B |
3092 |
2963 |
110.77 |
|
|
|
| 32 |
B |
3086 |
2958 |
50.38 |
|
|
|
| 33 |
B |
3076 |
2948 |
10.81 |
|
|
|
| 34 |
B |
3045 |
2918 |
50.72 |
|
|
|
| 35 |
B |
3037 |
2911 |
25.38 |
|
|
|
| 36 |
B |
3032 |
2906 |
26.16 |
|
|
|
| 37 |
B |
1486 |
1424 |
20.36 |
|
|
|
| 38 |
B |
1472 |
1410 |
9.87 |
|
|
|
| 39 |
B |
1463 |
1402 |
0.89 |
|
|
|
| 40 |
B |
1388 |
1330 |
0.46 |
|
|
|
| 41 |
B |
1380 |
1322 |
0.01 |
|
|
|
| 42 |
B |
1373 |
1316 |
0.86 |
|
|
|
| 43 |
B |
1340 |
1284 |
0.14 |
|
|
|
| 44 |
B |
1302 |
1248 |
1.17 |
|
|
|
| 45 |
B |
1226 |
1175 |
0.57 |
|
|
|
| 46 |
B |
1202 |
1152 |
1.05 |
|
|
|
| 47 |
B |
1156 |
1107 |
0.53 |
|
|
|
| 48 |
B |
1036 |
993 |
0.32 |
|
|
|
| 49 |
B |
1023 |
981 |
0.62 |
|
|
|
| 50 |
B |
965 |
924 |
2.56 |
|
|
|
| 51 |
B |
870 |
834 |
0.30 |
|
|
|
| 52 |
B |
820 |
786 |
3.36 |
|
|
|
| 53 |
B |
733 |
702 |
1.34 |
|
|
|
| 54 |
B |
485 |
465 |
0.40 |
|
|
|
| 55 |
B |
334 |
320 |
0.03 |
|
|
|
| 56 |
B |
272 |
261 |
0.09 |
|
|
|
| 57 |
B |
116 |
111 |
0.03 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 43736.4 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 41912.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.341 |
|
|
|
| 2 |
C |
0.472 |
|
|
|
| 3 |
C |
0.472 |
|
|
|
| 4 |
C |
0.787 |
|
|
|
| 5 |
C |
0.787 |
|
|
|
| 6 |
C |
0.393 |
|
|
|
| 7 |
C |
0.393 |
|
|
|
| 8 |
H |
-0.204 |
|
|
|
| 9 |
H |
-0.204 |
|
|
|
| 10 |
H |
-0.294 |
|
|
|
| 11 |
H |
-0.294 |
|
|
|
| 12 |
H |
-0.283 |
|
|
|
| 13 |
H |
-0.283 |
|
|
|
| 14 |
H |
-0.300 |
|
|
|
| 15 |
H |
-0.300 |
|
|
|
| 16 |
H |
-0.264 |
|
|
|
| 17 |
H |
-0.264 |
|
|
|
| 18 |
H |
-0.227 |
|
|
|
| 19 |
H |
-0.227 |
|
|
|
| 20 |
H |
-0.250 |
|
|
|
| 21 |
H |
-0.250 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-0.011 |
0.011 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-46.608 |
-0.183 |
0.000 |
| y |
-0.183 |
-46.732 |
0.000 |
| z |
0.000 |
0.000 |
-47.210 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.363 |
-0.183 |
0.000 |
| y |
-0.183 |
0.176 |
0.000 |
| z |
0.000 |
0.000 |
-0.540 |
|
| Polar |
|---|
| 3z2-r2 | -1.079 |
|---|
| x2-y2 | 0.125 |
|---|
| xy | -0.183 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.125 |
0.476 |
0.000 |
| y |
0.476 |
12.697 |
0.000 |
| z |
0.000 |
0.000 |
13.122 |
<r2> (average value of r
2) Å
2
| <r2> |
228.067 |
| (<r2>)1/2 |
15.102 |