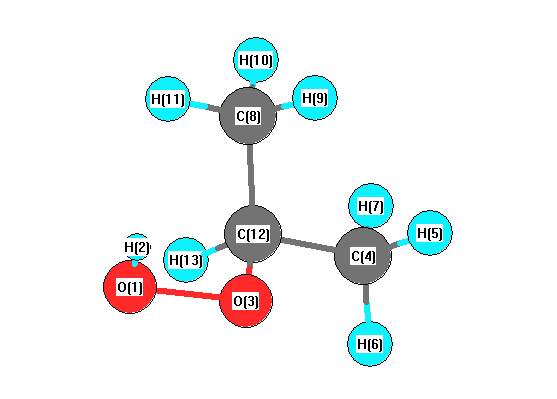
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3817 |
3658 |
35.82 |
|
|
|
| 2 |
A |
3164 |
3032 |
17.68 |
|
|
|
| 3 |
A |
3158 |
3026 |
17.98 |
|
|
|
| 4 |
A |
3153 |
3021 |
33.69 |
|
|
|
| 5 |
A |
3141 |
3010 |
13.07 |
|
|
|
| 6 |
A |
3067 |
2939 |
15.97 |
|
|
|
| 7 |
A |
3059 |
2931 |
21.55 |
|
|
|
| 8 |
A |
3055 |
2928 |
17.29 |
|
|
|
| 9 |
A |
1493 |
1431 |
7.73 |
|
|
|
| 10 |
A |
1473 |
1411 |
3.94 |
|
|
|
| 11 |
A |
1464 |
1403 |
1.42 |
|
|
|
| 12 |
A |
1460 |
1399 |
1.79 |
|
|
|
| 13 |
A |
1404 |
1345 |
3.26 |
|
|
|
| 14 |
A |
1396 |
1338 |
28.72 |
|
|
|
| 15 |
A |
1387 |
1329 |
43.14 |
|
|
|
| 16 |
A |
1360 |
1303 |
10.39 |
|
|
|
| 17 |
A |
1337 |
1281 |
14.77 |
|
|
|
| 18 |
A |
1190 |
1140 |
3.32 |
|
|
|
| 19 |
A |
1182 |
1133 |
29.34 |
|
|
|
| 20 |
A |
1147 |
1099 |
16.22 |
|
|
|
| 21 |
A |
995 |
954 |
3.60 |
|
|
|
| 22 |
A |
950 |
911 |
4.32 |
|
|
|
| 23 |
A |
925 |
887 |
4.50 |
|
|
|
| 24 |
A |
919 |
881 |
4.26 |
|
|
|
| 25 |
A |
836 |
801 |
3.42 |
|
|
|
| 26 |
A |
510 |
488 |
5.28 |
|
|
|
| 27 |
A |
469 |
450 |
4.55 |
|
|
|
| 28 |
A |
354 |
339 |
0.52 |
|
|
|
| 29 |
A |
290 |
278 |
4.94 |
|
|
|
| 30 |
A |
260 |
250 |
62.93 |
|
|
|
| 31 |
A |
226 |
216 |
52.45 |
|
|
|
| 32 |
A |
193 |
185 |
3.75 |
|
|
|
| 33 |
A |
132 |
126 |
5.12 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24482.0 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 23461.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.241 |
|
|
|
| 2 |
H |
0.177 |
|
|
|
| 3 |
O |
-0.491 |
|
|
|
| 4 |
C |
0.423 |
|
|
|
| 5 |
H |
-0.072 |
|
|
|
| 6 |
H |
-0.121 |
|
|
|
| 7 |
H |
-0.064 |
|
|
|
| 8 |
C |
0.421 |
|
|
|
| 9 |
H |
-0.039 |
|
|
|
| 10 |
H |
-0.095 |
|
|
|
| 11 |
H |
-0.168 |
|
|
|
| 12 |
C |
0.600 |
|
|
|
| 13 |
H |
-0.330 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.947 |
1.501 |
0.471 |
1.836 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-29.983 |
-2.661 |
-2.952 |
| y |
-2.661 |
-32.762 |
0.745 |
| z |
-2.952 |
0.745 |
-31.264 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.030 |
-2.661 |
-2.952 |
| y |
-2.661 |
-2.139 |
0.745 |
| z |
-2.952 |
0.745 |
0.109 |
|
| Polar |
|---|
| 3z2-r2 | 0.219 |
|---|
| x2-y2 | 2.779 |
|---|
| xy | -2.661 |
|---|
| xz | -2.952 |
|---|
| yz | 0.745 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.747 |
-0.319 |
-0.004 |
| y |
-0.319 |
7.458 |
0.023 |
| z |
-0.004 |
0.023 |
6.687 |
<r2> (average value of r
2) Å
2
| <r2> |
132.123 |
| (<r2>)1/2 |
11.494 |