return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3SO2NH2 (methanesulfonamide)

using model chemistry: QCISD(T)=FULL/aug-cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at QCISD(T)=FULL/aug-cc-pVDZ
 hartrees
Energy at 0K-643.416995
Energy at 298.15K-643.424579
HF Energy-642.435434
Nuclear repulsion energy272.391418
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)=FULL/aug-cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3459 3356        
2 A' 3175 3080        
3 A' 3056 2964        
4 A' 1569 1522        
5 A' 1432 1389        
6 A' 1318 1278        
7 A' 1072 1040        
8 A' 965 936        
9 A' 900 873        
10 A' 735 713        
11 A' 680 660        
12 A' 468 454        
13 A' 439 426        
14 A' 280 271        
15 A" 3568 3461        
16 A" 3182 3087        
17 A" 1432 1389        
18 A" 1272 1234        
19 A" 1096 1063        
20 A" 950 922        
21 A" 368 357        
22 A" 311 301        
23 A" 202 196        
24 A" 162 157        

Unscaled Zero Point Vibrational Energy (zpe) 16043.4 cm-1
Scaled (by 0.9701) Zero Point Vibrational Energy (zpe) 15563.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD(T)=FULL/aug-cc-pVDZ
ABC
0.14966 0.14100 0.13821

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD(T)=FULL/aug-cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -1.694 -0.041 0.000
S2 0.104 -0.138 0.000
N3 0.537 1.525 0.000
O4 0.537 -0.717 1.305
O5 0.537 -0.717 -1.305
H6 -2.053 -1.082 0.000
H7 -2.002 0.492 0.911
H8 -2.002 0.492 -0.911
H9 1.110 1.678 0.835
H10 1.110 1.678 -0.835

Atom - Atom Distances (Å)
  C1 S2 N3 O4 O5 H6 H7 H8 H9 H10
C11.79972.72532.67072.67071.10111.09991.09993.39303.3930
S21.79971.71841.49181.49182.35422.37942.37942.23752.2375
N32.72531.71842.59442.59443.67472.88882.88881.02451.0245
O42.67071.49182.59442.60932.92252.83953.58032.50703.2625
O52.67071.49182.59442.60932.92253.58032.83953.26252.5070
H61.10112.35423.67472.92252.92251.81921.81924.28004.2800
H71.09992.37942.88882.83953.58031.81921.82253.33163.7608
H81.09992.37942.88883.58032.83951.81921.82253.76083.3316
H93.39302.23751.02452.50703.26254.28003.33163.76081.6705
H103.39302.23751.02453.26252.50704.28003.76083.33161.6705

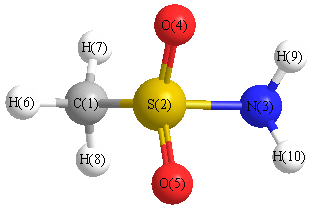
picture of methanesulfonamide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 S2 N3 101.526 C1 S2 O4 108.100
C1 S2 O5 108.100 S2 C1 H6 105.983
S2 C1 H7 107.841 S2 C1 H8 107.841
S2 N3 H9 106.564 S2 N3 H10 106.564
N3 S2 O4 107.627 N3 S2 O5 107.627
O4 S2 O5 121.977 H6 C1 H7 111.488
H6 C1 H8 111.488 H7 C1 H8 111.886
H9 N3 H10 109.224
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability