return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3SO2NH2 (methanesulfonamide)

using model chemistry: B2PLYP=FULLultrafine/aug-cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at B2PLYP=FULLultrafine/aug-cc-pVDZ
 hartrees
Energy at 0K-644.138178
Energy at 298.15K 
HF Energy-643.831182
Nuclear repulsion energy272.954181
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULLultrafine/aug-cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3490 3359 30.49      
2 A' 3205 3084 0.09      
3 A' 3086 2969 0.27      
4 A' 1569 1510 30.07      
5 A' 1435 1381 5.53      
6 A' 1323 1273 8.50      
7 A' 1077 1036 158.59      
8 A' 972 935 17.39      
9 A' 879 845 68.02      
10 A' 719 692 107.54      
11 A' 675 650 96.54      
12 A' 467 449 45.04      
13 A' 438 422 10.56      
14 A' 280 269 5.26      
15 A" 3602 3467 45.33      
16 A" 3212 3091 0.09      
17 A" 1436 1381 2.70      
18 A" 1275 1227 223.64      
19 A" 1090 1049 16.40      
20 A" 958 921 0.39      
21 A" 369 355 0.00      
22 A" 312 300 3.04      
23 A" 204 196 7.60      
24 A" 172 166 30.97      

Unscaled Zero Point Vibrational Energy (zpe) 16121.5 cm-1
Scaled (by 0.9623) Zero Point Vibrational Energy (zpe) 15513.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP=FULLultrafine/aug-cc-pVDZ
ABC
0.15053 0.14140 0.13852

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP=FULLultrafine/aug-cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -1.693 -0.047 0.000
S2 0.103 -0.136 0.000
N3 0.538 1.520 0.000
O4 0.538 -0.715 1.301
O5 0.538 -0.715 -1.301
H6 -2.047 -1.083 0.000
H7 -2.003 0.481 0.906
H8 -2.003 0.481 -0.906
H9 1.097 1.691 0.835
H10 1.097 1.691 -0.835

Atom - Atom Distances (Å)
  C1 S2 N3 O4 O5 H6 H7 H8 H9 H10
C11.79832.72602.66712.66711.09491.09381.09383.39113.3911
S21.79831.71251.48871.48872.34912.37492.37492.24122.2412
N32.72601.71252.58602.58603.66822.89102.89101.01961.0196
O42.66711.48872.58602.60272.91672.83573.57192.51293.2654
O52.66711.48872.58602.60272.91673.57192.83573.26542.5129
H61.09492.34913.66822.91672.91671.80831.80834.27474.2747
H71.09382.37492.89102.83573.57191.80831.81253.32863.7560
H81.09382.37492.89103.57192.83571.80831.81253.75603.3286
H93.39112.24121.01962.51293.26544.27473.32863.75601.6706
H103.39112.24121.01963.26542.51294.27473.75603.32861.6706

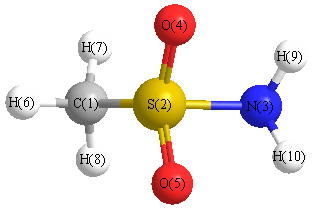
picture of methanesulfonamide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 S2 N3 101.846 C1 S2 O4 108.098
C1 S2 O5 108.098 S2 C1 H6 106.003
S2 C1 H7 107.909 S2 C1 H8 107.909
S2 N3 H9 107.510 S2 N3 H10 107.510
N3 S2 O4 107.555 N3 S2 O5 107.555
O4 S2 O5 121.886 H6 C1 H7 111.416
H6 C1 H8 111.416 H7 C1 H8 111.885
H9 N3 H10 110.023
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability