return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H6 (Trimethylenecycopropane)

using model chemistry: MP4/aug-cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes D3H 1A'
Energy calculated at MP4/aug-cc-pVDZ
 hartrees
Energy at 0K-231.476683
Energy at 298.15K-231.480910
HF Energy-230.574869
Nuclear repulsion energy186.506335
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A1' 3161 3088        
2 A1' 1875 1832        
3 A1' 1493 1459        
4 A1' 789 771        
5 A1" 741 724        
6 A2' 3250 3175        
7 A2' 1099 1074        
8 A2' 503 491        
9 A2" 851 831        
10 A2" 196 192        
11 E' 3251 3176        
11 E' 3251 3176        
12 E' 3161 3088        
12 E' 3161 3088        
13 E' 1670 1631        
13 E' 1670 1631        
14 E' 1457 1423        
14 E' 1457 1423        
15 E' 1136 1110        
15 E' 1136 1109        
16 E' 800 781        
16 E' 799 781        
17 E' 220 215        
17 E' 219 214        
18 E" 844 824        
18 E" 844 824        
19 E" 745 728        
19 E" 745 728        
20 E" 316 308        
20 E" 316 308        

Unscaled Zero Point Vibrational Energy (zpe) 20575.3 cm-1
Scaled (by 0.9769) Zero Point Vibrational Energy (zpe) 20100.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP4/aug-cc-pVDZ
ABC
0.13380 0.13380 0.06690

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP4/aug-cc-pVDZ

Point Group is D3h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.846 0.000
C2 0.733 -0.423 0.000
C3 -0.733 -0.423 0.000
C4 0.000 2.203 0.000
C5 1.908 -1.101 0.000
C6 -1.908 -1.101 0.000
H7 -0.941 2.764 0.000
H8 0.941 2.764 0.000
H9 2.864 -0.567 0.000
H10 1.923 -2.197 0.000
H11 -1.923 -2.197 0.000
H12 -2.864 -0.567 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12
C11.46541.46541.35652.72602.72602.13662.13663.19413.60013.60013.1941
C21.46541.46542.72601.35652.72603.60013.19412.13662.13663.19413.6001
C31.46541.46542.72602.72601.35653.19413.60013.60013.19412.13662.1366
C41.35652.72602.72603.81503.81501.09601.09603.98454.80184.80183.9845
C52.72601.35652.72603.81503.81504.80183.98451.09601.09603.98454.8018
C62.72602.72601.35653.81503.81503.98454.80184.80183.98451.09601.0960
H72.13663.60013.19411.09604.80183.98451.88255.05775.72905.05773.8465
H82.13663.19413.60011.09603.98454.80181.88253.84655.05775.72905.0577
H93.19412.13663.60013.98451.09604.80185.05773.84651.88255.05775.7290
H103.60012.13663.19414.80181.09603.98455.72905.05771.88253.84655.0577
H113.60013.19412.13664.80183.98451.09605.05775.72905.05773.84651.8825
H123.19413.60012.13663.98454.80181.09603.84655.05775.72905.05771.8825

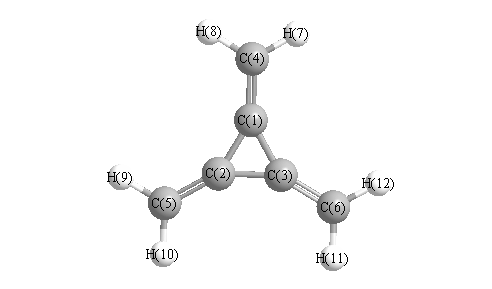
picture of Trimethylenecycopropane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 60.000 C1 C2 C5 150.000
C1 C3 C2 60.000 C1 C3 C6 150.000
C1 C4 H7 120.823 C1 C4 H8 120.823
C2 C1 C3 60.000 C2 C1 C4 150.000
C2 C3 C6 150.000 C2 C5 H9 120.823
C2 C5 H10 120.823 C3 C1 C4 150.000
C3 C2 C5 150.000 C3 C6 H11 120.823
C3 C6 H12 120.823 H7 C4 H8 118.354
H9 C5 H10 118.354 H11 C6 H12 118.354
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability