return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H6 (Propellane)

using model chemistry: MP4/aug-cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes D3H 1A1
Energy calculated at MP4/aug-cc-pVDZ
 hartrees
Energy at 0K-193.495281
Energy at 298.15K-193.501657
HF Energy-192.709747
Nuclear repulsion energy161.499061
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A1' 3146 3073        
2 A1' 1571 1535        
3 A1' 1150 1123        
4 A1' 916 895        
5 A1" 931 910        
6 A2' 3226 3152        
7 A2' 985 962        
8 A2" 1147 1120        
9 A2" 653 638        
10 E' 3229 3155        
10 E' 3229 3155        
11 E' 3142 3069        
11 E' 3142 3069        
12 E' 1525 1490        
12 E' 1525 1490        
13 E' 1239 1210        
13 E' 1239 1210        
14 E' 1122 1096        
14 E' 545 532        
15 E' 544 531        
15 E" 1166 1139        
16 E" 1166 1139        
16 E" 1122 1096        
17 E" 1095 1070        
17 E" 1095 1070        
18 E" 760 743        
18 E" 760 743        

Unscaled Zero Point Vibrational Energy (zpe) 20683.5 cm-1
Scaled (by 0.9769) Zero Point Vibrational Energy (zpe) 20205.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP4/aug-cc-pVDZ
ABC
0.28068 0.28068 0.19111

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP4/aug-cc-pVDZ

Point Group is D3h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 1.306 0.000
C2 0.000 0.000 0.815
C3 1.131 -0.653 0.000
C4 -1.131 -0.653 0.000
C5 0.000 0.000 -0.815
H6 0.928 1.891 0.000
H7 -0.928 1.891 0.000
H8 1.174 -1.750 0.000
H9 2.102 -0.142 0.000
H10 -2.102 -0.142 0.000
H11 -1.174 -1.750 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11
C11.53932.26142.26141.53931.09761.09763.27292.55222.55223.2729
C21.53931.53931.53931.63082.25912.25912.25912.25912.25912.2591
C32.26141.53932.26141.53932.55223.27291.09761.09763.27292.5522
C42.26141.53932.26141.53933.27292.55222.55223.27291.09761.0976
C51.53931.63081.53931.53932.25912.25912.25912.25912.25912.2591
H61.09762.25912.55223.27292.25911.85643.64922.34783.64924.2041
H71.09762.25913.27292.55222.25911.85644.20413.64922.34783.6492
H83.27292.25911.09762.55222.25913.64924.20411.85643.64922.3478
H92.55222.25911.09763.27292.25912.34783.64921.85644.20413.6492
H102.55222.25913.27291.09762.25913.64922.34783.64924.20411.8564
H113.27292.25912.55221.09762.25914.20413.64922.34783.64921.8564

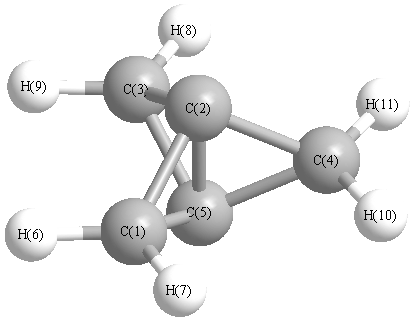
picture of Propellane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 94.538 C1 C2 C4 94.538
C1 C2 C5 58.014 C1 C5 C2 58.014
C1 C5 C3 94.538 C1 C5 C4 94.538
C2 C1 C5 63.971 C2 C1 H6 116.915
C2 C1 H7 116.915 C2 C3 C5 63.971
C2 C3 H8 116.915 C2 C3 H9 116.915
C2 C4 C5 63.971 C2 C4 H10 116.915
C2 C4 H11 116.915 C3 C2 C4 94.538
C3 C2 C5 58.014 C3 C5 C4 94.538
C4 C2 C5 58.014 C5 C1 H6 116.915
C5 C1 H7 116.915 C5 C3 H8 116.915
C5 C3 H9 116.915 C5 C4 H10 116.915
C5 C4 H11 116.915 H6 C1 H7 115.490
H8 C3 H9 115.490 H10 C4 H11 115.490
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability