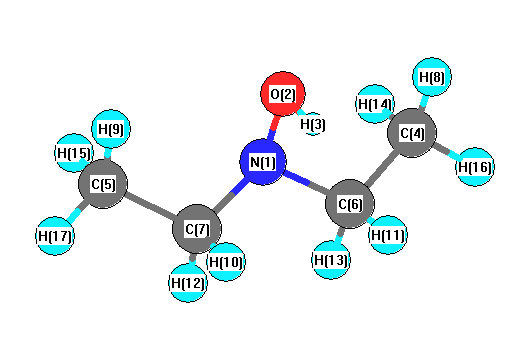
Vibrational Frequencies calculated at HF/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
4015 |
3656 |
5.15 |
|
|
|
| 2 |
A |
3269 |
2977 |
85.12 |
|
|
|
| 3 |
A |
3242 |
2952 |
67.11 |
|
|
|
| 4 |
A |
3232 |
2943 |
30.24 |
|
|
|
| 5 |
A |
3177 |
2893 |
2.22 |
|
|
|
| 6 |
A |
3075 |
2800 |
167.15 |
|
|
|
| 7 |
A |
1617 |
1472 |
11.18 |
|
|
|
| 8 |
A |
1595 |
1452 |
1.87 |
|
|
|
| 9 |
A |
1583 |
1441 |
11.87 |
|
|
|
| 10 |
A |
1556 |
1417 |
21.37 |
|
|
|
| 11 |
A |
1528 |
1392 |
40.24 |
|
|
|
| 12 |
A |
1501 |
1367 |
0.36 |
|
|
|
| 13 |
A |
1419 |
1292 |
7.74 |
|
|
|
| 14 |
A |
1319 |
1201 |
4.11 |
|
|
|
| 15 |
A |
1263 |
1151 |
12.53 |
|
|
|
| 16 |
A |
1172 |
1067 |
2.24 |
|
|
|
| 17 |
A |
1065 |
970 |
19.00 |
|
|
|
| 18 |
A |
885 |
806 |
3.36 |
|
|
|
| 19 |
A |
860 |
783 |
15.45 |
|
|
|
| 20 |
A |
497 |
452 |
4.05 |
|
|
|
| 21 |
A |
452 |
412 |
0.04 |
|
|
|
| 22 |
A |
227 |
207 |
0.22 |
|
|
|
| 23 |
A |
224 |
204 |
2.68 |
|
|
|
| 24 |
A |
109 |
99 |
0.46 |
|
|
|
| 25 |
A |
3268 |
2975 |
0.13 |
|
|
|
| 26 |
A |
3241 |
2952 |
19.29 |
|
|
|
| 27 |
A |
3214 |
2927 |
0.01 |
|
|
|
| 28 |
A |
3176 |
2892 |
72.27 |
|
|
|
| 29 |
A |
3061 |
2787 |
3.03 |
|
|
|
| 30 |
A |
1607 |
1464 |
0.58 |
|
|
|
| 31 |
A |
1588 |
1446 |
2.60 |
|
|
|
| 32 |
A |
1582 |
1440 |
0.17 |
|
|
|
| 33 |
A |
1530 |
1394 |
36.40 |
|
|
|
| 34 |
A |
1496 |
1362 |
13.14 |
|
|
|
| 35 |
A |
1401 |
1275 |
1.65 |
|
|
|
| 36 |
A |
1308 |
1191 |
4.12 |
|
|
|
| 37 |
A |
1187 |
1081 |
14.76 |
|
|
|
| 38 |
A |
1162 |
1059 |
8.92 |
|
|
|
| 39 |
A |
1014 |
923 |
0.01 |
|
|
|
| 40 |
A |
859 |
782 |
0.58 |
|
|
|
| 41 |
A |
585 |
533 |
22.72 |
|
|
|
| 42 |
A |
440 |
401 |
115.84 |
|
|
|
| 43 |
A |
368 |
335 |
6.64 |
|
|
|
| 44 |
A |
264 |
241 |
0.03 |
|
|
|
| 45 |
A |
131 |
119 |
0.12 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35680.2 cm
-1
Scaled (by 0.9106) Zero Point Vibrational Energy (zpe) 32490.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.368 |
|
|
|
| 2 |
O |
-0.679 |
|
|
|
| 3 |
H |
0.150 |
|
|
|
| 4 |
C |
0.438 |
|
|
|
| 5 |
C |
0.438 |
|
|
|
| 6 |
C |
0.602 |
|
|
|
| 7 |
C |
0.602 |
|
|
|
| 8 |
H |
-0.052 |
|
|
|
| 9 |
H |
-0.052 |
|
|
|
| 10 |
H |
-0.183 |
|
|
|
| 11 |
H |
-0.183 |
|
|
|
| 12 |
H |
-0.277 |
|
|
|
| 13 |
H |
-0.277 |
|
|
|
| 14 |
H |
-0.050 |
|
|
|
| 15 |
H |
-0.050 |
|
|
|
| 16 |
H |
-0.031 |
|
|
|
| 17 |
H |
-0.031 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.397 |
-1.961 |
0.000 |
2.407 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.974 |
2.400 |
0.000 |
| y |
2.400 |
-38.792 |
0.000 |
| z |
0.000 |
0.000 |
-38.429 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.363 |
2.400 |
0.000 |
| y |
2.400 |
0.409 |
0.000 |
| z |
0.000 |
0.000 |
0.954 |
|
| Polar |
|---|
| 3z2-r2 | 1.908 |
|---|
| x2-y2 | -1.181 |
|---|
| xy | 2.400 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
210.979 |
| (<r2>)1/2 |
14.525 |