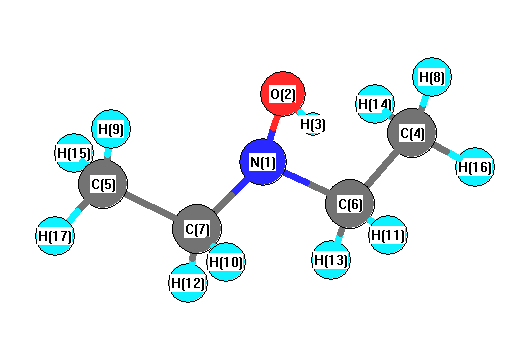
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3631 |
3479 |
3.47 |
|
|
|
| 2 |
A |
3167 |
3035 |
38.31 |
|
|
|
| 3 |
A |
3152 |
3020 |
33.99 |
|
|
|
| 4 |
A |
3107 |
2978 |
31.20 |
|
|
|
| 5 |
A |
3070 |
2942 |
3.97 |
|
|
|
| 6 |
A |
2930 |
2808 |
157.32 |
|
|
|
| 7 |
A |
1492 |
1430 |
12.32 |
|
|
|
| 8 |
A |
1470 |
1409 |
1.07 |
|
|
|
| 9 |
A |
1464 |
1403 |
21.54 |
|
|
|
| 10 |
A |
1428 |
1369 |
35.64 |
|
|
|
| 11 |
A |
1411 |
1353 |
19.01 |
|
|
|
| 12 |
A |
1377 |
1319 |
0.96 |
|
|
|
| 13 |
A |
1301 |
1247 |
4.23 |
|
|
|
| 14 |
A |
1194 |
1144 |
3.86 |
|
|
|
| 15 |
A |
1163 |
1114 |
12.52 |
|
|
|
| 16 |
A |
1107 |
1061 |
2.47 |
|
|
|
| 17 |
A |
972 |
932 |
19.05 |
|
|
|
| 18 |
A |
823 |
789 |
2.99 |
|
|
|
| 19 |
A |
795 |
762 |
15.85 |
|
|
|
| 20 |
A |
456 |
437 |
4.03 |
|
|
|
| 21 |
A |
427 |
409 |
0.25 |
|
|
|
| 22 |
A |
210 |
201 |
2.04 |
|
|
|
| 23 |
A |
207 |
199 |
0.62 |
|
|
|
| 24 |
A |
110 |
106 |
0.40 |
|
|
|
| 25 |
A |
3167 |
3034 |
0.00 |
|
|
|
| 26 |
A |
3152 |
3020 |
12.63 |
|
|
|
| 27 |
A |
3095 |
2966 |
0.39 |
|
|
|
| 28 |
A |
3069 |
2941 |
48.18 |
|
|
|
| 29 |
A |
2918 |
2796 |
5.90 |
|
|
|
| 30 |
A |
1488 |
1426 |
0.35 |
|
|
|
| 31 |
A |
1464 |
1403 |
0.83 |
|
|
|
| 32 |
A |
1457 |
1396 |
1.08 |
|
|
|
| 33 |
A |
1397 |
1339 |
33.73 |
|
|
|
| 34 |
A |
1370 |
1313 |
11.19 |
|
|
|
| 35 |
A |
1294 |
1240 |
1.65 |
|
|
|
| 36 |
A |
1211 |
1160 |
4.08 |
|
|
|
| 37 |
A |
1101 |
1055 |
5.28 |
|
|
|
| 38 |
A |
1078 |
1033 |
15.44 |
|
|
|
| 39 |
A |
968 |
928 |
0.28 |
|
|
|
| 40 |
A |
799 |
765 |
0.10 |
|
|
|
| 41 |
A |
550 |
527 |
20.61 |
|
|
|
| 42 |
A |
391 |
375 |
99.44 |
|
|
|
| 43 |
A |
342 |
328 |
1.26 |
|
|
|
| 44 |
A |
240 |
230 |
0.00 |
|
|
|
| 45 |
A |
133 |
127 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 33572.9 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 32172.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.153 |
|
|
|
| 2 |
O |
-0.532 |
|
|
|
| 3 |
H |
0.139 |
|
|
|
| 4 |
C |
0.497 |
|
|
|
| 5 |
C |
0.497 |
|
|
|
| 6 |
C |
0.771 |
|
|
|
| 7 |
C |
0.771 |
|
|
|
| 8 |
H |
-0.107 |
|
|
|
| 9 |
H |
-0.107 |
|
|
|
| 10 |
H |
-0.294 |
|
|
|
| 11 |
H |
-0.294 |
|
|
|
| 12 |
H |
-0.400 |
|
|
|
| 13 |
H |
-0.400 |
|
|
|
| 14 |
H |
-0.106 |
|
|
|
| 15 |
H |
-0.106 |
|
|
|
| 16 |
H |
-0.087 |
|
|
|
| 17 |
H |
-0.087 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.073 |
2.221 |
0.000 |
2.222 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.331 |
0.745 |
0.000 |
| y |
0.745 |
-41.541 |
0.000 |
| z |
0.000 |
0.000 |
-38.081 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.480 |
0.745 |
0.000 |
| y |
0.745 |
-3.835 |
0.000 |
| z |
0.000 |
0.000 |
1.355 |
|
| Polar |
|---|
| 3z2-r2 | 2.709 |
|---|
| x2-y2 | 4.210 |
|---|
| xy | 0.745 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.555 |
-0.016 |
0.000 |
| y |
-0.016 |
9.416 |
0.000 |
| z |
0.000 |
0.000 |
11.680 |
<r2> (average value of r
2) Å
2
| <r2> |
210.950 |
| (<r2>)1/2 |
14.524 |