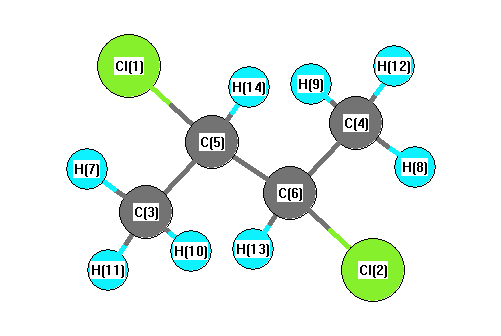
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3174 |
3042 |
0.00 |
|
|
|
| 2 |
Ag |
3165 |
3033 |
0.00 |
|
|
|
| 3 |
Ag |
3118 |
2988 |
0.00 |
|
|
|
| 4 |
Ag |
3075 |
2947 |
0.00 |
|
|
|
| 5 |
Ag |
1472 |
1411 |
0.00 |
|
|
|
| 6 |
Ag |
1470 |
1408 |
0.00 |
|
|
|
| 7 |
Ag |
1403 |
1345 |
0.00 |
|
|
|
| 8 |
Ag |
1382 |
1324 |
0.00 |
|
|
|
| 9 |
Ag |
1266 |
1213 |
0.00 |
|
|
|
| 10 |
Ag |
1183 |
1134 |
0.00 |
|
|
|
| 11 |
Ag |
1152 |
1104 |
0.00 |
|
|
|
| 12 |
Ag |
1025 |
982 |
0.00 |
|
|
|
| 13 |
Ag |
861 |
825 |
0.00 |
|
|
|
| 14 |
Ag |
706 |
676 |
0.00 |
|
|
|
| 15 |
Ag |
480 |
460 |
0.00 |
|
|
|
| 16 |
Ag |
345 |
331 |
0.00 |
|
|
|
| 17 |
Ag |
283 |
272 |
0.00 |
|
|
|
| 18 |
Ag |
229 |
220 |
0.00 |
|
|
|
| 19 |
Au |
3175 |
3043 |
15.69 |
|
|
|
| 20 |
Au |
3167 |
3035 |
24.37 |
|
|
|
| 21 |
Au |
3133 |
3003 |
5.32 |
|
|
|
| 22 |
Au |
3075 |
2946 |
18.50 |
|
|
|
| 23 |
Au |
1472 |
1411 |
16.98 |
|
|
|
| 24 |
Au |
1465 |
1404 |
10.76 |
|
|
|
| 25 |
Au |
1397 |
1339 |
25.51 |
|
|
|
| 26 |
Au |
1307 |
1253 |
3.18 |
|
|
|
| 27 |
Au |
1208 |
1158 |
34.06 |
|
|
|
| 28 |
Au |
1101 |
1055 |
4.96 |
|
|
|
| 29 |
Au |
1021 |
979 |
42.13 |
|
|
|
| 30 |
Au |
974 |
934 |
12.66 |
|
|
|
| 31 |
Au |
665 |
638 |
79.07 |
|
|
|
| 32 |
Au |
359 |
344 |
3.39 |
|
|
|
| 33 |
Au |
333 |
320 |
2.57 |
|
|
|
| 34 |
Au |
242 |
232 |
3.07 |
|
|
|
| 35 |
Au |
203 |
194 |
3.07 |
|
|
|
| 36 |
Au |
67 |
64 |
3.06 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25077.1 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 24031.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
Cl |
-0.122 |
|
|
|
| 2 |
Cl |
-0.122 |
|
|
|
| 3 |
C |
0.708 |
|
|
|
| 4 |
C |
0.708 |
|
|
|
| 5 |
C |
-0.091 |
|
|
|
| 6 |
C |
-0.091 |
|
|
|
| 7 |
H |
-0.096 |
|
|
|
| 8 |
H |
-0.096 |
|
|
|
| 9 |
H |
-0.088 |
|
|
|
| 10 |
H |
-0.088 |
|
|
|
| 11 |
H |
-0.062 |
|
|
|
| 12 |
H |
-0.062 |
|
|
|
| 13 |
H |
-0.248 |
|
|
|
| 14 |
H |
-0.248 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-50.532 |
1.987 |
-3.652 |
| y |
1.987 |
-53.288 |
3.527 |
| z |
-3.652 |
3.527 |
-54.876 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.550 |
1.987 |
-3.652 |
| y |
1.987 |
-0.584 |
3.527 |
| z |
-3.652 |
3.527 |
-2.966 |
|
| Polar |
|---|
| 3z2-r2 | -5.932 |
|---|
| x2-y2 | 2.756 |
|---|
| xy | 1.987 |
|---|
| xz | -3.652 |
|---|
| yz | 3.527 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.922 |
-0.363 |
0.631 |
| y |
-0.363 |
10.998 |
-2.081 |
| z |
0.631 |
-2.081 |
12.352 |
<r2> (average value of r
2) Å
2
| <r2> |
291.070 |
| (<r2>)1/2 |
17.061 |