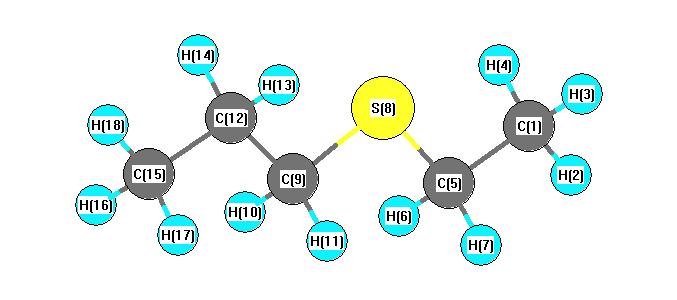
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3143 |
3012 |
21.88 |
|
|
|
| 2 |
A' |
3140 |
3009 |
33.06 |
|
|
|
| 3 |
A' |
3065 |
2938 |
28.88 |
|
|
|
| 4 |
A' |
3061 |
2933 |
28.00 |
|
|
|
| 5 |
A' |
3060 |
2932 |
30.92 |
|
|
|
| 6 |
A' |
3053 |
2926 |
41.06 |
|
|
|
| 7 |
A' |
3048 |
2921 |
12.71 |
|
|
|
| 8 |
A' |
1487 |
1425 |
5.35 |
|
|
|
| 9 |
A' |
1480 |
1419 |
2.88 |
|
|
|
| 10 |
A' |
1472 |
1411 |
1.06 |
|
|
|
| 11 |
A' |
1466 |
1405 |
0.62 |
|
|
|
| 12 |
A' |
1458 |
1397 |
7.37 |
|
|
|
| 13 |
A' |
1397 |
1338 |
2.13 |
|
|
|
| 14 |
A' |
1391 |
1333 |
3.46 |
|
|
|
| 15 |
A' |
1364 |
1307 |
1.59 |
|
|
|
| 16 |
A' |
1288 |
1234 |
14.26 |
|
|
|
| 17 |
A' |
1238 |
1186 |
28.63 |
|
|
|
| 18 |
A' |
1125 |
1078 |
2.47 |
|
|
|
| 19 |
A' |
1078 |
1033 |
1.51 |
|
|
|
| 20 |
A' |
1070 |
1025 |
0.67 |
|
|
|
| 21 |
A' |
1004 |
962 |
6.03 |
|
|
|
| 22 |
A' |
919 |
881 |
2.89 |
|
|
|
| 23 |
A' |
779 |
747 |
2.01 |
|
|
|
| 24 |
A' |
705 |
675 |
0.92 |
|
|
|
| 25 |
A' |
384 |
368 |
0.32 |
|
|
|
| 26 |
A' |
303 |
290 |
0.54 |
|
|
|
| 27 |
A' |
271 |
260 |
0.82 |
|
|
|
| 28 |
A' |
104 |
100 |
0.30 |
|
|
|
| 29 |
A" |
3151 |
3020 |
23.57 |
|
|
|
| 30 |
A" |
3135 |
3005 |
54.71 |
|
|
|
| 31 |
A" |
3114 |
2985 |
0.55 |
|
|
|
| 32 |
A" |
3110 |
2980 |
12.91 |
|
|
|
| 33 |
A" |
3093 |
2964 |
1.88 |
|
|
|
| 34 |
A" |
1477 |
1416 |
7.78 |
|
|
|
| 35 |
A" |
1468 |
1407 |
9.17 |
|
|
|
| 36 |
A" |
1311 |
1257 |
0.31 |
|
|
|
| 37 |
A" |
1255 |
1203 |
0.00 |
|
|
|
| 38 |
A" |
1230 |
1178 |
0.01 |
|
|
|
| 39 |
A" |
1057 |
1013 |
0.65 |
|
|
|
| 40 |
A" |
1029 |
986 |
0.13 |
|
|
|
| 41 |
A" |
862 |
826 |
0.03 |
|
|
|
| 42 |
A" |
783 |
750 |
4.22 |
|
|
|
| 43 |
A" |
741 |
710 |
2.78 |
|
|
|
| 44 |
A" |
241 |
231 |
0.04 |
|
|
|
| 45 |
A" |
236 |
227 |
0.06 |
|
|
|
| 46 |
A" |
110 |
106 |
0.94 |
|
|
|
| 47 |
A" |
62 |
59 |
0.55 |
|
|
|
| 48 |
A" |
41 |
39 |
0.28 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35428.2 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 33950.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.426 |
|
|
|
| 2 |
H |
-0.101 |
|
|
|
| 3 |
H |
-0.114 |
|
|
|
| 4 |
H |
-0.114 |
|
|
|
| 5 |
C |
0.263 |
|
|
|
| 6 |
H |
-0.209 |
|
|
|
| 7 |
H |
-0.209 |
|
|
|
| 8 |
S |
0.254 |
|
|
|
| 9 |
C |
0.127 |
|
|
|
| 10 |
H |
-0.252 |
|
|
|
| 11 |
H |
-0.252 |
|
|
|
| 12 |
C |
0.693 |
|
|
|
| 13 |
H |
-0.244 |
|
|
|
| 14 |
H |
-0.244 |
|
|
|
| 15 |
C |
0.365 |
|
|
|
| 16 |
H |
-0.125 |
|
|
|
| 17 |
H |
-0.125 |
|
|
|
| 18 |
H |
-0.138 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.957 |
-1.309 |
0.000 |
1.621 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-43.963 |
2.530 |
0.000 |
| y |
2.530 |
-47.874 |
0.000 |
| z |
0.000 |
0.000 |
-48.082 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.016 |
2.530 |
0.000 |
| y |
2.530 |
-1.852 |
0.000 |
| z |
0.000 |
0.000 |
-2.164 |
|
| Polar |
|---|
| 3z2-r2 | -4.327 |
|---|
| x2-y2 | 3.912 |
|---|
| xy | 2.530 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
14.388 |
2.596 |
0.000 |
| y |
2.596 |
14.265 |
0.000 |
| z |
0.000 |
0.000 |
10.550 |
<r2> (average value of r
2) Å
2
| <r2> |
353.609 |
| (<r2>)1/2 |
18.804 |