return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3NNCH3 ((E)-1,2-Dimethyldiazene)

using model chemistry: MP4/aug-cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2H 1AG
Energy calculated at MP4/aug-cc-pVDZ
 hartrees
Energy at 0K-188.806414
Energy at 298.15K-188.813739
HF Energy-188.091899
Nuclear repulsion energy118.626369
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 Ag 3156 3083        
2 Ag 3054 2984        
3 Ag 1582 1545        
4 Ag 1511 1476        
5 Ag 1444 1411        
6 Ag 1225 1197        
7 Ag 941 920        
8 Ag 598 584        
9 Au 3147 3074        
10 Au 1519 1483        
11 Au 1151 1124        
12 Au 288 282        
13 Au 143 140        
14 Bg 3147 3074        
15 Bg 1518 1483        
16 Bg 1053 1029        
17 Bg 210 205        
18 Bu 3155 3082        
19 Bu 3054 2983        
20 Bu 1523 1488        
21 Bu 1447 1414        
22 Bu 1157 1130        
23 Bu 1039 1015        
24 Bu 354 346        

Unscaled Zero Point Vibrational Energy (zpe) 18707.5 cm-1
Scaled (by 0.9769) Zero Point Vibrational Energy (zpe) 18275.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP4/aug-cc-pVDZ
ABC
1.34382 0.14589 0.13855

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP4/aug-cc-pVDZ

Point Group is C2h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 0.382 0.503 0.000
N2 -0.382 -0.503 0.000
C3 -0.382 1.771 0.000
C4 0.382 -1.771 0.000
H5 -1.472 1.590 0.000
H6 1.472 -1.590 0.000
H7 -0.078 2.345 0.893
H8 -0.078 2.345 -0.893
H9 0.078 -2.345 0.893
H10 0.078 -2.345 -0.893

Atom - Atom Distances (Å)
  N1 N2 C3 C4 H5 H6 H7 H8 H9 H10
N11.26401.48042.27502.14882.36012.09812.09813.00103.0010
N21.26402.27501.48042.36012.14883.00103.00102.09812.0981
C31.48042.27503.62441.10493.83881.10421.10424.23774.2377
C42.27501.48043.62443.83881.10494.23774.23771.10421.1042
H52.14882.36011.10493.83884.33331.81971.81974.32284.3228
H62.36012.14883.83881.10494.33334.32284.32281.81971.8197
H72.09813.00101.10424.23771.81974.32281.78604.69355.0218
H82.09813.00101.10424.23771.81974.32281.78605.02184.6935
H93.00102.09814.23771.10424.32281.81974.69355.02181.7860
H103.00102.09814.23771.10424.32281.81975.02184.69351.7860

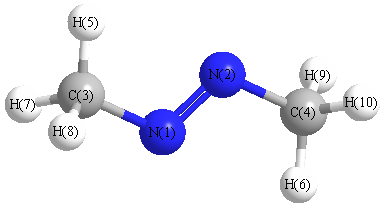
picture of (E)-1,2-Dimethyldiazene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 N2 C4 111.741 N1 C3 H5 111.613
N1 C3 H7 107.644 N1 C3 H8 107.644
N2 N1 C3 111.741 N2 C4 H6 111.613
N2 C4 H9 107.644 N2 C4 H10 107.644
H5 C3 H7 110.917 H5 C3 H8 110.917
H6 C4 H9 110.917 H6 C4 H10 110.917
H7 C3 H8 107.940 H9 C4 H10 107.940
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability