All results from a given calculation for CH3NNCH3 ((Z)-1,2-Dimethyldiazene)
using model chemistry: CCSD=FULL/aug-cc-pVDZ
19 10 17 12 22
States and conformations
| State |
Conformation |
minimum conformation |
conformer description |
state description |
| 1 |
1 |
yes |
C2V |
1A1 |
Energy calculated at CCSD=FULL/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -188.774586 |
| Energy at 298.15K | |
| HF Energy | -188.074857 |
| Nuclear repulsion energy | 121.660707 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD=FULL/aug-cc-pVDZ
Geometric Data calculated at CCSD=FULL/aug-cc-pVDZ
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
0.000 |
0.626 |
-0.790 |
| N2 |
0.000 |
-0.626 |
-0.790 |
| C3 |
0.000 |
1.351 |
0.508 |
| C4 |
0.000 |
-1.351 |
0.508 |
| H5 |
0.000 |
2.425 |
0.281 |
| H6 |
0.000 |
-2.425 |
0.281 |
| H7 |
-0.897 |
1.103 |
1.101 |
| H8 |
0.897 |
1.103 |
1.101 |
| H9 |
0.897 |
-1.103 |
1.101 |
| H10 |
-0.897 |
-1.103 |
1.101 |
Atom - Atom Distances (Å)
| |
N1 |
N2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| N1 | | 1.2520 | 1.4866 | 2.3650 | 2.0936 | 3.2333 | 2.1467 | 2.1467 | 2.7149 | 2.7149 |
N2 | 1.2520 | | 2.3650 | 1.4866 | 3.2333 | 2.0936 | 2.7149 | 2.7149 | 2.1467 | 2.1467 | C3 | 1.4866 | 2.3650 | | 2.7022 | 1.0971 | 3.7825 | 1.1035 | 1.1035 | 2.6794 | 2.6794 | C4 | 2.3650 | 1.4866 | 2.7022 | | 3.7825 | 1.0971 | 2.6794 | 2.6794 | 1.1035 | 1.1035 | H5 | 2.0936 | 3.2333 | 1.0971 | 3.7825 | | 4.8492 | 1.7950 | 1.7950 | 3.7311 | 3.7311 | H6 | 3.2333 | 2.0936 | 3.7825 | 1.0971 | 4.8492 | | 3.7311 | 3.7311 | 1.7950 | 1.7950 | H7 | 2.1467 | 2.7149 | 1.1035 | 2.6794 | 1.7950 | 3.7311 | | 1.7934 | 2.8432 | 2.2063 | H8 | 2.1467 | 2.7149 | 1.1035 | 2.6794 | 1.7950 | 3.7311 | 1.7934 | | 2.2063 | 2.8432 | H9 | 2.7149 | 2.1467 | 2.6794 | 1.1035 | 3.7311 | 1.7950 | 2.8432 | 2.2063 | | 1.7934 | H10 | 2.7149 | 2.1467 | 2.6794 | 1.1035 | 3.7311 | 1.7950 | 2.2063 | 2.8432 | 1.7934 | |
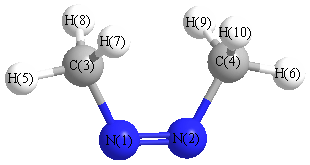 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
N2 |
C4 |
119.192 |
|
N1 |
C3 |
H5 |
107.292 |
| N1 |
C3 |
H7 |
111.092 |
|
N1 |
C3 |
H8 |
111.092 |
| N2 |
N1 |
C3 |
119.192 |
|
N2 |
C4 |
H6 |
107.292 |
| N2 |
C4 |
H9 |
111.092 |
|
N2 |
C4 |
H10 |
111.092 |
| H5 |
C3 |
H7 |
109.316 |
|
H5 |
C3 |
H8 |
109.316 |
| H6 |
C4 |
H9 |
109.316 |
|
H6 |
C4 |
H10 |
109.316 |
| H7 |
C3 |
H8 |
108.701 |
|
H9 |
C4 |
H10 |
108.701 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
