Jump to
S1C2
Energy calculated at HF/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -147.936456 |
| Energy at 298.15K | -147.938886 |
| Nuclear repulsion energy | 59.859566 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3764 |
3428 |
51.90 |
|
|
|
| 2 |
A' |
2577 |
2346 |
147.72 |
|
|
|
| 3 |
A' |
1752 |
1596 |
41.54 |
|
|
|
| 4 |
A' |
1145 |
1043 |
6.68 |
|
|
|
| 5 |
A' |
687 |
625 |
191.01 |
|
|
|
| 6 |
A' |
536 |
489 |
51.56 |
|
|
|
| 7 |
A" |
3869 |
3523 |
73.12 |
|
|
|
| 8 |
A" |
1306 |
1189 |
0.01 |
|
|
|
| 9 |
A" |
462 |
421 |
1.77 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 8048.9 cm
-1
Scaled (by 0.9106) Zero Point Vibrational Energy (zpe) 7329.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.226 |
0.000 |
| N2 |
-0.063 |
1.363 |
0.000 |
| N3 |
0.124 |
-1.114 |
0.000 |
| H4 |
-0.214 |
-1.551 |
0.833 |
| H5 |
-0.214 |
-1.551 |
-0.833 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
N3 |
H4 |
H5 |
| C1 | | 1.1390 | 1.3452 | 1.9741 | 1.9741 |
N2 | 1.1390 | | 2.4838 | 3.0346 | 3.0346 | N3 | 1.3452 | 2.4838 | | 0.9999 | 0.9999 | H4 | 1.9741 | 3.0346 | 0.9999 | | 1.6664 | H5 | 1.9741 | 3.0346 | 0.9999 | 1.6664 | |
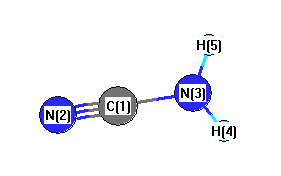 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H4 |
113.844 |
|
C1 |
N3 |
H5 |
113.844 |
| N2 |
C1 |
N3 |
177.868 |
|
H4 |
N3 |
H5 |
112.877 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.212 |
|
|
|
| 2 |
N |
-0.279 |
|
|
|
| 3 |
N |
-0.086 |
|
|
|
| 4 |
H |
0.077 |
|
|
|
| 5 |
H |
0.077 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.910 |
-4.557 |
0.000 |
4.647 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-18.792 |
2.115 |
0.000 |
| y |
2.115 |
-19.114 |
0.000 |
| z |
0.000 |
0.000 |
-15.290 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.590 |
2.115 |
0.000 |
| y |
2.115 |
-2.073 |
0.000 |
| z |
0.000 |
0.000 |
3.663 |
|
| Polar |
|---|
| 3z2-r2 | 7.326 |
|---|
| x2-y2 | 0.322 |
|---|
| xy | 2.115 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.800 |
-0.081 |
0.000 |
| y |
-0.081 |
5.115 |
0.000 |
| z |
0.000 |
0.000 |
2.926 |
<r2> (average value of r
2) Å
2
| <r2> |
39.495 |
| (<r2>)1/2 |
6.285 |
Jump to
S1C1
Energy calculated at HF/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -147.934646 |
| Energy at 298.15K | |
| HF Energy | -147.934646 |
| Nuclear repulsion energy | 60.042313 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3833 |
3490 |
83.94 |
|
|
|
| 2 |
A1 |
2562 |
2333 |
197.10 |
|
|
|
| 3 |
A1 |
1744 |
1588 |
48.32 |
|
|
|
| 4 |
A1 |
1187 |
1081 |
14.01 |
|
|
|
| 5 |
B1 |
592 |
539 |
3.51 |
|
|
|
| 6 |
B1 |
472i |
429i |
275.42 |
|
|
|
| 7 |
B2 |
3971 |
3616 |
112.65 |
|
|
|
| 8 |
B2 |
1235 |
1124 |
1.50 |
|
|
|
| 9 |
B2 |
456 |
416 |
0.16 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 7554.0 cm
-1
Scaled (by 0.9106) Zero Point Vibrational Energy (zpe) 6878.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/aug-cc-pVDZ
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.225 |
| N2 |
0.000 |
0.000 |
1.366 |
| N3 |
0.000 |
0.000 |
-1.102 |
| H4 |
0.000 |
0.860 |
-1.600 |
| H5 |
0.000 |
-0.860 |
-1.600 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
N3 |
H4 |
H5 |
| C1 | | 1.1408 | 1.3275 | 2.0174 | 2.0174 |
N2 | 1.1408 | | 2.4684 | 3.0880 | 3.0880 | N3 | 1.3275 | 2.4684 | | 0.9932 | 0.9932 | H4 | 2.0174 | 3.0880 | 0.9932 | | 1.7193 | H5 | 2.0174 | 3.0880 | 0.9932 | 1.7193 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H4 |
120.063 |
|
C1 |
N3 |
H5 |
120.063 |
| N2 |
C1 |
N3 |
180.000 |
|
H4 |
N3 |
H5 |
119.874 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.190 |
|
|
|
| 2 |
N |
-0.278 |
|
|
|
| 3 |
N |
0.029 |
|
|
|
| 4 |
H |
0.030 |
|
|
|
| 5 |
H |
0.030 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-4.915 |
4.915 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-19.019 |
0.000 |
0.000 |
| y |
0.000 |
-15.011 |
0.000 |
| z |
0.000 |
0.000 |
-18.446 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.290 |
0.000 |
0.000 |
| y |
0.000 |
3.721 |
0.000 |
| z |
0.000 |
0.000 |
-1.431 |
|
| Polar |
|---|
| 3z2-r2 | -2.861 |
|---|
| x2-y2 | -4.007 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.790 |
0.000 |
0.000 |
| y |
0.000 |
2.919 |
0.000 |
| z |
0.000 |
0.000 |
5.185 |
<r2> (average value of r
2) Å
2
| <r2> |
39.395 |
| (<r2>)1/2 |
6.277 |
